return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (2H-Thiopyran, tetrahydro-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-593.689951
Energy at 298.15K-593.702342
HF Energy-592.711689
Nuclear repulsion energy312.959342
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3173 2997 16.65      
2 A 3159 2984 23.16      
3 A 3146 2971 20.75      
4 A 3099 2927 12.98      
5 A 3093 2922 18.39      
6 A 3087 2916 14.61      
7 A 1519 1434 2.00      
8 A 1505 1422 10.71      
9 A 1488 1406 6.59      
10 A 1394 1316 0.85      
11 A 1354 1279 4.79      
12 A 1282 1211 0.82      
13 A 1258 1188 6.05      
14 A 1097 1037 1.76      
15 A 1051 993 0.54      
16 A 992 937 7.92      
17 A 849 802 4.74      
18 A 841 794 0.43      
19 A 691 653 3.07      
20 A 502 475 0.60      
21 A 369 348 0.44      
22 A 342 323 0.16      
23 A 184 173 1.74      
24 A 3174 2998 2.38      
25 A 3154 2979 11.47      
26 A 3103 2931 14.79      
27 A 3094 2922 28.55      
28 A 1497 1414 2.45      
29 A 1484 1402 0.77      
30 A 1402 1324 0.05      
31 A 1381 1305 0.03      
32 A 1313 1241 17.59      
33 A 1304 1232 2.20      
34 A 1177 1112 0.17      
35 A 1124 1062 0.09      
36 A 1108 1046 0.48      
37 A 967 914 0.47      
38 A 923 872 3.73      
39 A 810 765 0.02      
40 A 722 682 0.17      
41 A 398 376 0.27      
42 A 243 229 0.58      

Unscaled Zero Point Vibrational Energy (zpe) 31927.1 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 30155.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.13415 0.10057 0.06426

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 1.503 0.000
C2 0.066 -1.716 0.000
C3 0.392 -0.935 1.273
C4 0.392 -0.935 -1.273
C5 -0.392 0.371 1.360
C6 -0.392 0.371 -1.360
H7 0.614 -2.662 0.000
H8 -1.000 -1.973 0.000
H9 -1.468 0.173 1.360
H10 -1.468 0.173 -1.360
H11 0.157 -1.548 2.149
H12 0.157 -1.548 -2.149
H13 1.462 -0.712 1.307
H14 1.462 -0.712 -1.307
H15 -0.155 0.915 -2.276
H16 -0.155 0.915 2.276

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S13.22012.77822.77821.81201.81204.21033.61682.40262.40263.73543.73542.95832.95832.35592.3559
C23.22011.52891.52892.53282.53281.09331.09642.78822.78822.15792.15792.16022.16023.48623.4862
C32.77821.52892.54621.52543.04142.15722.15282.16693.40871.09503.48501.09392.80234.03952.1742
C42.77821.52892.54623.04141.52542.15722.15283.40872.16693.48501.09502.80231.09392.17424.0395
C51.81202.53281.52543.04142.71913.47282.77691.09452.93122.14644.03682.14743.42343.68371.0919
C61.81202.53283.04141.52542.71913.47282.77692.93121.09454.03682.14643.42342.14741.09193.6837
H74.21031.09332.15722.15723.47283.47281.75553.77193.77192.46402.46402.49612.49614.30914.3091
H83.61681.09642.15282.15282.77692.77691.75552.58342.58342.47762.47763.05943.05943.77283.7728
H92.40262.78822.16693.40871.09452.93123.77192.58342.71932.49544.23293.06154.05983.93621.7649
H102.40262.78823.40872.16692.93121.09453.77192.58342.71934.23292.49544.05983.06151.76493.9362
H113.73542.15791.09503.48502.14644.03682.46402.47762.49544.23294.29891.76433.78835.07432.4858
H123.73542.15793.48501.09504.03682.14642.46402.47764.23292.49544.29893.78831.76432.48585.0743
H132.95832.16021.09392.80232.14743.42342.49613.05943.06154.05981.76433.78832.61424.25432.4898
H142.95832.16022.80231.09393.42342.14742.49613.05944.05983.06153.78831.76432.61422.48984.2543
H152.35593.48624.03952.17423.68371.09194.30913.77283.93621.76495.07432.48584.25432.48984.5521
H162.35593.48622.17424.03951.09193.68374.30913.77281.76493.93622.48585.07432.48984.25434.5521

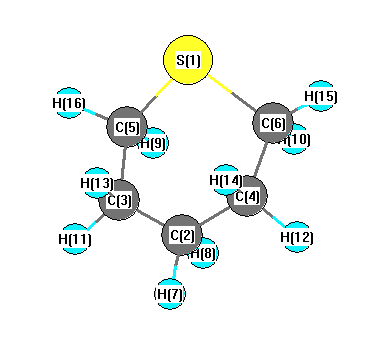
picture of 2H-Thiopyran, tetrahydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C5 C3 112.418 S1 C5 H9 108.994
S1 C5 H16 105.758 S1 C6 C4 112.418
S1 C6 H10 108.994 S1 C6 H15 105.758
C2 C3 C5 112.046 C2 C3 H11 109.549
C2 C3 H13 109.791 C2 C4 C6 112.046
C2 C4 H12 109.549 C2 C4 H14 109.791
C3 C2 C4 112.757 C3 C2 H7 109.587
C3 C2 H8 109.064 C3 C5 H9 110.528
C3 C5 H16 111.266 C4 C2 H7 109.587
C4 C2 H8 109.064 C4 C6 H10 110.528
C4 C6 H15 111.266 C5 S1 C6 97.230
C5 C3 H11 108.884 C5 C3 H13 109.031
C6 C4 H12 108.884 C6 C4 H14 109.031
H7 C2 H8 106.591 H9 C5 H16 107.655
H10 C6 H15 107.655 H11 C3 H13 107.423
H12 C4 H14 107.423
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability