return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4 ((Z)-Hexa-1,5-diyne-3-ene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-230.286332
Energy at 298.15K-230.287250
HF Energy-229.403223
Nuclear repulsion energy166.452262
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3535 3338 80.46      
2 A1 3239 3059 5.06      
3 A1 2138 2019 8.39      
4 A1 1628 1538 2.22      
5 A1 1239 1171 0.21      
6 A1 906 855 4.68      
7 A1 610 576 20.21      
8 A1 422 399 3.31      
9 A1 98 93 0.08      
10 A2 950 898 0.00      
11 A2 594 561 0.00      
12 A2 497 470 0.00      
13 A2 211 199 0.00      
14 B1 778 735 14.67      
15 B1 558 527 90.47      
16 B1 330 312 5.93      
17 B2 3534 3338 62.83      
18 B2 3222 3043 4.09      
19 B2 2153 2034 3.82      
20 B2 1418 1339 1.85      
21 B2 1040 982 6.68      
22 B2 725 685 8.31      
23 B2 601 567 48.80      
24 B2 230 218 2.92      

Unscaled Zero Point Vibrational Energy (zpe) 15327.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 14477.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.22938 0.08815 0.06368

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.676 1.114
C2 0.000 -0.676 1.114
C3 0.000 1.459 -0.066
C4 0.000 -1.459 -0.066
H5 0.000 1.198 2.065
H6 0.000 -1.198 2.065
C7 0.000 2.152 -1.067
C8 0.000 -2.152 -1.067
H9 0.000 2.742 -1.950
H10 0.000 -2.742 -1.950

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 C7 C8 H9 H10
C11.35261.41562.43921.08492.10202.63373.57183.69534.5906
C21.35262.43921.41562.10201.08493.57182.63374.59063.6953
C31.41562.43922.91712.14673.40581.21823.74692.27984.6039
C42.43921.41562.91713.40582.14673.74691.21824.60392.2798
H51.08492.10202.14673.40582.39663.27444.58664.30155.6255
H62.10201.08493.40582.14672.39664.58663.27445.62554.3015
C72.63373.57181.21823.74693.27444.58664.30401.06164.9731
C83.57182.63373.74691.21824.58663.27444.30404.97311.0616
H93.69534.59062.27984.60394.30155.62551.06164.97315.4845
H104.59063.69534.60392.27985.62554.30154.97311.06165.4845

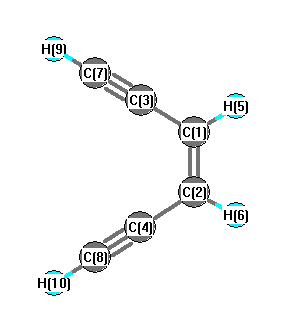
picture of (Z)-Hexa-1,5-diyne-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 123.545 C1 C2 H6 118.762
C1 C3 C7 178.849 C2 C1 C3 123.545
C2 C1 H5 118.762 C2 C4 C8 178.849
C3 C1 H5 117.693 C3 C7 H9 179.082
C4 C2 H6 117.693 C4 C8 H10 179.082
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability