Jump to
S1C2
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -246.716038 |
| Energy at 298.15K | -246.721653 |
| HF Energy | -245.816155 |
| Nuclear repulsion energy | 157.334516 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3192 |
3024 |
14.55 |
|
|
|
| 2 |
A' |
3046 |
2886 |
38.84 |
|
|
|
| 3 |
A' |
3039 |
2879 |
18.31 |
|
|
|
| 4 |
A' |
2365 |
2240 |
0.95 |
|
|
|
| 5 |
A' |
1550 |
1469 |
3.57 |
|
|
|
| 6 |
A' |
1531 |
1451 |
6.73 |
|
|
|
| 7 |
A' |
1511 |
1431 |
0.21 |
|
|
|
| 8 |
A' |
1444 |
1368 |
43.50 |
|
|
|
| 9 |
A' |
1261 |
1195 |
64.28 |
|
|
|
| 10 |
A' |
1197 |
1134 |
104.55 |
|
|
|
| 11 |
A' |
1029 |
975 |
28.76 |
|
|
|
| 12 |
A' |
962 |
911 |
7.66 |
|
|
|
| 13 |
A' |
549 |
520 |
1.75 |
|
|
|
| 14 |
A' |
388 |
368 |
2.46 |
|
|
|
| 15 |
A' |
183 |
174 |
2.74 |
|
|
|
| 16 |
A" |
3104 |
2940 |
37.40 |
|
|
|
| 17 |
A" |
3076 |
2914 |
23.97 |
|
|
|
| 18 |
A" |
1522 |
1442 |
7.18 |
|
|
|
| 19 |
A" |
1288 |
1221 |
3.48 |
|
|
|
| 20 |
A" |
1207 |
1143 |
3.18 |
|
|
|
| 21 |
A" |
1056 |
1001 |
1.70 |
|
|
|
| 22 |
A" |
377 |
357 |
2.71 |
|
|
|
| 23 |
A" |
234 |
222 |
3.89 |
|
|
|
| 24 |
A" |
88 |
84 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17599.0 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 16673.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.126 |
-0.330 |
0.000 |
| O2 |
-0.739 |
-0.599 |
0.000 |
| C3 |
0.000 |
0.589 |
0.000 |
| C4 |
1.433 |
0.250 |
0.000 |
| N5 |
2.569 |
0.021 |
0.000 |
| H6 |
-2.635 |
-1.292 |
0.000 |
| H7 |
-2.424 |
0.236 |
0.892 |
| H8 |
-2.424 |
0.236 |
-0.892 |
| H9 |
-0.213 |
1.202 |
-0.887 |
| H10 |
-0.213 |
1.202 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4125 | 2.3161 | 3.6058 | 4.7078 | 1.0891 | 1.0975 | 1.0975 | 2.6064 | 2.6064 |
O2 | 1.4125 | | 1.3996 | 2.3324 | 3.3657 | 2.0186 | 2.0811 | 2.0811 | 2.0757 | 2.0757 | C3 | 2.3161 | 1.3996 | | 1.4726 | 2.6310 | 3.2381 | 2.6069 | 2.6069 | 1.0990 | 1.0990 | C4 | 3.6058 | 2.3324 | 1.4726 | | 1.1587 | 4.3507 | 3.9588 | 3.9588 | 2.0982 | 2.0982 | N5 | 4.7078 | 3.3657 | 2.6310 | 1.1587 | | 5.3670 | 5.0764 | 5.0764 | 3.1498 | 3.1498 | H6 | 1.0891 | 2.0186 | 3.2381 | 4.3507 | 5.3670 | | 1.7824 | 1.7824 | 3.5885 | 3.5885 | H7 | 1.0975 | 2.0811 | 2.6069 | 3.9588 | 5.0764 | 1.7824 | | 1.7836 | 2.9976 | 2.4128 | H8 | 1.0975 | 2.0811 | 2.6069 | 3.9588 | 5.0764 | 1.7824 | 1.7836 | | 2.4128 | 2.9976 | H9 | 2.6064 | 2.0757 | 1.0990 | 2.0982 | 3.1498 | 3.5885 | 2.9976 | 2.4128 | | 1.7739 | H10 | 2.6064 | 2.0757 | 1.0990 | 2.0982 | 3.1498 | 3.5885 | 2.4128 | 2.9976 | 1.7739 | |
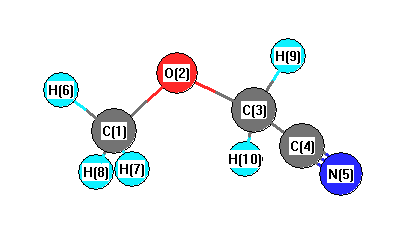 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
110.899 |
|
O2 |
C1 |
H6 |
106.883 |
| O2 |
C1 |
H7 |
111.403 |
|
O2 |
C1 |
H8 |
111.403 |
| O2 |
C3 |
C4 |
108.568 |
|
O2 |
C3 |
H9 |
111.787 |
| O2 |
C3 |
H10 |
111.787 |
|
C3 |
C4 |
N5 |
178.093 |
| C4 |
C3 |
H9 |
108.488 |
|
C4 |
C3 |
H10 |
108.488 |
| H6 |
C1 |
H7 |
109.207 |
|
H6 |
C1 |
H8 |
109.207 |
| H7 |
C1 |
H8 |
108.688 |
|
H9 |
C3 |
H10 |
107.624 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -246.718667 |
| Energy at 298.15K | -246.724404 |
| HF Energy | -245.817817 |
| Nuclear repulsion energy | 160.208684 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3194 |
3026 |
14.94 |
|
|
|
| 2 |
A |
3160 |
2994 |
4.53 |
|
|
|
| 3 |
A |
3120 |
2956 |
28.86 |
|
|
|
| 4 |
A |
3062 |
2901 |
23.06 |
|
|
|
| 5 |
A |
3052 |
2892 |
28.17 |
|
|
|
| 6 |
A |
2349 |
2225 |
1.48 |
|
|
|
| 7 |
A |
1541 |
1460 |
8.47 |
|
|
|
| 8 |
A |
1525 |
1444 |
5.73 |
|
|
|
| 9 |
A |
1519 |
1439 |
3.22 |
|
|
|
| 10 |
A |
1505 |
1426 |
0.84 |
|
|
|
| 11 |
A |
1421 |
1346 |
18.93 |
|
|
|
| 12 |
A |
1342 |
1271 |
7.68 |
|
|
|
| 13 |
A |
1254 |
1188 |
62.12 |
|
|
|
| 14 |
A |
1210 |
1146 |
13.10 |
|
|
|
| 15 |
A |
1193 |
1130 |
73.52 |
|
|
|
| 16 |
A |
1055 |
999 |
6.56 |
|
|
|
| 17 |
A |
978 |
926 |
27.14 |
|
|
|
| 18 |
A |
910 |
862 |
19.56 |
|
|
|
| 19 |
A |
602 |
570 |
1.52 |
|
|
|
| 20 |
A |
405 |
383 |
2.45 |
|
|
|
| 21 |
A |
371 |
352 |
1.74 |
|
|
|
| 22 |
A |
267 |
253 |
10.69 |
|
|
|
| 23 |
A |
183 |
173 |
4.19 |
|
|
|
| 24 |
A |
121 |
114 |
8.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17667.9 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 16738.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.551 |
-0.774 |
0.137 |
| O2 |
1.081 |
0.431 |
-0.441 |
| C3 |
-0.049 |
0.928 |
0.212 |
| C4 |
-1.238 |
0.057 |
0.034 |
| N5 |
-2.163 |
-0.631 |
-0.096 |
| H6 |
2.446 |
-1.056 |
-0.416 |
| H7 |
0.808 |
-1.575 |
0.054 |
| H8 |
1.807 |
-0.632 |
1.194 |
| H9 |
-0.269 |
1.906 |
-0.220 |
| H10 |
0.119 |
1.054 |
1.289 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4166 | 2.3372 | 2.9122 | 3.7238 | 1.0890 | 1.0958 | 1.0972 | 3.2590 | 2.5923 |
O2 | 1.4166 | | 1.3961 | 2.3969 | 3.4306 | 2.0187 | 2.0844 | 2.0807 | 2.0114 | 2.0749 | C3 | 2.3372 | 1.3961 | | 1.4849 | 2.6444 | 3.2488 | 2.6503 | 2.6158 | 1.0910 | 1.0982 | C4 | 2.9122 | 2.3969 | 1.4849 | | 1.1596 | 3.8749 | 2.6172 | 3.3303 | 2.1030 | 2.1009 | N5 | 3.7238 | 3.4306 | 2.6444 | 1.1596 | | 4.6394 | 3.1206 | 4.1738 | 3.1679 | 3.1573 | H6 | 1.0890 | 2.0187 | 3.2488 | 3.8749 | 4.6394 | | 1.7818 | 1.7835 | 4.0226 | 3.5739 | H7 | 1.0958 | 2.0844 | 2.6503 | 2.6172 | 3.1206 | 1.7818 | | 1.7851 | 3.6538 | 2.9853 | H8 | 1.0972 | 2.0807 | 2.6158 | 3.3303 | 4.1738 | 1.7835 | 1.7851 | | 3.5703 | 2.3871 | H9 | 3.2590 | 2.0114 | 1.0910 | 2.1030 | 3.1679 | 4.0226 | 3.6538 | 3.5703 | | 1.7757 | H10 | 2.5923 | 2.0749 | 1.0982 | 2.1009 | 3.1573 | 3.5739 | 2.9853 | 2.3871 | 1.7757 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.390 |
|
O2 |
C1 |
H6 |
106.619 |
| O2 |
C1 |
H7 |
111.485 |
|
O2 |
C1 |
H8 |
111.093 |
| O2 |
C3 |
C4 |
112.568 |
|
O2 |
C3 |
H9 |
107.309 |
| O2 |
C3 |
H10 |
112.031 |
|
C3 |
C4 |
N5 |
179.428 |
| C4 |
C3 |
H9 |
108.488 |
|
C4 |
C3 |
H10 |
107.909 |
| H6 |
C1 |
H7 |
109.283 |
|
H6 |
C1 |
H8 |
109.332 |
| H7 |
C1 |
H8 |
108.976 |
|
H9 |
C3 |
H10 |
108.411 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
