Jump to
S1C2
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -246.707505 |
| Energy at 298.15K | -246.713135 |
| HF Energy | -245.816805 |
| Nuclear repulsion energy | 157.535292 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3206 |
3036 |
13.34 |
|
|
|
| 2 |
A' |
3059 |
2897 |
36.95 |
|
|
|
| 3 |
A' |
3052 |
2891 |
15.55 |
|
|
|
| 4 |
A' |
2399 |
2272 |
1.13 |
|
|
|
| 5 |
A' |
1554 |
1472 |
4.04 |
|
|
|
| 6 |
A' |
1535 |
1454 |
6.58 |
|
|
|
| 7 |
A' |
1517 |
1437 |
0.10 |
|
|
|
| 8 |
A' |
1451 |
1374 |
43.33 |
|
|
|
| 9 |
A' |
1270 |
1203 |
71.95 |
|
|
|
| 10 |
A' |
1207 |
1143 |
95.35 |
|
|
|
| 11 |
A' |
1039 |
984 |
28.73 |
|
|
|
| 12 |
A' |
969 |
918 |
7.97 |
|
|
|
| 13 |
A' |
553 |
524 |
1.85 |
|
|
|
| 14 |
A' |
390 |
370 |
2.55 |
|
|
|
| 15 |
A' |
184 |
175 |
2.75 |
|
|
|
| 16 |
A" |
3120 |
2956 |
35.16 |
|
|
|
| 17 |
A" |
3091 |
2928 |
20.90 |
|
|
|
| 18 |
A" |
1527 |
1446 |
7.48 |
|
|
|
| 19 |
A" |
1295 |
1227 |
3.49 |
|
|
|
| 20 |
A" |
1211 |
1147 |
3.20 |
|
|
|
| 21 |
A" |
1060 |
1004 |
1.71 |
|
|
|
| 22 |
A" |
379 |
359 |
2.74 |
|
|
|
| 23 |
A" |
233 |
221 |
3.99 |
|
|
|
| 24 |
A" |
89 |
84 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17694.3 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 16760.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.123 |
-0.333 |
0.000 |
| O2 |
-0.738 |
-0.598 |
0.000 |
| C3 |
0.000 |
0.589 |
0.000 |
| C4 |
1.432 |
0.252 |
0.000 |
| N5 |
2.565 |
0.025 |
0.000 |
| H6 |
-2.631 |
-1.296 |
0.000 |
| H7 |
-2.423 |
0.231 |
0.891 |
| H8 |
-2.423 |
0.231 |
-0.891 |
| H9 |
-0.214 |
1.201 |
-0.886 |
| H10 |
-0.214 |
1.201 |
0.886 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4104 | 2.3148 | 3.6035 | 4.7021 | 1.0884 | 1.0967 | 1.0967 | 2.6049 | 2.6049 |
O2 | 1.4104 | | 1.3977 | 2.3311 | 3.3615 | 2.0170 | 2.0788 | 2.0788 | 2.0731 | 2.0731 | C3 | 2.3148 | 1.3977 | | 1.4713 | 2.6262 | 3.2361 | 2.6062 | 2.6062 | 1.0983 | 1.0983 | C4 | 3.6035 | 2.3311 | 1.4713 | | 1.1553 | 4.3479 | 3.9569 | 3.9569 | 2.0965 | 2.0965 | N5 | 4.7021 | 3.3615 | 2.6262 | 1.1553 | | 5.3610 | 5.0711 | 5.0711 | 3.1449 | 3.1449 | H6 | 1.0884 | 2.0170 | 3.2361 | 4.3479 | 5.3610 | | 1.7807 | 1.7807 | 3.5864 | 3.5864 | H7 | 1.0967 | 2.0788 | 2.6062 | 3.9569 | 5.0711 | 1.7807 | | 1.7825 | 2.9966 | 2.4125 | H8 | 1.0967 | 2.0788 | 2.6062 | 3.9569 | 5.0711 | 1.7807 | 1.7825 | | 2.4125 | 2.9966 | H9 | 2.6049 | 2.0731 | 1.0983 | 2.0965 | 3.1449 | 3.5864 | 2.9966 | 2.4125 | | 1.7727 | H10 | 2.6049 | 2.0731 | 1.0983 | 2.0965 | 3.1449 | 3.5864 | 2.4125 | 2.9966 | 1.7727 | |
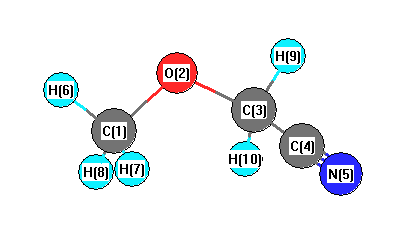 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.039 |
|
O2 |
C1 |
H6 |
106.939 |
| O2 |
C1 |
H7 |
111.411 |
|
O2 |
C1 |
H8 |
111.411 |
| O2 |
C3 |
C4 |
108.654 |
|
O2 |
C3 |
H9 |
111.753 |
| O2 |
C3 |
H10 |
111.753 |
|
C3 |
C4 |
N5 |
178.115 |
| C4 |
C3 |
H9 |
108.481 |
|
C4 |
C3 |
H10 |
108.481 |
| H6 |
C1 |
H7 |
109.159 |
|
H6 |
C1 |
H8 |
109.159 |
| H7 |
C1 |
H8 |
108.710 |
|
H9 |
C3 |
H10 |
107.619 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -246.710013 |
| Energy at 298.15K | -246.715764 |
| HF Energy | -245.818482 |
| Nuclear repulsion energy | 160.393010 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3206 |
3037 |
13.70 |
|
|
|
| 2 |
A |
3170 |
3003 |
3.92 |
|
|
|
| 3 |
A |
3136 |
2970 |
27.11 |
|
|
|
| 4 |
A |
3075 |
2913 |
20.54 |
|
|
|
| 5 |
A |
3065 |
2903 |
26.45 |
|
|
|
| 6 |
A |
2384 |
2258 |
1.70 |
|
|
|
| 7 |
A |
1545 |
1463 |
8.62 |
|
|
|
| 8 |
A |
1530 |
1449 |
5.73 |
|
|
|
| 9 |
A |
1525 |
1444 |
3.47 |
|
|
|
| 10 |
A |
1510 |
1430 |
1.04 |
|
|
|
| 11 |
A |
1427 |
1352 |
18.84 |
|
|
|
| 12 |
A |
1348 |
1276 |
8.35 |
|
|
|
| 13 |
A |
1261 |
1195 |
70.86 |
|
|
|
| 14 |
A |
1215 |
1151 |
17.37 |
|
|
|
| 15 |
A |
1202 |
1138 |
59.43 |
|
|
|
| 16 |
A |
1059 |
1003 |
6.98 |
|
|
|
| 17 |
A |
986 |
934 |
26.99 |
|
|
|
| 18 |
A |
918 |
870 |
19.12 |
|
|
|
| 19 |
A |
607 |
575 |
1.66 |
|
|
|
| 20 |
A |
406 |
384 |
2.57 |
|
|
|
| 21 |
A |
373 |
354 |
1.74 |
|
|
|
| 22 |
A |
268 |
253 |
11.04 |
|
|
|
| 23 |
A |
183 |
174 |
4.20 |
|
|
|
| 24 |
A |
120 |
113 |
8.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17758.7 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 16821.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.553 |
-0.771 |
0.136 |
| O2 |
1.079 |
0.431 |
-0.439 |
| C3 |
-0.052 |
0.927 |
0.210 |
| C4 |
-1.239 |
0.055 |
0.034 |
| N5 |
-2.159 |
-0.632 |
-0.096 |
| H6 |
2.448 |
-1.051 |
-0.416 |
| H7 |
0.812 |
-1.574 |
0.053 |
| H8 |
1.808 |
-0.629 |
1.193 |
| H9 |
-0.273 |
1.904 |
-0.221 |
| H10 |
0.116 |
1.056 |
1.287 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4145 | 2.3373 | 2.9126 | 3.7218 | 1.0885 | 1.0951 | 1.0964 | 3.2582 | 2.5934 |
O2 | 1.4145 | | 1.3947 | 2.3949 | 3.4255 | 2.0182 | 2.0819 | 2.0781 | 2.0108 | 2.0725 | C3 | 2.3373 | 1.3947 | | 1.4829 | 2.6389 | 3.2487 | 2.6505 | 2.6166 | 1.0906 | 1.0975 | C4 | 2.9126 | 2.3949 | 1.4829 | | 1.1560 | 3.8753 | 2.6193 | 3.3307 | 2.1009 | 2.0991 | N5 | 3.7218 | 3.4255 | 2.6389 | 1.1560 | | 4.6377 | 3.1211 | 4.1714 | 3.1624 | 3.1522 | H6 | 1.0885 | 2.0182 | 3.2487 | 3.8753 | 4.6377 | | 1.7802 | 1.7817 | 4.0218 | 3.5744 | H7 | 1.0951 | 2.0819 | 2.6505 | 2.6193 | 3.1211 | 1.7802 | | 1.7839 | 3.6535 | 2.9869 | H8 | 1.0964 | 2.0781 | 2.6166 | 3.3307 | 4.1714 | 1.7817 | 1.7839 | | 3.5704 | 2.3898 | H9 | 3.2582 | 2.0108 | 1.0906 | 2.1009 | 3.1624 | 4.0218 | 3.6535 | 3.5704 | | 1.7737 | H10 | 2.5934 | 2.0725 | 1.0975 | 2.0991 | 3.1522 | 3.5744 | 2.9869 | 2.3898 | 1.7737 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
112.608 |
|
O2 |
C1 |
H6 |
106.752 |
| O2 |
C1 |
H7 |
111.482 |
|
O2 |
C1 |
H8 |
111.082 |
| O2 |
C3 |
C4 |
112.622 |
|
O2 |
C3 |
H9 |
107.377 |
| O2 |
C3 |
H10 |
111.975 |
|
C3 |
C4 |
N5 |
179.424 |
| C4 |
C3 |
H9 |
108.477 |
|
C4 |
C3 |
H10 |
107.943 |
| H6 |
C1 |
H7 |
109.227 |
|
H6 |
C1 |
H8 |
109.266 |
| H7 |
C1 |
H8 |
108.978 |
|
H9 |
C3 |
H10 |
108.315 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
