Jump to
S1C2
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -247.191063 |
| Energy at 298.15K | -247.196662 |
| HF Energy | -247.191063 |
| Nuclear repulsion energy | 157.681661 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3166 |
3166 |
16.80 |
|
|
|
| 2 |
A' |
2994 |
2994 |
49.92 |
|
|
|
| 3 |
A' |
2985 |
2985 |
21.90 |
|
|
|
| 4 |
A' |
2410 |
2410 |
0.54 |
|
|
|
| 5 |
A' |
1520 |
1520 |
5.96 |
|
|
|
| 6 |
A' |
1501 |
1501 |
4.69 |
|
|
|
| 7 |
A' |
1485 |
1485 |
3.66 |
|
|
|
| 8 |
A' |
1425 |
1425 |
41.72 |
|
|
|
| 9 |
A' |
1252 |
1252 |
82.68 |
|
|
|
| 10 |
A' |
1200 |
1200 |
102.22 |
|
|
|
| 11 |
A' |
1030 |
1030 |
31.52 |
|
|
|
| 12 |
A' |
960 |
960 |
6.70 |
|
|
|
| 13 |
A' |
558 |
558 |
2.03 |
|
|
|
| 14 |
A' |
393 |
393 |
2.23 |
|
|
|
| 15 |
A' |
187 |
187 |
2.87 |
|
|
|
| 16 |
A" |
3052 |
3052 |
43.42 |
|
|
|
| 17 |
A" |
3020 |
3020 |
32.33 |
|
|
|
| 18 |
A" |
1492 |
1492 |
8.90 |
|
|
|
| 19 |
A" |
1274 |
1274 |
4.44 |
|
|
|
| 20 |
A" |
1192 |
1192 |
3.00 |
|
|
|
| 21 |
A" |
1033 |
1033 |
1.23 |
|
|
|
| 22 |
A" |
385 |
385 |
2.45 |
|
|
|
| 23 |
A" |
218 |
218 |
3.66 |
|
|
|
| 24 |
A" |
86 |
86 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17408.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17408.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.121 |
-0.353 |
0.000 |
| O2 |
-0.737 |
-0.600 |
0.000 |
| C3 |
0.000 |
0.583 |
0.000 |
| C4 |
1.432 |
0.263 |
0.000 |
| N5 |
2.563 |
0.050 |
0.000 |
| H6 |
-2.618 |
-1.323 |
0.000 |
| H7 |
-2.431 |
0.210 |
0.893 |
| H8 |
-2.431 |
0.210 |
-0.893 |
| H9 |
-0.216 |
1.200 |
-0.887 |
| H10 |
-0.216 |
1.200 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4053 | 2.3179 | 3.6056 | 4.7012 | 1.0904 | 1.1003 | 1.1003 | 2.6125 | 2.6125 |
O2 | 1.4053 | | 1.3939 | 2.3348 | 3.3640 | 2.0150 | 2.0789 | 2.0789 | 2.0734 | 2.0734 | C3 | 2.3179 | 1.3939 | | 1.4671 | 2.6180 | 3.2383 | 2.6162 | 2.6162 | 1.1022 | 1.1022 | C4 | 3.6056 | 2.3348 | 1.4671 | | 1.1513 | 4.3497 | 3.9648 | 3.9648 | 2.0930 | 2.0930 | N5 | 4.7012 | 3.3640 | 2.6180 | 1.1513 | | 5.3603 | 5.0757 | 5.0757 | 3.1360 | 3.1360 | H6 | 1.0904 | 2.0150 | 3.2383 | 4.3497 | 5.3603 | | 1.7843 | 1.7843 | 3.5947 | 3.5947 | H7 | 1.1003 | 2.0789 | 2.6162 | 3.9648 | 5.0757 | 1.7843 | | 1.7862 | 3.0088 | 2.4255 | H8 | 1.1003 | 2.0789 | 2.6162 | 3.9648 | 5.0757 | 1.7843 | 1.7862 | | 2.4255 | 3.0088 | H9 | 2.6125 | 2.0734 | 1.1022 | 2.0930 | 3.1360 | 3.5947 | 3.0088 | 2.4255 | | 1.7745 | H10 | 2.6125 | 2.0734 | 1.1022 | 2.0930 | 3.1360 | 3.5947 | 2.4255 | 3.0088 | 1.7745 | |
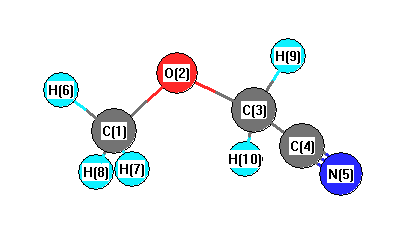 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
111.798 |
|
O2 |
C1 |
H6 |
107.009 |
| O2 |
C1 |
H7 |
111.556 |
|
O2 |
C1 |
H8 |
111.556 |
| O2 |
C3 |
C4 |
109.363 |
|
O2 |
C3 |
H9 |
111.802 |
| O2 |
C3 |
H10 |
111.802 |
|
C3 |
C4 |
N5 |
178.104 |
| C4 |
C3 |
H9 |
108.261 |
|
C4 |
C3 |
H10 |
108.261 |
| H6 |
C1 |
H7 |
109.074 |
|
H6 |
C1 |
H8 |
109.074 |
| H7 |
C1 |
H8 |
108.516 |
|
H9 |
C3 |
H10 |
107.219 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.268 |
|
|
|
| 2 |
O |
-0.251 |
|
|
|
| 3 |
C |
-0.191 |
|
|
|
| 4 |
C |
0.262 |
|
|
|
| 5 |
N |
-0.300 |
|
|
|
| 6 |
H |
0.168 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
| 9 |
H |
0.160 |
|
|
|
| 10 |
H |
0.160 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.335 |
1.670 |
0.000 |
4.646 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.817 |
0.683 |
0.000 |
| y |
0.683 |
-28.643 |
0.000 |
| z |
0.000 |
0.000 |
-28.679 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.156 |
0.683 |
0.000 |
| y |
0.683 |
3.605 |
0.000 |
| z |
0.000 |
0.000 |
3.550 |
|
| Polar |
|---|
| 3z2-r2 | 7.101 |
|---|
| x2-y2 | -7.174 |
|---|
| xy | 0.683 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.117 |
0.027 |
0.000 |
| y |
0.027 |
4.912 |
0.000 |
| z |
0.000 |
0.000 |
4.688 |
<r2> (average value of r
2) Å
2
| <r2> |
141.770 |
| (<r2>)1/2 |
11.907 |
Jump to
S1C1
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -247.193834 |
| Energy at 298.15K | -247.199581 |
| HF Energy | -247.193834 |
| Nuclear repulsion energy | 160.533467 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3167 |
3167 |
17.02 |
|
|
|
| 2 |
A |
3126 |
3126 |
5.05 |
|
|
|
| 3 |
A |
3075 |
3075 |
33.51 |
|
|
|
| 4 |
A |
3009 |
3009 |
34.47 |
|
|
|
| 5 |
A |
3006 |
3006 |
31.38 |
|
|
|
| 6 |
A |
2392 |
2392 |
0.20 |
|
|
|
| 7 |
A |
1517 |
1517 |
9.66 |
|
|
|
| 8 |
A |
1499 |
1499 |
5.80 |
|
|
|
| 9 |
A |
1496 |
1496 |
5.92 |
|
|
|
| 10 |
A |
1482 |
1482 |
2.29 |
|
|
|
| 11 |
A |
1407 |
1407 |
17.71 |
|
|
|
| 12 |
A |
1330 |
1330 |
9.25 |
|
|
|
| 13 |
A |
1246 |
1246 |
81.23 |
|
|
|
| 14 |
A |
1200 |
1200 |
26.64 |
|
|
|
| 15 |
A |
1191 |
1191 |
49.58 |
|
|
|
| 16 |
A |
1045 |
1045 |
6.80 |
|
|
|
| 17 |
A |
978 |
978 |
28.13 |
|
|
|
| 18 |
A |
906 |
906 |
19.27 |
|
|
|
| 19 |
A |
609 |
609 |
1.41 |
|
|
|
| 20 |
A |
406 |
406 |
1.99 |
|
|
|
| 21 |
A |
378 |
378 |
2.14 |
|
|
|
| 22 |
A |
264 |
264 |
10.08 |
|
|
|
| 23 |
A |
185 |
185 |
3.92 |
|
|
|
| 24 |
A |
124 |
124 |
8.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17518.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17518.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.556 |
-0.770 |
0.132 |
| O2 |
1.079 |
0.434 |
-0.427 |
| C3 |
-0.058 |
0.927 |
0.204 |
| C4 |
-1.240 |
0.054 |
0.033 |
| N5 |
-2.155 |
-0.636 |
-0.094 |
| H6 |
2.465 |
-1.030 |
-0.411 |
| H7 |
0.825 |
-1.582 |
0.026 |
| H8 |
1.794 |
-0.645 |
1.199 |
| H9 |
-0.279 |
1.901 |
-0.239 |
| H10 |
0.097 |
1.070 |
1.285 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
N5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4100 | 2.3424 | 2.9171 | 3.7199 | 1.0903 | 1.0983 | 1.0998 | 3.2613 | 2.6158 |
O2 | 1.4100 | | 1.3904 | 2.3949 | 3.4223 | 2.0157 | 2.0815 | 2.0780 | 2.0079 | 2.0736 | C3 | 2.3424 | 1.3904 | | 1.4799 | 2.6321 | 3.2512 | 2.6654 | 2.6250 | 1.0927 | 1.1013 | C4 | 2.9171 | 2.3949 | 1.4799 | | 1.1523 | 3.8861 | 2.6348 | 3.3255 | 2.0995 | 2.0952 | N5 | 3.7199 | 3.4223 | 2.6321 | 1.1523 | | 4.6471 | 3.1282 | 4.1550 | 3.1580 | 3.1435 | H6 | 1.0903 | 2.0157 | 3.2512 | 3.8861 | 4.6471 | | 1.7848 | 1.7857 | 4.0184 | 3.5907 | H7 | 1.0983 | 2.0815 | 2.6654 | 2.6348 | 3.1282 | 1.7848 | | 1.7871 | 3.6630 | 3.0249 | H8 | 1.0998 | 2.0780 | 2.6250 | 3.3255 | 4.1550 | 1.7857 | 1.7871 | | 3.5845 | 2.4147 | H9 | 3.2613 | 2.0079 | 1.0927 | 2.0995 | 3.1580 | 4.0184 | 3.6630 | 3.5845 | | 1.7761 | H10 | 2.6158 | 2.0736 | 1.1013 | 2.0952 | 3.1435 | 3.5907 | 3.0249 | 2.4147 | 1.7761 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
C3 |
113.535 |
|
O2 |
C1 |
H6 |
106.750 |
| O2 |
C1 |
H7 |
111.563 |
|
O2 |
C1 |
H8 |
111.177 |
| O2 |
C3 |
C4 |
113.066 |
|
O2 |
C3 |
H9 |
107.317 |
| O2 |
C3 |
H10 |
112.125 |
|
C3 |
C4 |
N5 |
179.274 |
| C4 |
C3 |
H9 |
108.456 |
|
C4 |
C3 |
H10 |
107.626 |
| H6 |
C1 |
H7 |
109.279 |
|
H6 |
C1 |
H8 |
109.240 |
| H7 |
C1 |
H8 |
108.782 |
|
H9 |
C3 |
H10 |
108.092 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.282 |
|
|
|
| 2 |
O |
-0.244 |
|
|
|
| 3 |
C |
-0.195 |
|
|
|
| 4 |
C |
0.233 |
|
|
|
| 5 |
N |
-0.305 |
|
|
|
| 6 |
H |
0.165 |
|
|
|
| 7 |
H |
0.152 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
H |
0.190 |
|
|
|
| 10 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.578 |
1.285 |
1.305 |
3.163 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.746 |
-5.379 |
0.510 |
| y |
-5.379 |
-27.552 |
0.533 |
| z |
0.510 |
0.533 |
-29.132 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.404 |
-5.379 |
0.510 |
| y |
-5.379 |
4.887 |
0.533 |
| z |
0.510 |
0.533 |
2.517 |
|
| Polar |
|---|
| 3z2-r2 | 5.035 |
|---|
| x2-y2 | -8.194 |
|---|
| xy | -5.379 |
|---|
| xz | 0.510 |
|---|
| yz | 0.533 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.968 |
0.389 |
0.174 |
| y |
0.389 |
5.695 |
0.192 |
| z |
0.174 |
0.192 |
4.662 |
<r2> (average value of r
2) Å
2
| <r2> |
121.892 |
| (<r2>)1/2 |
11.040 |
