return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (Thiophene, tetrahydro-2-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-593.690834
Energy at 298.15K-593.702731
HF Energy-592.710635
Nuclear repulsion energy312.186139
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3205 3027 12.04      
2 A 3196 3019 8.07      
3 A 3189 3012 12.01      
4 A 3174 2998 15.29      
5 A 3161 2986 14.11      
6 A 3128 2954 18.56      
7 A 3113 2940 8.68      
8 A 3111 2938 16.73      
9 A 3098 2926 26.64      
10 A 3096 2924 5.64      
11 A 1525 1441 6.29      
12 A 1523 1439 0.12      
13 A 1516 1432 4.80      
14 A 1506 1422 4.89      
15 A 1501 1417 2.51      
16 A 1425 1346 3.42      
17 A 1388 1311 1.58      
18 A 1364 1289 0.41      
19 A 1328 1254 0.19      
20 A 1313 1240 4.09      
21 A 1300 1228 21.68      
22 A 1252 1182 4.10      
23 A 1210 1143 6.82      
24 A 1173 1108 0.30      
25 A 1126 1064 0.15      
26 A 1088 1028 1.83      
27 A 1057 998 0.45      
28 A 1054 996 3.59      
29 A 983 928 2.45      
30 A 971 917 1.95      
31 A 924 873 0.83      
32 A 873 824 0.98      
33 A 868 820 2.87      
34 A 719 679 1.54      
35 A 665 628 1.92      
36 A 530 500 0.66      
37 A 461 435 0.06      
38 A 415 392 0.48      
39 A 288 272 0.77      
40 A 266 251 0.18      
41 A 253 239 0.20      
42 A 91 86 1.43      

Unscaled Zero Point Vibrational Energy (zpe) 31711.5 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 29951.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.14160 0.09365 0.06203

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.259 0.133 0.211
H2 -2.189 0.011 1.293
H3 -2.830 1.040 -0.005
H4 -2.813 -0.717 -0.192
S5 0.110 -1.284 -0.072
C6 1.654 -0.375 0.295
H7 2.487 -0.863 -0.210
H8 1.837 -0.397 1.371
C9 -0.870 0.233 -0.402
H10 -0.954 0.343 -1.487
C11 1.421 1.054 -0.193
H12 1.556 1.101 -1.277
H13 2.129 1.748 0.267
C14 -0.027 1.376 0.156
H15 -0.356 2.336 -0.253
H16 -0.147 1.413 1.244

Atom - Atom Distances (Å)
  C1 H2 H3 H4 S5 C6 H7 H8 C9 H10 C11 H12 H13 C14 H15 H16
C11.09111.09321.09112.77523.94714.86844.29011.52162.15113.81534.20814.67632.55582.94802.6775
H21.09111.77611.76722.97063.98904.98914.04712.15853.05904.04074.67084.76592.79823.34012.4777
H31.09321.77611.76633.74854.71205.65195.07362.15702.48984.25584.56795.01722.82822.80472.9835
H41.09111.76721.76632.98004.50635.30234.91572.17252.50034.58934.85475.54133.50183.91883.7022
S52.77522.97063.74852.98001.82852.41802.41851.83542.40442.68303.03763.65802.67353.65363.0121
C63.94713.98904.71204.50631.82851.08991.09122.68793.23951.52822.15822.17582.43193.41872.7099
H74.86844.98915.65195.30232.41801.08991.77203.53683.86402.19402.42072.67843.38724.27973.7733
H84.29014.04715.07364.91572.41851.09121.77203.29604.06232.17313.05432.42962.84493.86142.6886
C91.52162.15852.15702.17251.83542.68793.53683.29601.09382.44282.72143.42591.52642.17002.1506
H102.15113.05902.48982.50032.40443.23953.86404.06231.09382.79692.63103.81522.15092.41833.0420
C113.81534.04074.25584.58932.68301.52822.19402.17312.44282.79691.09341.09261.52412.19132.1569
H124.20814.67084.56794.85473.03762.15822.42073.05432.72142.63101.09341.76932.15342.49563.0584
H134.67634.76595.01725.54133.65802.17582.67842.42963.42593.81521.09261.76932.19062.60562.4993
C142.55582.79822.82823.50182.67352.43193.38722.84491.52642.15091.52412.15342.19061.09341.0951
H152.94803.34012.80473.91883.65363.41874.27973.86142.17002.41832.19132.49562.60561.09341.7712
H162.67752.47772.98353.70223.01212.70993.77332.68862.15063.04202.15693.05842.49931.09511.7712

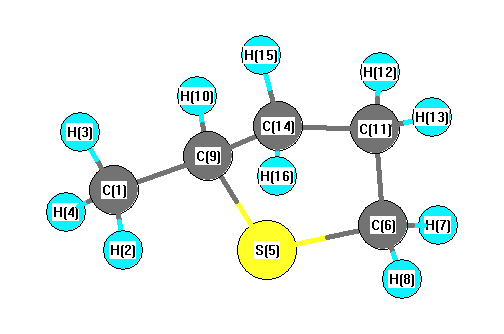
picture of Thiophene, tetrahydro-2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C9 S5 111.182 C1 C9 H10 109.580
C1 C9 C14 113.965 H2 C1 H3 108.806
H2 C1 H4 108.161 H2 C1 C9 110.331
H3 C1 H4 107.932 H3 C1 C9 110.083
H4 C1 C9 111.445 S5 C6 H7 109.275
S5 C6 H8 109.243 S5 C6 C11 105.781
S5 C9 H10 107.630 S5 C9 C14 104.989
C6 S5 C9 94.385 C6 C11 H12 109.704
C6 C11 H13 111.148 C6 C11 C14 105.637
H7 C6 H8 108.662 H7 C6 C11 112.785
H8 C6 C11 111.014 C9 C14 C11 106.410
C9 C14 H15 110.761 C9 C14 H16 109.136
H10 C9 C14 109.235 C11 C14 H15 112.649
C11 C14 H16 109.792 H12 C11 H13 108.070
H12 C11 C14 109.612 H13 C11 C14 112.645
H15 C14 H16 108.054
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability