Jump to
S1C2
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Geometric Data calculated at MP2/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -235.182517 |
| Energy at 298.15K | -235.196062 |
| Nuclear repulsion energy | 255.887558 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3205 |
3028 |
34.56 |
|
|
|
| 2 |
A' |
3184 |
3008 |
26.50 |
|
|
|
| 3 |
A' |
3181 |
3004 |
20.36 |
|
|
|
| 4 |
A' |
3175 |
2999 |
7.86 |
|
|
|
| 5 |
A' |
3138 |
2963 |
23.96 |
|
|
|
| 6 |
A' |
3113 |
2941 |
4.90 |
|
|
|
| 7 |
A' |
3087 |
2915 |
19.03 |
|
|
|
| 8 |
A' |
3082 |
2911 |
18.91 |
|
|
|
| 9 |
A' |
1533 |
1448 |
2.60 |
|
|
|
| 10 |
A' |
1529 |
1445 |
0.82 |
|
|
|
| 11 |
A' |
1518 |
1434 |
5.91 |
|
|
|
| 12 |
A' |
1501 |
1418 |
0.27 |
|
|
|
| 13 |
A' |
1431 |
1352 |
1.99 |
|
|
|
| 14 |
A' |
1417 |
1338 |
10.20 |
|
|
|
| 15 |
A' |
1337 |
1263 |
4.53 |
|
|
|
| 16 |
A' |
1325 |
1252 |
5.11 |
|
|
|
| 17 |
A' |
1251 |
1182 |
0.62 |
|
|
|
| 18 |
A' |
1222 |
1154 |
0.26 |
|
|
|
| 19 |
A' |
1035 |
977 |
0.97 |
|
|
|
| 20 |
A' |
1011 |
955 |
0.52 |
|
|
|
| 21 |
A' |
968 |
914 |
0.36 |
|
|
|
| 22 |
A' |
946 |
893 |
0.40 |
|
|
|
| 23 |
A' |
892 |
843 |
0.43 |
|
|
|
| 24 |
A' |
717 |
677 |
0.96 |
|
|
|
| 25 |
A' |
604 |
571 |
1.67 |
|
|
|
| 26 |
A' |
391 |
370 |
0.32 |
|
|
|
| 27 |
A' |
349 |
330 |
0.03 |
|
|
|
| 28 |
A' |
187 |
176 |
0.00 |
|
|
|
| 29 |
A" |
3180 |
3003 |
17.73 |
|
|
|
| 30 |
A" |
3177 |
3000 |
17.93 |
|
|
|
| 31 |
A" |
3172 |
2996 |
6.52 |
|
|
|
| 32 |
A" |
3110 |
2938 |
47.16 |
|
|
|
| 33 |
A" |
1534 |
1448 |
6.83 |
|
|
|
| 34 |
A" |
1510 |
1426 |
0.08 |
|
|
|
| 35 |
A" |
1494 |
1411 |
2.60 |
|
|
|
| 36 |
A" |
1288 |
1217 |
0.11 |
|
|
|
| 37 |
A" |
1270 |
1199 |
0.29 |
|
|
|
| 38 |
A" |
1237 |
1168 |
0.12 |
|
|
|
| 39 |
A" |
1197 |
1131 |
2.86 |
|
|
|
| 40 |
A" |
1098 |
1037 |
0.06 |
|
|
|
| 41 |
A" |
996 |
940 |
0.01 |
|
|
|
| 42 |
A" |
964 |
910 |
1.55 |
|
|
|
| 43 |
A" |
903 |
853 |
0.04 |
|
|
|
| 44 |
A" |
797 |
753 |
0.26 |
|
|
|
| 45 |
A" |
380 |
359 |
0.07 |
|
|
|
| 46 |
A" |
298 |
282 |
0.02 |
|
|
|
| 47 |
A" |
274 |
259 |
0.00 |
|
|
|
| 48 |
A" |
232 |
219 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37220.6 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 35154.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.110 |
0.400 |
0.000 |
| C2 |
0.404 |
-1.706 |
0.000 |
| C3 |
-0.110 |
-0.720 |
1.076 |
| C4 |
-0.110 |
-0.720 |
-1.076 |
| C5 |
1.206 |
1.168 |
0.000 |
| C6 |
-1.291 |
1.352 |
0.000 |
| H7 |
1.492 |
-1.779 |
0.000 |
| H8 |
-0.011 |
-2.713 |
0.000 |
| H9 |
0.500 |
-0.562 |
1.968 |
| H10 |
0.500 |
-0.562 |
-1.968 |
| H11 |
-1.129 |
-0.967 |
1.382 |
| H12 |
-1.129 |
-0.967 |
-1.382 |
| H13 |
2.066 |
0.495 |
0.000 |
| H14 |
-2.236 |
0.803 |
0.000 |
| H15 |
1.280 |
1.807 |
-0.885 |
| H16 |
1.280 |
1.807 |
0.885 |
| H17 |
-1.275 |
1.996 |
0.884 |
| H18 |
-1.275 |
1.996 |
-0.884 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1679 | 1.5534 | 1.5534 | 1.5232 | 1.5168 | 2.7045 | 3.1153 | 2.2736 | 2.2736 | 2.1953 | 2.1953 | 2.1773 | 2.1637 | 2.1659 | 2.1659 | 2.1644 | 2.1644 |
C2 | 2.1679 | | 1.5468 | 1.5468 | 2.9838 | 3.4958 | 1.0901 | 1.0897 | 2.2779 | 2.2779 | 2.1919 | 2.1919 | 2.7572 | 3.6414 | 3.7264 | 3.7264 | 4.1596 | 4.1596 | C3 | 1.5534 | 1.5468 | | 2.1517 | 2.5405 | 2.6164 | 2.2006 | 2.2671 | 1.0918 | 3.1079 | 1.0925 | 2.6721 | 2.7138 | 2.8277 | 3.4870 | 2.8901 | 2.9619 | 3.5461 | C4 | 1.5534 | 1.5468 | 2.1517 | | 2.5405 | 2.6164 | 2.2006 | 2.2671 | 3.1079 | 1.0918 | 2.6721 | 1.0925 | 2.7138 | 2.8277 | 2.8901 | 3.4870 | 3.5461 | 2.9619 | C5 | 1.5232 | 2.9838 | 2.5405 | 2.5405 | | 2.5038 | 2.9612 | 4.0679 | 2.7135 | 2.7135 | 3.4528 | 3.4528 | 1.0923 | 3.4609 | 1.0933 | 1.0933 | 2.7604 | 2.7604 | C6 | 1.5168 | 3.4958 | 2.6164 | 2.6164 | 2.5038 | | 4.1887 | 4.2616 | 3.2774 | 3.2774 | 2.7043 | 2.7043 | 3.4645 | 1.0924 | 2.7570 | 2.7570 | 1.0937 | 1.0937 | H7 | 2.7045 | 1.0901 | 2.2006 | 2.2006 | 2.9612 | 4.1887 | | 1.7695 | 2.5171 | 2.5171 | 3.0717 | 3.0717 | 2.3450 | 4.5340 | 3.6992 | 3.6992 | 4.7630 | 4.7630 | H8 | 3.1153 | 1.0897 | 2.2671 | 2.2671 | 4.0679 | 4.2616 | 1.7695 | | 2.9599 | 2.9599 | 2.4914 | 2.4914 | 3.8214 | 4.1605 | 4.7832 | 4.7832 | 4.9553 | 4.9553 | H9 | 2.2736 | 2.2779 | 1.0918 | 3.1079 | 2.7135 | 3.2774 | 2.5171 | 2.9599 | | 3.9352 | 1.7779 | 3.7466 | 2.7274 | 3.6356 | 3.7885 | 2.7187 | 3.2968 | 4.2217 | H10 | 2.2736 | 2.2779 | 3.1079 | 1.0918 | 2.7135 | 3.2774 | 2.5171 | 2.9599 | 3.9352 | | 3.7466 | 1.7779 | 2.7274 | 3.6356 | 2.7187 | 3.7885 | 4.2217 | 3.2968 | H11 | 2.1953 | 2.1919 | 1.0925 | 2.6721 | 3.4528 | 2.7043 | 3.0717 | 2.4914 | 1.7779 | 3.7466 | | 2.7638 | 3.7751 | 2.5034 | 4.3168 | 3.7075 | 3.0085 | 3.7330 | H12 | 2.1953 | 2.1919 | 2.6721 | 1.0925 | 3.4528 | 2.7043 | 3.0717 | 2.4914 | 3.7466 | 1.7779 | 2.7638 | | 3.7751 | 2.5034 | 3.7075 | 4.3168 | 3.7330 | 3.0085 | H13 | 2.1773 | 2.7572 | 2.7138 | 2.7138 | 1.0923 | 3.4645 | 2.3450 | 3.8214 | 2.7274 | 2.7274 | 3.7751 | 3.7751 | | 4.3123 | 1.7665 | 1.7665 | 3.7675 | 3.7675 | H14 | 2.1637 | 3.6414 | 2.8277 | 2.8277 | 3.4609 | 1.0924 | 4.5340 | 4.1605 | 3.6356 | 3.6356 | 2.5034 | 2.5034 | 4.3123 | | 3.7618 | 3.7618 | 1.7686 | 1.7686 | H15 | 2.1659 | 3.7264 | 3.4870 | 2.8901 | 1.0933 | 2.7570 | 3.6992 | 4.7832 | 3.7885 | 2.7187 | 4.3168 | 3.7075 | 1.7665 | 3.7618 | | 1.7691 | 3.1129 | 2.5621 | H16 | 2.1659 | 3.7264 | 2.8901 | 3.4870 | 1.0933 | 2.7570 | 3.6992 | 4.7832 | 2.7187 | 3.7885 | 3.7075 | 4.3168 | 1.7665 | 3.7618 | 1.7691 | | 2.5621 | 3.1129 | H17 | 2.1644 | 4.1596 | 2.9619 | 3.5461 | 2.7604 | 1.0937 | 4.7630 | 4.9553 | 3.2968 | 4.2217 | 3.0085 | 3.7330 | 3.7675 | 1.7686 | 3.1129 | 2.5621 | | 1.7671 | H18 | 2.1644 | 4.1596 | 3.5461 | 2.9619 | 2.7604 | 1.0937 | 4.7630 | 4.9553 | 4.2217 | 3.2968 | 3.7330 | 3.0085 | 3.7675 | 1.7686 | 2.5621 | 3.1129 | 1.7671 | |
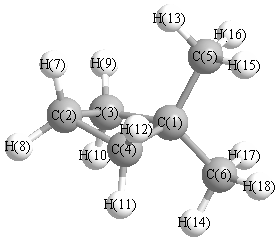 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.736 |
|
C1 |
C3 |
H9 |
117.459 |
| C1 |
C3 |
H11 |
110.931 |
|
C1 |
C4 |
C2 |
88.736 |
| C1 |
C4 |
H10 |
117.459 |
|
C1 |
C4 |
H12 |
110.931 |
| C1 |
C5 |
H13 |
111.646 |
|
C1 |
C5 |
H15 |
110.670 |
| C1 |
C5 |
H16 |
110.670 |
|
C1 |
C6 |
H14 |
110.998 |
| C1 |
C6 |
H17 |
110.971 |
|
C1 |
C6 |
H18 |
110.971 |
| C2 |
C3 |
H9 |
118.357 |
|
C2 |
C3 |
H11 |
111.131 |
| C2 |
C4 |
H10 |
118.357 |
|
C2 |
C4 |
H12 |
111.131 |
| C3 |
C1 |
C4 |
87.665 |
|
C3 |
C1 |
C5 |
111.325 |
| C3 |
C1 |
C6 |
116.894 |
|
C3 |
C2 |
C4 |
88.137 |
| C3 |
C2 |
H7 |
111.975 |
|
C3 |
C2 |
H8 |
117.554 |
| C4 |
C1 |
C5 |
111.325 |
|
C4 |
C1 |
C6 |
116.894 |
| C4 |
C2 |
H7 |
111.975 |
|
C4 |
C2 |
H8 |
117.554 |
| C5 |
C1 |
C6 |
110.894 |
|
H7 |
C2 |
H8 |
108.537 |
| H9 |
C3 |
H11 |
108.963 |
|
H10 |
C4 |
H12 |
108.963 |
| H13 |
C5 |
H15 |
107.855 |
|
H13 |
C5 |
H16 |
107.855 |
| H14 |
C6 |
H17 |
107.998 |
|
H14 |
C6 |
H18 |
107.998 |
| H15 |
C5 |
H16 |
108.004 |
|
H17 |
C6 |
H18 |
107.768 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
