return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Butanenitrile, 2-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-250.041162
Energy at 298.15K-250.050656
HF Energy-249.054865
Nuclear repulsion energy220.347008
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3219 3041 9.15      
2 A 3209 3031 11.05      
3 A 3208 3030 15.19      
4 A 3199 3021 15.28      
5 A 3165 2990 5.26      
6 A 3114 2941 5.96      
7 A 3113 2940 19.24      
8 A 3108 2935 6.95      
9 A 3102 2930 1.14      
10 A 2207 2084 0.01      
11 A 1542 1457 2.50      
12 A 1532 1447 8.29      
13 A 1527 1442 7.78      
14 A 1522 1437 7.30      
15 A 1510 1426 0.18      
16 A 1434 1355 2.80      
17 A 1428 1349 4.92      
18 A 1392 1315 0.33      
19 A 1361 1286 1.14      
20 A 1330 1256 2.29      
21 A 1295 1223 1.16      
22 A 1202 1135 2.00      
23 A 1166 1101 0.16      
24 A 1129 1067 4.98      
25 A 1075 1015 1.22      
26 A 1015 959 1.61      
27 A 983 928 4.35      
28 A 919 868 0.39      
29 A 823 777 0.52      
30 A 779 736 3.29      
31 A 558 527 0.09      
32 A 534 504 0.87      
33 A 401 378 0.26      
34 A 323 305 0.75      
35 A 300 283 0.08      
36 A 222 209 0.06      
37 A 204 192 2.48      
38 A 160 151 5.45      
39 A 93 88 1.52      

Unscaled Zero Point Vibrational Energy (zpe) 29200.2 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 27579.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.21901 0.07384 0.05920

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.516 -0.280 0.121
N2 -2.601 -0.669 -0.102
C3 -0.011 1.632 -0.153
H4 -0.105 1.647 -1.240
H5 -0.781 2.279 0.266
H6 0.962 2.039 0.119
C7 2.299 -0.420 0.063
H8 2.456 -0.341 1.141
H9 2.973 -1.189 -0.316
H10 2.593 0.526 -0.392
C11 0.851 -0.779 -0.254
H12 0.691 -0.798 -1.336
H13 0.631 -1.785 0.113
C14 -0.157 0.199 0.373
H15 -0.010 0.201 1.459

Atom - Atom Distances (Å)
  C1 N2 C3 H4 H5 H6 C7 H8 H9 H10 C11 H12 H13 C14 H15
C11.17432.44902.74962.66703.39493.81854.10134.60154.21912.44802.69492.62231.46332.0712
N21.17433.46543.59083.48484.48194.90985.21825.60315.33823.45733.51813.42672.63753.1482
C32.44903.46541.09141.09021.08953.09693.41324.10942.83862.56232.79213.48661.53312.1550
H42.74963.59081.09141.76791.77193.42764.02234.28623.04202.78792.57303.76192.16843.0634
H52.66703.48481.09021.76791.76614.10074.25575.14433.85883.50563.76874.30552.17452.5172
H63.39494.48191.08951.77191.76612.79982.99053.82842.28232.84573.20043.83882.16902.4740
C73.81854.90983.09693.42764.10072.79981.09231.09061.09011.52582.16502.15602.55182.7691
H84.10135.21823.41324.02234.25572.99051.09231.76331.76652.17133.07562.54352.77632.5448
H94.60155.60314.10944.28625.14433.82841.09061.76331.75872.16272.53052.45453.49293.7395
H104.21915.33822.83863.04203.85882.28231.09011.76651.75872.18182.50313.07352.87303.2104
C112.44803.45732.56232.78793.50562.84571.52582.17132.16272.18181.09381.09281.53822.1534
H122.69493.51812.79212.57303.76873.20042.16503.07562.53052.50311.09381.75422.15273.0498
H132.62233.42673.48663.76194.30553.83882.15602.54352.45453.07351.09281.75422.15072.4835
C141.46332.63751.53312.16842.17452.16902.55182.77633.49292.87301.53822.15272.15071.0957
H152.07123.14822.15503.06342.51722.47402.76912.54483.73953.21042.15343.04982.48351.0957

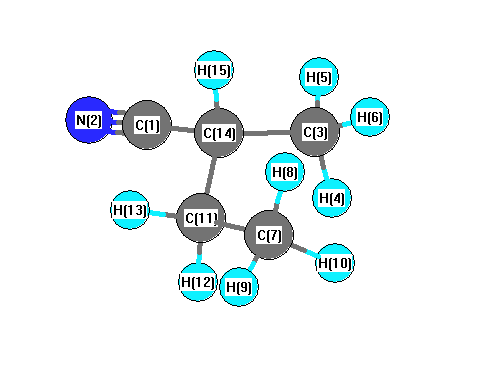
picture of Butanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C14 C3 109.614 C1 C14 C11 109.262
C1 C14 H15 107.202 N2 C1 C14 178.899
C3 C14 C11 113.079 C3 C14 H15 108.989
H4 C3 H5 108.265 H4 C3 H6 108.673
H4 C3 C14 110.288 H5 C3 H6 108.239
H5 C3 C14 110.849 H6 C3 C14 110.452
C7 C11 H12 110.385 C7 C11 H13 109.734
C7 C11 C14 112.784 H8 C7 H9 107.753
H8 C7 H10 108.079 H8 C7 C11 110.981
H9 C7 H10 107.510 H9 C7 C11 110.393
H10 C7 C11 111.958 C11 C14 H15 108.519
H12 C11 H13 106.694 H12 C11 C14 108.568
H13 C11 C14 108.472
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability