Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -1590.757267 |
| Energy at 298.15K | |
| HF Energy | -1590.030171 |
| Nuclear repulsion energy | 353.571709 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
549 |
516 |
0.00 |
33.83 |
0.04 |
0.07 |
| 2 |
A1 |
217 |
204 |
0.00 |
1.79 |
0.75 |
0.86 |
| 3 |
B1 |
492 |
462 |
0.00 |
15.25 |
0.75 |
0.86 |
| 4 |
B2 |
307 |
289 |
0.00 |
8.15 |
0.75 |
0.86 |
| 5 |
E |
462 |
434 |
0.00 |
4.28 |
0.75 |
0.86 |
| 5 |
E |
462 |
434 |
0.00 |
4.28 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1243.5 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 1168.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.415 |
0.322 |
| S2 |
0.000 |
-1.415 |
0.322 |
| S3 |
-1.415 |
0.000 |
-0.322 |
| S4 |
1.415 |
0.000 |
-0.322 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8293 | 2.1019 | 2.1019 |
S2 | 2.8293 | | 2.1019 | 2.1019 | S3 | 2.1019 | 2.1019 | | 2.8293 | S4 | 2.1019 | 2.1019 | 2.8293 | |
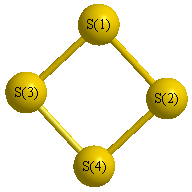 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.698 |
|
S1 |
S3 |
S4 |
47.698 |
| S2 |
S1 |
S3 |
47.698 |
|
S2 |
S4 |
S3 |
47.698 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -1590.823258 |
| Energy at 298.15K | |
| HF Energy | -1589.963982 |
| Nuclear repulsion energy | 330.010389 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
672 |
632 |
0.00 |
0.00 |
0.33 |
0.50 |
| 2 |
A1 |
433 |
407 |
0.11 |
61911.48 |
0.33 |
0.50 |
| 3 |
A1 |
233 |
219 |
0.00 |
0.00 |
0.33 |
0.50 |
| 4 |
A2 |
233 |
219 |
0.00 |
0.58 |
0.75 |
0.86 |
| 5 |
B2 |
5226 |
4912 |
0.00 |
778.56 |
0.75 |
0.86 |
| 6 |
B2 |
317 |
298 |
0.00 |
8179.56 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3557.3 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 3343.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.291 |
0.961 |
| S2 |
0.000 |
-1.291 |
0.961 |
| S3 |
0.000 |
1.291 |
-0.961 |
| S4 |
0.000 |
-1.291 |
-0.961 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5814 | 1.9230 | 3.2191 |
S2 | 2.5814 | | 3.2191 | 1.9230 | S3 | 1.9230 | 3.2191 | | 2.5819 | S4 | 3.2191 | 1.9230 | 2.5819 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.008 |
|
S1 |
S3 |
S4 |
89.992 |
| S2 |
S1 |
S3 |
90.008 |
|
S2 |
S4 |
S3 |
89.992 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -1590.823258 |
| Energy at 298.15K | |
| HF Energy | -1589.964021 |
| Nuclear repulsion energy | 330.009975 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
673 |
633 |
0.00 |
0.00 |
0.33 |
0.50 |
| 2 |
Ag |
233 |
219 |
0.00 |
0.00 |
0.33 |
0.50 |
| 3 |
Au |
233 |
219 |
0.00 |
9573.09 |
0.75 |
0.86 |
| 4 |
B1u |
5481 |
5152 |
0.00 |
0.00 |
0.38 |
0.55 |
| 5 |
B2u |
432 |
406 |
0.10 |
0.02 |
0.71 |
0.83 |
| 6 |
B3g |
317 |
298 |
0.00 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3684.6 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 3463.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.961 |
1.291 |
| S2 |
0.000 |
0.961 |
-1.291 |
| S3 |
0.000 |
-0.961 |
1.291 |
| S4 |
0.000 |
-0.961 |
-1.291 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5822 | 1.9226 | 3.2193 |
S2 | 2.5822 | | 3.2193 | 1.9226 | S3 | 1.9226 | 3.2193 | | 2.5822 | S4 | 3.2193 | 1.9226 | 2.5822 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
