Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -133.602526 |
| Energy at 298.15K | -133.608004 |
| HF Energy | -133.090916 |
| Nuclear repulsion energy | 70.855251 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3512 |
3317 |
0.02 |
|
|
|
| 2 |
A' |
3230 |
3051 |
4.36 |
|
|
|
| 3 |
A' |
3105 |
2933 |
8.00 |
|
|
|
| 4 |
A' |
3092 |
2921 |
44.68 |
|
|
|
| 5 |
A' |
1711 |
1616 |
39.73 |
|
|
|
| 6 |
A' |
1501 |
1418 |
10.00 |
|
|
|
| 7 |
A' |
1435 |
1356 |
17.68 |
|
|
|
| 8 |
A' |
1401 |
1324 |
10.54 |
|
|
|
| 9 |
A' |
1280 |
1209 |
35.77 |
|
|
|
| 10 |
A' |
1067 |
1008 |
28.93 |
|
|
|
| 11 |
A' |
942 |
890 |
5.67 |
|
|
|
| 12 |
A' |
483 |
456 |
17.05 |
|
|
|
| 13 |
A" |
3185 |
3008 |
7.23 |
|
|
|
| 14 |
A" |
1508 |
1425 |
7.56 |
|
|
|
| 15 |
A" |
1134 |
1071 |
7.98 |
|
|
|
| 16 |
A" |
1090 |
1030 |
9.85 |
|
|
|
| 17 |
A" |
688 |
650 |
49.92 |
|
|
|
| 18 |
A" |
198 |
187 |
1.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15281.7 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 14433.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.037 |
-0.625 |
0.000 |
| C2 |
0.000 |
0.453 |
0.000 |
| N3 |
1.247 |
0.170 |
0.000 |
| H4 |
-0.555 |
-1.601 |
0.000 |
| H5 |
-1.680 |
-0.535 |
0.878 |
| H6 |
-1.680 |
-0.535 |
-0.878 |
| H7 |
-0.381 |
1.481 |
0.000 |
| H8 |
1.793 |
1.032 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4962 | 2.4180 | 1.0885 | 1.0925 | 1.0925 | 2.2064 | 3.2798 |
C2 | 1.4962 | | 1.2785 | 2.1280 | 2.1384 | 2.1384 | 1.0963 | 1.8843 | N3 | 2.4180 | 1.2785 | | 2.5259 | 3.1362 | 3.1362 | 2.0904 | 1.0212 | H4 | 1.0885 | 2.1280 | 2.5259 | | 1.7817 | 1.7817 | 3.0873 | 3.5280 | H5 | 1.0925 | 2.1384 | 3.1362 | 1.7817 | | 1.7568 | 2.5548 | 3.9108 | H6 | 1.0925 | 2.1384 | 3.1362 | 1.7817 | 1.7568 | | 2.5548 | 3.9108 | H7 | 2.2064 | 1.0963 | 2.0904 | 3.0873 | 2.5548 | 2.5548 | | 2.2201 | H8 | 3.2798 | 1.8843 | 1.0212 | 3.5280 | 3.9108 | 3.9108 | 2.2201 | |
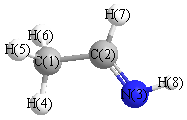 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.051 |
|
C1 |
C2 |
H7 |
115.790 |
| C2 |
C1 |
H4 |
109.827 |
|
C2 |
C1 |
H5 |
110.416 |
| C2 |
C1 |
H6 |
110.416 |
|
C2 |
N3 |
H8 |
109.537 |
| N3 |
C2 |
H7 |
123.159 |
|
H4 |
C1 |
H5 |
109.552 |
| H4 |
C1 |
H6 |
109.552 |
|
H5 |
C1 |
H6 |
107.033 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -133.601864 |
| Energy at 298.15K | -133.607317 |
| HF Energy | -133.090051 |
| Nuclear repulsion energy | 70.794844 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3478 |
3285 |
1.04 |
|
|
|
| 2 |
A' |
3206 |
3028 |
13.44 |
|
|
|
| 3 |
A' |
3155 |
2980 |
21.17 |
|
|
|
| 4 |
A' |
3098 |
2926 |
4.49 |
|
|
|
| 5 |
A' |
1704 |
1609 |
36.83 |
|
|
|
| 6 |
A' |
1507 |
1424 |
22.34 |
|
|
|
| 7 |
A' |
1441 |
1361 |
20.54 |
|
|
|
| 8 |
A' |
1415 |
1336 |
5.03 |
|
|
|
| 9 |
A' |
1283 |
1212 |
54.41 |
|
|
|
| 10 |
A' |
1076 |
1016 |
24.35 |
|
|
|
| 11 |
A' |
925 |
874 |
3.75 |
|
|
|
| 12 |
A' |
491 |
463 |
7.54 |
|
|
|
| 13 |
A" |
3186 |
3009 |
5.16 |
|
|
|
| 14 |
A" |
1502 |
1418 |
7.54 |
|
|
|
| 15 |
A" |
1166 |
1101 |
40.35 |
|
|
|
| 16 |
A" |
1082 |
1022 |
9.68 |
|
|
|
| 17 |
A" |
692 |
654 |
8.66 |
|
|
|
| 18 |
A" |
176 |
166 |
0.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15291.1 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 14442.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.024 |
-0.614 |
0.000 |
| C2 |
0.000 |
0.483 |
0.000 |
| N3 |
1.273 |
0.367 |
0.000 |
| H4 |
-0.552 |
-1.597 |
0.000 |
| H5 |
-1.669 |
-0.531 |
0.878 |
| H6 |
-1.669 |
-0.531 |
-0.878 |
| H7 |
-0.396 |
1.501 |
0.000 |
| H8 |
1.521 |
-0.626 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5008 | 2.4975 | 1.0909 | 1.0926 | 1.0926 | 2.2065 | 2.5451 |
C2 | 1.5008 | | 1.2781 | 2.1525 | 2.1414 | 2.1414 | 1.0922 | 1.8827 | N3 | 2.4975 | 1.2781 | | 2.6810 | 3.1986 | 3.1986 | 2.0177 | 1.0237 | H4 | 1.0909 | 2.1525 | 2.6810 | | 1.7762 | 1.7762 | 3.1025 | 2.2891 | H5 | 1.0926 | 2.1414 | 3.1986 | 1.7762 | | 1.7563 | 2.5540 | 3.3098 | H6 | 1.0926 | 2.1414 | 3.1986 | 1.7762 | 1.7563 | | 2.5540 | 3.3098 | H7 | 2.2065 | 1.0922 | 2.0177 | 3.1025 | 2.5540 | 2.5540 | | 2.8637 | H8 | 2.5451 | 1.8827 | 1.0237 | 2.2891 | 3.3098 | 3.3098 | 2.8637 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.800 |
|
C1 |
C2 |
H7 |
115.728 |
| C2 |
C1 |
H4 |
111.331 |
|
C2 |
C1 |
H5 |
110.332 |
| C2 |
C1 |
H6 |
110.332 |
|
C2 |
N3 |
H8 |
109.256 |
| N3 |
C2 |
H7 |
116.472 |
|
H4 |
C1 |
H5 |
108.874 |
| H4 |
C1 |
H6 |
108.874 |
|
H5 |
C1 |
H6 |
106.980 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
