Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Geometric Data calculated at QCISD/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -133.634667 |
| Energy at 298.15K | -133.640122 |
| HF Energy | -133.090392 |
| Nuclear repulsion energy | 70.776430 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3473 |
3290 |
2.68 |
|
|
|
| 2 |
A' |
3173 |
3006 |
22.40 |
|
|
|
| 3 |
A' |
3138 |
2973 |
18.70 |
|
|
|
| 4 |
A' |
3078 |
2916 |
7.03 |
|
|
|
| 5 |
A' |
1744 |
1652 |
50.01 |
|
|
|
| 6 |
A' |
1512 |
1433 |
20.95 |
|
|
|
| 7 |
A' |
1453 |
1376 |
21.45 |
|
|
|
| 8 |
A' |
1427 |
1352 |
3.28 |
|
|
|
| 9 |
A' |
1300 |
1231 |
56.00 |
|
|
|
| 10 |
A' |
1082 |
1025 |
21.66 |
|
|
|
| 11 |
A' |
925 |
876 |
2.80 |
|
|
|
| 12 |
A' |
493 |
467 |
7.74 |
|
|
|
| 13 |
A" |
3152 |
2986 |
9.60 |
|
|
|
| 14 |
A" |
1505 |
1426 |
6.62 |
|
|
|
| 15 |
A" |
1166 |
1105 |
40.12 |
|
|
|
| 16 |
A" |
1091 |
1033 |
9.68 |
|
|
|
| 17 |
A" |
698 |
661 |
9.73 |
|
|
|
| 18 |
A" |
172 |
163 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15290.4 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 14486.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.021 |
-0.622 |
0.000 |
| C2 |
0.000 |
0.485 |
0.000 |
| N3 |
1.269 |
0.375 |
0.000 |
| H4 |
-0.543 |
-1.604 |
0.000 |
| H5 |
-1.667 |
-0.544 |
0.880 |
| H6 |
-1.667 |
-0.544 |
-0.880 |
| H7 |
-0.405 |
1.500 |
0.000 |
| H8 |
1.530 |
-0.614 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5065 | 2.4980 | 1.0923 | 1.0940 | 1.0940 | 2.2096 | 2.5510 |
C2 | 1.5065 | | 1.2735 | 2.1589 | 2.1473 | 2.1473 | 1.0926 | 1.8834 | N3 | 2.4980 | 1.2735 | | 2.6837 | 3.1994 | 3.1994 | 2.0167 | 1.0228 | H4 | 1.0923 | 2.1589 | 2.6837 | | 1.7780 | 1.7780 | 3.1072 | 2.2974 | H5 | 1.0940 | 2.1473 | 3.1994 | 1.7780 | | 1.7596 | 2.5576 | 3.3160 | H6 | 1.0940 | 2.1473 | 3.1994 | 1.7780 | 1.7596 | | 2.5576 | 3.3160 | H7 | 2.2096 | 1.0926 | 2.0167 | 3.1072 | 2.5576 | 2.5576 | | 2.8655 | H8 | 2.5510 | 1.8834 | 1.0228 | 2.2974 | 3.3160 | 3.3160 | 2.8655 | |
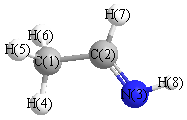 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.737 |
|
C1 |
C2 |
H7 |
115.535 |
| C2 |
C1 |
H4 |
111.355 |
|
C2 |
C1 |
H5 |
110.316 |
| C2 |
C1 |
H6 |
110.316 |
|
C2 |
N3 |
H8 |
109.726 |
| N3 |
C2 |
H7 |
116.727 |
|
H4 |
C1 |
H5 |
108.831 |
| H4 |
C1 |
H6 |
108.831 |
|
H5 |
C1 |
H6 |
107.075 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
