return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (2-Butanethiol, 3-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-594.872429
Energy at 298.15K-594.885482
HF Energy-593.871283
Nuclear repulsion energy317.122403
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3209 3031 14.45      
2 A 3205 3027 17.34      
3 A 3201 3024 10.76      
4 A 3193 3016 17.73      
5 A 3189 3012 29.69      
6 A 3184 3007 0.17      
7 A 3109 2936 20.15      
8 A 3102 2930 11.31      
9 A 3099 2927 12.99      
10 A 3097 2925 8.60      
11 A 3088 2917 1.62      
12 A 2782 2628 2.17      
13 A 1548 1462 3.12      
14 A 1532 1447 3.04      
15 A 1529 1445 18.99      
16 A 1518 1434 3.03      
17 A 1515 1431 3.38      
18 A 1510 1426 1.12      
19 A 1440 1360 5.95      
20 A 1425 1346 8.86      
21 A 1418 1339 3.22      
22 A 1392 1315 0.98      
23 A 1361 1285 0.41      
24 A 1329 1256 3.69      
25 A 1266 1195 23.70      
26 A 1219 1152 2.19      
27 A 1194 1128 6.65      
28 A 1175 1110 0.98      
29 A 1107 1046 0.86      
30 A 1060 1001 9.26      
31 A 995 940 2.94      
32 A 984 930 0.71      
33 A 952 900 1.91      
34 A 939 887 0.94      
35 A 909 859 3.63      
36 A 815 770 2.38      
37 A 717 677 1.28      
38 A 481 454 0.29      
39 A 410 387 0.44      
40 A 379 358 0.11      
41 A 373 352 0.32      
42 A 337 318 0.90      
43 A 262 248 1.24      
44 A 247 233 3.49      
45 A 236 223 0.57      
46 A 203 191 3.68      
47 A 186 175 11.53      
48 A 68 64 1.68      

Unscaled Zero Point Vibrational Energy (zpe) 35743.6 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 33759.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.12365 0.07180 0.04861

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.171 -1.643 0.121
H2 -2.073 -2.040 -0.348
H3 -0.340 -2.299 -0.140
H4 -1.308 -1.690 1.205
C5 0.546 1.862 -0.032
H6 -0.264 2.502 0.324
H7 1.473 2.235 0.405
H8 0.621 1.958 -1.117
S9 1.749 -0.620 -0.103
H10 2.676 0.196 0.410
C11 0.303 0.408 0.357
H12 0.171 0.331 1.442
C13 -0.936 -0.197 -0.320
H14 -0.759 -0.181 -1.402
C15 -2.183 0.634 -0.015
H16 -2.330 0.725 1.065
H17 -2.131 1.637 -0.438
H18 -3.068 0.148 -0.429

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 S9 H10 C11 H12 C13 H14 C15 H16 H17 H18
C11.09181.09001.09353.90584.24774.70224.20803.10164.27322.53652.72771.52922.15102.49572.80073.46302.6660
H21.09181.76391.76624.71044.93535.60574.88204.08455.30343.48393.72322.16572.50902.69773.11663.67922.4050
H31.09001.76391.76474.25624.82414.91404.47242.68093.95302.82693.11152.19172.50093.46653.81564.33473.6755
H41.09351.76621.76474.19364.40944.87694.73503.49264.47892.77802.51592.16623.06212.76732.62693.80103.0242
C53.90584.71044.25624.19361.09111.09131.09192.75912.74051.52462.15772.55352.78492.99233.28122.71614.0195
H64.24774.93534.82414.40941.09111.75951.77593.73853.73742.16932.48012.85543.22862.69932.82422.19363.7381
H74.70225.60574.91404.87691.09131.75951.76662.91262.36742.17022.52883.49963.75354.01364.14513.74935.0674
H84.20804.88204.47244.73501.09191.77591.76662.99073.10782.16223.06502.77552.56133.29073.87162.85224.1663
S93.10164.08452.68093.49262.75913.73852.91262.99071.33691.83262.40382.72602.85794.12794.45094.50064.8883
H104.27325.30343.95304.47892.74053.73742.36743.10781.33692.38312.71263.70553.90154.89725.07655.08915.8050
C112.53653.48392.82692.77801.52462.16932.17022.16221.83262.38311.09521.53602.13782.52382.74502.83973.4711
H122.72773.72323.11152.51592.15772.48012.52883.06502.40382.71261.09522.14643.03552.78452.55973.24553.7448
C131.52922.16572.19172.16622.55352.85543.49962.77552.72603.70551.53602.14641.09681.53022.17162.19282.1629
H142.15102.50902.50093.06212.78493.22863.75352.56132.85793.90152.13783.03551.09682.14953.06292.47412.5276
C152.49572.69773.46652.76732.99232.69934.01363.29074.12794.89722.52382.78451.53022.14951.09361.08971.0917
H162.80073.11663.81562.62693.28122.82424.14513.87164.45095.07652.74502.55972.17163.06291.09361.76901.7639
H173.46303.67924.33473.80102.71612.19363.74932.85224.50065.08912.83973.24552.19282.47411.08971.76901.7601
H182.66602.40503.67553.02424.01953.73815.06744.16634.88835.80503.47113.74482.16292.52761.09171.76391.7601

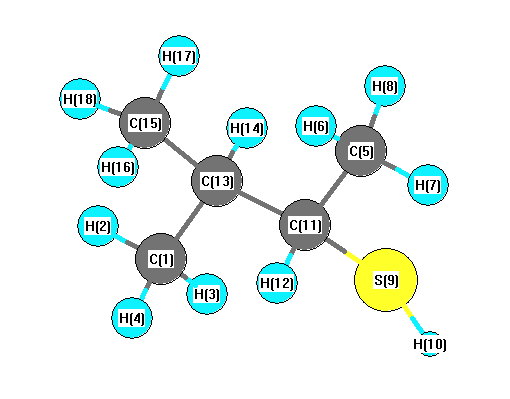
picture of 2-Butanethiol, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C13 C11 111.688 C1 C13 H14 108.873
C1 C13 C15 109.323 H2 C1 H3 107.892
H2 C1 H4 107.844 H2 C1 C13 110.320
H3 C1 H4 107.842 H3 C1 C13 112.523
H4 C1 C13 110.263 C5 C11 S9 110.203
C5 C11 H12 109.808 C5 C11 C13 113.083
H6 C5 H7 107.456 H6 C5 H8 108.878
H6 C5 C11 110.970 H7 C5 H8 108.036
H7 C5 C11 111.034 H8 C5 C11 110.361
S9 C11 H12 107.702 S9 C11 C13 107.714
H10 S9 C11 96.251 C11 C13 H14 107.400
C11 C13 C15 110.786 H12 C11 C13 108.157
C13 C15 H16 110.607 C13 C15 H17 112.563
C13 C15 H18 110.035 H14 C13 C15 108.692
H16 C15 H17 108.241 H16 C15 H18 107.635
H17 C15 H18 107.587
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability