Jump to
S1C2
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Geometric Data calculated at MP2/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -192.933303 |
| Energy at 298.15K | -192.938236 |
| HF Energy | -192.211138 |
| Nuclear repulsion energy | 149.457081 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3325 |
3140 |
0.06 |
|
|
|
| 2 |
A1 |
3310 |
3126 |
1.68 |
|
|
|
| 3 |
A1 |
3280 |
3098 |
0.78 |
|
|
|
| 4 |
A1 |
1579 |
1491 |
0.37 |
|
|
|
| 5 |
A1 |
1476 |
1394 |
4.08 |
|
|
|
| 6 |
A1 |
1185 |
1119 |
2.14 |
|
|
|
| 7 |
A1 |
1107 |
1045 |
9.13 |
|
|
|
| 8 |
A1 |
1073 |
1013 |
12.20 |
|
|
|
| 9 |
A1 |
849 |
802 |
0.30 |
|
|
|
| 10 |
A2 |
917 |
866 |
0.00 |
|
|
|
| 11 |
A2 |
875 |
827 |
0.00 |
|
|
|
| 12 |
A2 |
541 |
511 |
0.00 |
|
|
|
| 13 |
B1 |
963 |
910 |
4.73 |
|
|
|
| 14 |
B1 |
779 |
736 |
25.73 |
|
|
|
| 15 |
B1 |
707 |
668 |
58.91 |
|
|
|
| 16 |
B1 |
521 |
492 |
9.84 |
|
|
|
| 17 |
B2 |
5751 |
5431 |
55621.81 |
|
|
|
| 18 |
B2 |
3289 |
3107 |
37.10 |
|
|
|
| 19 |
B2 |
3278 |
3096 |
31.57 |
|
|
|
| 20 |
B2 |
1383 |
1306 |
98.95 |
|
|
|
| 21 |
B2 |
1306 |
1233 |
0.57 |
|
|
|
| 22 |
B2 |
1167 |
1102 |
10.28 |
|
|
|
| 23 |
B2 |
970 |
916 |
5.31 |
|
|
|
| 24 |
B2 |
856 |
808 |
8.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20241.7 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 19118.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.216 |
| C2 |
0.000 |
1.113 |
0.391 |
| C3 |
0.000 |
0.684 |
-1.001 |
| C4 |
0.000 |
-0.684 |
-1.001 |
| C5 |
0.000 |
-1.113 |
0.391 |
| H6 |
0.000 |
0.000 |
2.294 |
| H7 |
0.000 |
2.145 |
0.715 |
| H8 |
0.000 |
1.341 |
-1.857 |
| H9 |
0.000 |
-1.341 |
-1.857 |
| H10 |
0.000 |
-2.145 |
0.715 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3855 | 2.3202 | 2.3202 | 1.3855 | 1.0774 | 2.2024 | 3.3530 | 3.3530 | 2.2024 |
C2 | 1.3855 | | 1.4566 | 2.2730 | 2.2261 | 2.2041 | 1.0813 | 2.2596 | 3.3280 | 3.2738 | C3 | 2.3202 | 1.4566 | | 1.3678 | 2.2730 | 3.3647 | 2.2535 | 1.0792 | 2.1984 | 3.3084 | C4 | 2.3202 | 2.2730 | 1.3678 | | 1.4566 | 3.3647 | 3.3084 | 2.1984 | 1.0792 | 2.2535 | C5 | 1.3855 | 2.2261 | 2.2730 | 1.4566 | | 2.2041 | 3.2738 | 3.3280 | 2.2596 | 1.0813 | H6 | 1.0774 | 2.2041 | 3.3647 | 3.3647 | 2.2041 | | 2.6629 | 4.3618 | 4.3618 | 2.6629 | H7 | 2.2024 | 1.0813 | 2.2535 | 3.3084 | 3.2738 | 2.6629 | | 2.6948 | 4.3318 | 4.2893 | H8 | 3.3530 | 2.2596 | 1.0792 | 2.1984 | 3.3280 | 4.3618 | 2.6948 | | 2.6818 | 4.3318 | H9 | 3.3530 | 3.3280 | 2.1984 | 1.0792 | 2.2596 | 4.3618 | 4.3318 | 2.6818 | | 2.6948 | H10 | 2.2024 | 3.2738 | 3.3084 | 2.2535 | 1.0813 | 2.6629 | 4.2893 | 4.3318 | 2.6948 | |
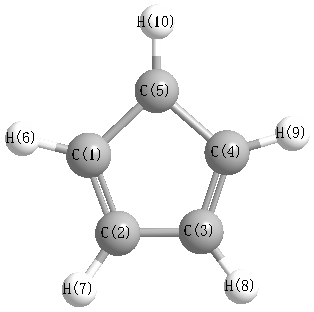 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
109.416 |
|
C1 |
C2 |
H7 |
126.015 |
| C1 |
C5 |
C4 |
109.416 |
|
C1 |
C5 |
H10 |
126.015 |
| C2 |
C1 |
C5 |
106.901 |
|
C2 |
C1 |
H6 |
126.550 |
| C2 |
C3 |
C4 |
107.134 |
|
C2 |
C3 |
H8 |
125.366 |
| C3 |
C2 |
H7 |
124.569 |
|
C3 |
C4 |
C5 |
107.134 |
| C3 |
C4 |
H9 |
127.501 |
|
C4 |
C3 |
H8 |
127.501 |
| C4 |
C5 |
H10 |
124.569 |
|
C5 |
C1 |
H6 |
126.550 |
| C5 |
C4 |
H9 |
125.366 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
