Jump to
S1C2
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Geometric Data calculated at MP2/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -235.180082 |
| Energy at 298.15K | -235.193568 |
| HF Energy | -234.198893 |
| Nuclear repulsion energy | 250.615587 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3189 |
3012 |
46.99 |
|
|
|
| 2 |
A' |
3185 |
3009 |
20.79 |
|
|
|
| 3 |
A' |
3178 |
3001 |
18.94 |
|
|
|
| 4 |
A' |
3131 |
2957 |
18.59 |
|
|
|
| 5 |
A' |
3127 |
2953 |
14.24 |
|
|
|
| 6 |
A' |
3109 |
2937 |
3.22 |
|
|
|
| 7 |
A' |
3094 |
2922 |
17.75 |
|
|
|
| 8 |
A' |
3090 |
2919 |
19.95 |
|
|
|
| 9 |
A' |
1529 |
1444 |
9.46 |
|
|
|
| 10 |
A' |
1524 |
1439 |
2.36 |
|
|
|
| 11 |
A' |
1517 |
1433 |
0.74 |
|
|
|
| 12 |
A' |
1429 |
1349 |
8.45 |
|
|
|
| 13 |
A' |
1425 |
1346 |
0.61 |
|
|
|
| 14 |
A' |
1396 |
1318 |
0.16 |
|
|
|
| 15 |
A' |
1376 |
1300 |
10.44 |
|
|
|
| 16 |
A' |
1287 |
1216 |
2.46 |
|
|
|
| 17 |
A' |
1268 |
1198 |
0.56 |
|
|
|
| 18 |
A' |
1214 |
1146 |
0.27 |
|
|
|
| 19 |
A' |
1164 |
1100 |
0.41 |
|
|
|
| 20 |
A' |
1067 |
1008 |
0.38 |
|
|
|
| 21 |
A' |
973 |
919 |
1.04 |
|
|
|
| 22 |
A' |
968 |
915 |
1.16 |
|
|
|
| 23 |
A' |
878 |
829 |
1.00 |
|
|
|
| 24 |
A' |
823 |
777 |
0.90 |
|
|
|
| 25 |
A' |
649 |
613 |
2.12 |
|
|
|
| 26 |
A' |
433 |
409 |
0.08 |
|
|
|
| 27 |
A' |
328 |
310 |
0.15 |
|
|
|
| 28 |
A' |
129 |
122 |
0.01 |
|
|
|
| 29 |
A" |
3185 |
3009 |
18.83 |
|
|
|
| 30 |
A" |
3183 |
3006 |
15.32 |
|
|
|
| 31 |
A" |
3176 |
3000 |
9.04 |
|
|
|
| 32 |
A" |
3107 |
2935 |
45.84 |
|
|
|
| 33 |
A" |
1526 |
1442 |
3.63 |
|
|
|
| 34 |
A" |
1525 |
1440 |
0.14 |
|
|
|
| 35 |
A" |
1492 |
1409 |
2.65 |
|
|
|
| 36 |
A" |
1305 |
1232 |
0.52 |
|
|
|
| 37 |
A" |
1284 |
1212 |
0.00 |
|
|
|
| 38 |
A" |
1239 |
1170 |
0.09 |
|
|
|
| 39 |
A" |
1154 |
1090 |
0.45 |
|
|
|
| 40 |
A" |
1145 |
1081 |
0.06 |
|
|
|
| 41 |
A" |
1003 |
947 |
3.28 |
|
|
|
| 42 |
A" |
928 |
876 |
0.00 |
|
|
|
| 43 |
A" |
919 |
868 |
0.00 |
|
|
|
| 44 |
A" |
816 |
771 |
0.34 |
|
|
|
| 45 |
A" |
369 |
348 |
0.01 |
|
|
|
| 46 |
A" |
262 |
248 |
0.14 |
|
|
|
| 47 |
A" |
254 |
240 |
0.01 |
|
|
|
| 48 |
A" |
238 |
224 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37293.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 35223.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.286 |
0.108 |
1.078 |
| C2 |
0.286 |
0.108 |
-1.078 |
| C3 |
-0.674 |
0.670 |
0.000 |
| C4 |
0.888 |
-0.831 |
0.000 |
| C5 |
-0.963 |
2.158 |
0.000 |
| C6 |
0.286 |
-2.226 |
0.000 |
| H7 |
1.001 |
0.864 |
1.413 |
| H8 |
-0.161 |
-0.366 |
1.954 |
| H9 |
1.001 |
0.864 |
-1.413 |
| H10 |
-0.161 |
-0.366 |
-1.954 |
| H11 |
-1.613 |
0.110 |
0.000 |
| H12 |
1.978 |
-0.896 |
0.000 |
| H13 |
-0.031 |
2.729 |
0.000 |
| H14 |
-0.806 |
-2.182 |
0.000 |
| H15 |
-1.536 |
2.449 |
-0.884 |
| H16 |
-1.536 |
2.449 |
0.884 |
| H17 |
0.599 |
-2.786 |
-0.885 |
| H18 |
0.599 |
-2.786 |
0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1550 | 1.5488 | 1.5505 | 2.6313 | 2.5710 | 1.0929 | 1.0924 | 2.6988 | 3.1009 | 2.1836 | 2.2428 | 2.8520 | 2.7564 | 3.5562 | 2.9728 | 3.5105 | 2.9171 |
C2 | 2.1550 | | 1.5488 | 1.5505 | 2.6313 | 2.5710 | 2.6988 | 3.1009 | 1.0929 | 1.0924 | 2.1836 | 2.2428 | 2.8520 | 2.7564 | 2.9728 | 3.5562 | 2.9171 | 3.5105 | C3 | 1.5488 | 1.5488 | | 2.1657 | 1.5160 | 3.0512 | 2.1998 | 2.2700 | 2.1998 | 2.2700 | 1.0932 | 3.0794 | 2.1576 | 2.8548 | 2.1654 | 2.1654 | 3.7874 | 3.7874 | C4 | 1.5505 | 1.5505 | 2.1657 | | 3.5153 | 1.5195 | 2.2092 | 2.2662 | 2.2092 | 2.2662 | 2.6717 | 1.0921 | 3.6770 | 2.1665 | 4.1726 | 4.1726 | 2.1654 | 2.1654 | C5 | 2.6313 | 2.6313 | 1.5160 | 3.5153 | | 4.5588 | 2.7437 | 3.2907 | 2.7437 | 3.2907 | 2.1487 | 4.2395 | 1.0928 | 4.3428 | 1.0928 | 1.0928 | 5.2596 | 5.2596 | C6 | 2.5710 | 2.5710 | 3.0512 | 1.5195 | 4.5588 | | 3.4720 | 2.7352 | 3.4720 | 2.7352 | 3.0110 | 2.1519 | 4.9660 | 1.0932 | 5.0950 | 5.0950 | 1.0925 | 1.0925 | H7 | 1.0929 | 2.6988 | 2.1998 | 2.2092 | 2.7437 | 3.4720 | | 1.7765 | 2.8256 | 3.7681 | 3.0655 | 2.4589 | 2.5579 | 3.8128 | 3.7714 | 3.0379 | 4.3314 | 3.7095 | H8 | 1.0924 | 3.1009 | 2.2700 | 2.2662 | 3.2907 | 2.7352 | 1.7765 | | 3.7681 | 3.9081 | 2.4801 | 2.9456 | 3.6626 | 2.7446 | 4.2265 | 3.3098 | 3.8073 | 2.7530 | H9 | 2.6988 | 1.0929 | 2.1998 | 2.2092 | 2.7437 | 3.4720 | 2.8256 | 3.7681 | | 1.7765 | 3.0655 | 2.4589 | 2.5579 | 3.8128 | 3.0379 | 3.7714 | 3.7095 | 4.3314 | H10 | 3.1009 | 1.0924 | 2.2700 | 2.2662 | 3.2907 | 2.7352 | 3.7681 | 3.9081 | 1.7765 | | 2.4801 | 2.9456 | 3.6626 | 2.7446 | 3.3098 | 4.2265 | 2.7530 | 3.8073 | H11 | 2.1836 | 2.1836 | 1.0932 | 2.6717 | 2.1487 | 3.0110 | 3.0655 | 2.4801 | 3.0655 | 2.4801 | | 3.7290 | 3.0599 | 2.4299 | 2.5013 | 2.5013 | 3.7498 | 3.7498 | H12 | 2.2428 | 2.2428 | 3.0794 | 1.0921 | 4.2395 | 2.1519 | 2.4589 | 2.9456 | 2.4589 | 2.9456 | 3.7290 | | 4.1449 | 3.0664 | 4.9310 | 4.9310 | 2.5014 | 2.5014 | H13 | 2.8520 | 2.8520 | 2.1576 | 3.6770 | 1.0928 | 4.9660 | 2.5579 | 3.6626 | 2.5579 | 3.6626 | 3.0599 | 4.1449 | | 4.9722 | 1.7675 | 1.7675 | 5.6214 | 5.6214 | H14 | 2.7564 | 2.7564 | 2.8548 | 2.1665 | 4.3428 | 1.0932 | 3.8128 | 2.7446 | 3.8128 | 2.7446 | 2.4299 | 3.0664 | 4.9722 | | 4.7705 | 4.7705 | 1.7665 | 1.7665 | H15 | 3.5562 | 2.9728 | 2.1654 | 4.1726 | 1.0928 | 5.0950 | 3.7714 | 4.2265 | 3.0379 | 3.3098 | 2.5013 | 4.9310 | 1.7675 | 4.7705 | | 1.7677 | 5.6532 | 5.9234 | H16 | 2.9728 | 3.5562 | 2.1654 | 4.1726 | 1.0928 | 5.0950 | 3.0379 | 3.3098 | 3.7714 | 4.2265 | 2.5013 | 4.9310 | 1.7675 | 4.7705 | 1.7677 | | 5.9234 | 5.6532 | H17 | 3.5105 | 2.9171 | 3.7874 | 2.1654 | 5.2596 | 1.0925 | 4.3314 | 3.8073 | 3.7095 | 2.7530 | 3.7498 | 2.5014 | 5.6214 | 1.7665 | 5.6532 | 5.9234 | | 1.7698 | H18 | 2.9171 | 3.5105 | 3.7874 | 2.1654 | 5.2596 | 1.0925 | 3.7095 | 2.7530 | 4.3314 | 3.8073 | 3.7498 | 2.5014 | 5.6214 | 1.7665 | 5.9234 | 5.6532 | 1.7698 | |
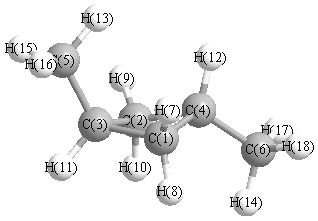 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.171 |
|
C1 |
C3 |
C5 |
118.310 |
| C1 |
C3 |
H11 |
110.292 |
|
C1 |
C4 |
C2 |
88.046 |
| C1 |
C4 |
C6 |
113.743 |
|
C1 |
C4 |
H12 |
115.039 |
| C2 |
C3 |
C5 |
118.310 |
|
C2 |
C3 |
H11 |
110.292 |
| C2 |
C4 |
C6 |
113.743 |
|
C2 |
C4 |
H12 |
115.039 |
| C3 |
C1 |
C4 |
88.659 |
|
C3 |
C1 |
H7 |
111.598 |
| C3 |
C1 |
H8 |
117.472 |
|
C3 |
C2 |
C4 |
88.659 |
| C3 |
C2 |
H9 |
111.598 |
|
C3 |
C2 |
H10 |
117.472 |
| C3 |
C5 |
H13 |
110.543 |
|
C3 |
C5 |
H15 |
111.169 |
| C3 |
C5 |
H16 |
111.169 |
|
C4 |
C1 |
H7 |
112.226 |
| C4 |
C1 |
H8 |
117.011 |
|
C4 |
C2 |
H9 |
112.226 |
| C4 |
C2 |
H10 |
117.011 |
|
C4 |
C6 |
H14 |
110.983 |
| C4 |
C6 |
H17 |
110.936 |
|
C4 |
C6 |
H18 |
110.936 |
| C5 |
C3 |
H11 |
109.813 |
|
C6 |
C4 |
H12 |
109.891 |
| H7 |
C1 |
H8 |
108.771 |
|
H9 |
C2 |
H10 |
108.771 |
| H13 |
C5 |
H15 |
107.936 |
|
H13 |
C5 |
H16 |
107.936 |
| H14 |
C6 |
H17 |
107.838 |
|
H14 |
C6 |
H18 |
107.838 |
| H15 |
C5 |
H16 |
107.954 |
|
H17 |
C6 |
H18 |
108.177 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
