Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -100.131296 |
| Energy at 298.15K | -100.130294 |
| HF Energy | -99.778062 |
| Nuclear repulsion energy | 27.418383 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
2078 |
1953 |
28.84 |
|
|
|
| 2 |
Σ |
635 |
597 |
109.65 |
|
|
|
| 3 |
Π |
150 |
141 |
39.59 |
|
|
|
| 3 |
Π |
150 |
141 |
39.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1506.4 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 1415.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.064 |
| C2 |
0.000 |
0.000 |
-0.160 |
| N3 |
0.000 |
0.000 |
1.021 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9037 | 3.0851 |
C2 | 1.9037 | | 1.1814 | N3 | 3.0851 | 1.1814 | |
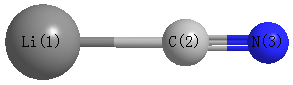 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -100.136981 |
| Energy at 298.15K | -100.136278 |
| HF Energy | -99.787836 |
| Nuclear repulsion energy | 29.129041 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
1.429 |
-0.593 |
0.000 |
| C2 |
-0.714 |
-0.375 |
0.000 |
| N3 |
0.000 |
0.575 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.1543 | 1.8453 |
C2 | 2.1543 | | 1.1887 | N3 | 1.8453 | 1.1887 | |
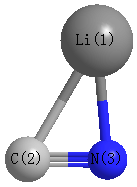 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
58.857 |
|
Li1 |
N3 |
C2 |
87.684 |
| C2 |
Li1 |
N3 |
33.459 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -100.133085 |
| Energy at 298.15K | -100.131932 |
| HF Energy | -99.791244 |
| Nuclear repulsion energy | 28.236623 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.883 |
| C2 |
0.000 |
0.000 |
-1.073 |
| N3 |
0.000 |
0.000 |
0.113 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9559 | 1.7695 |
C2 | 2.9559 | | 1.1864 | N3 | 1.7695 | 1.1864 | |
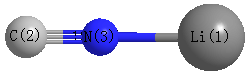 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
