return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Pentalene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-307.572402
Energy at 298.15K-307.578018
HF Energy-306.372196
Nuclear repulsion energy314.765040
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3293 3110 0.00      
2 Ag 3271 3089 0.00      
3 Ag 3253 3072 0.00      
4 Ag 1626 1536 0.00      
5 Ag 1539 1453 0.00      
6 Ag 1394 1317 0.00      
7 Ag 1322 1249 0.00      
8 Ag 1199 1132 0.00      
9 Ag 1110 1049 0.00      
10 Ag 980 925 0.00      
11 Ag 861 813 0.00      
12 Ag 710 671 0.00      
13 Ag 659 622 0.00      
14 Au 887 837 7.76      
15 Au 863 815 11.72      
16 Au 723 683 73.70      
17 Au 498 471 5.88      
18 Au 296 280 1.09      
19 Au 146 138 4.86      
20 Bg 905 854 0.00      
21 Bg 884 835 0.00      
22 Bg 774 731 0.00      
23 Bg 658 621 0.00      
24 Bg 540 510 0.00      
25 Bu 3293 3110 13.01      
26 Bu 3271 3090 3.08      
27 Bu 3254 3073 9.82      
28 Bu 1630 1540 36.72      
29 Bu 1518 1434 10.65      
30 Bu 1342 1268 16.53      
31 Bu 1271 1201 18.05      
32 Bu 1135 1072 34.92      
33 Bu 1058 999 4.40      
34 Bu 966 912 4.92      
35 Bu 839 792 5.02      
36 Bu 443 419 3.14      

Unscaled Zero Point Vibrational Energy (zpe) 24204.6 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 22861.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.17498 0.07238 0.05120

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.343 1.093 0.000
C2 2.163 -0.158 0.000
C3 1.343 -1.238 0.000
C4 -1.343 -1.093 0.000
C5 -2.163 0.158 0.000
C6 -1.343 1.238 0.000
C7 0.038 0.730 0.000
C8 -0.038 -0.730 0.000
H9 1.764 2.091 0.000
H10 3.245 -0.168 0.000
H11 1.639 -2.277 0.000
H12 -1.764 -2.091 0.000
H13 -3.245 0.168 0.000
H14 -1.639 2.277 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.49642.33113.46283.62842.68921.35442.28691.08302.28243.38334.44864.68023.2077
C21.49641.35623.62844.33803.77362.30352.27412.28421.08202.18304.37715.41824.5150
C32.33111.35622.68923.77363.65252.36121.47093.35542.18291.08063.22174.79834.6091
C43.46283.62842.68921.49642.33112.28691.35444.44864.68023.20771.08302.28243.3833
C53.62844.33803.77361.49641.35622.27412.30354.37715.41824.51502.28421.08202.1830
C62.68923.77363.65252.33111.35621.47092.36123.22174.79834.60913.35542.18291.0806
C71.35442.30352.36122.28692.27411.47091.46202.19813.33063.40673.34733.33082.2813
C82.28692.27411.47091.35442.30352.36121.46203.34733.33082.28132.19813.33063.4067
H91.08302.28423.35544.44864.37713.22172.19813.34732.70084.36985.47135.36573.4078
H102.28241.08202.18294.68025.41824.79833.33063.33082.70082.65165.36576.49895.4615
H113.38332.18301.08063.20774.51504.60913.40672.28134.36982.65163.40785.46155.6109
H124.44864.37713.22171.08302.28423.35543.34732.19815.47135.36573.40782.70084.3698
H134.68025.41824.79832.28241.08202.18293.33083.33065.36576.49895.46152.70082.6516
H143.20774.51504.60913.38332.18301.08062.28133.40673.40785.46155.61094.36982.6516

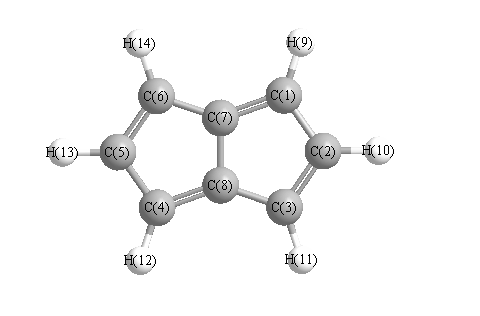
picture of Pentalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 109.511 C1 C2 H10 123.758
C1 C7 C6 144.246 C1 C7 C8 108.521
C2 C1 C7 107.701 C2 C1 H9 123.843
C2 C3 C8 107.034 C2 C3 H11 126.868
C3 C2 H10 126.731 C3 C8 C4 144.246
C3 C8 C7 107.233 C4 C5 C6 109.511
C4 C5 H13 123.758 C4 C8 C7 108.521
C5 C4 C8 107.701 C5 C4 H12 123.843
C5 C6 C7 107.034 C5 C6 H14 126.868
C6 C5 H13 126.731 C6 C7 C8 107.233
C7 C1 H9 128.456 C7 C6 H14 126.098
C8 C3 H11 126.098 C8 C4 H12 128.456
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability