return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3C(OH)=NH (Ethaninidic acid)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-208.742535
Energy at 298.15K-208.748925
HF Energy-207.981893
Nuclear repulsion energy121.811057
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3837 3635 54.29      
2 A' 3584 3396 2.90      
3 A' 3200 3032 9.47      
4 A' 3100 2937 4.23      
5 A' 1788 1694 200.47      
6 A' 1518 1438 24.10      
7 A' 1484 1406 60.83      
8 A' 1419 1345 1.08      
9 A' 1297 1229 93.92      
10 A' 1137 1077 176.34      
11 A' 1036 982 28.54      
12 A' 889 842 0.74      
13 A' 555 526 40.04      
14 A' 423 401 1.55      
15 A" 3176 3009 4.75      
16 A" 1502 1423 6.40      
17 A" 1086 1029 7.98      
18 A" 875 829 24.66      
19 A" 632 599 119.55      
20 A" 526 498 21.79      
21 A" 132 125 0.60      

Unscaled Zero Point Vibrational Energy (zpe) 16597.5 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 15724.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
0.36299 0.31118 0.17292

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.135 0.000
C2 0.908 -1.064 0.000
N3 0.295 1.369 0.000
O4 -1.297 -0.245 0.000
H5 1.955 -0.764 0.000
H6 0.706 -1.677 0.881
H7 0.706 -1.677 -0.881
H8 1.302 1.494 0.000
H9 -1.805 0.574 0.000

Atom - Atom Distances (Å)
  C1 C2 N3 O4 H5 H6 H7 H8 H9
C11.50341.26911.35122.15152.13452.13451.88241.8577
C21.50342.50892.35141.08941.09211.09212.58813.1687
N31.26912.50892.26692.70293.19753.19751.01502.2453
O41.35122.35142.26693.29272.61442.61443.12710.9641
H52.15151.08942.70293.29271.78061.78062.35043.9908
H62.13451.09213.19752.61441.78061.76163.34483.4851
H72.13451.09213.19752.61441.78061.76163.34483.4851
H81.88242.58811.01503.12712.35043.34483.34483.2404
H91.85773.16872.24530.96413.99083.48513.48513.2404

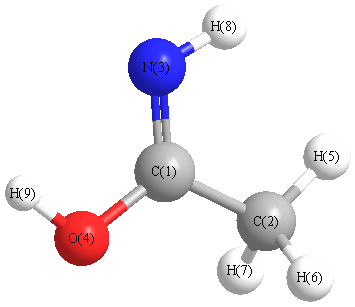
picture of Ethaninidic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 111.154 C1 C2 H6 109.631
C1 C2 H7 109.631 C1 N3 H8 110.502
C1 O4 H9 105.486 C2 C1 N3 129.430
C2 C1 O4 110.807 N3 C1 O4 119.763
H5 C2 H6 109.421 H5 C2 H7 109.421
H6 C2 H7 107.509
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability