return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2OH (aminomethanol)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes 1A
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-170.715868
Energy at 298.15K-170.722488
HF Energy-170.095163
Nuclear repulsion energy82.389698
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3893 3689 18.80      
2 A 3639 3447 2.04      
3 A 3552 3365 0.73      
4 A 3161 2995 26.87      
5 A 3083 2920 50.54      
6 A 1703 1613 23.88      
7 A 1545 1463 0.19      
8 A 1459 1382 44.40      
9 A 1420 1345 0.72      
10 A 1412 1338 3.36      
11 A 1192 1130 20.33      
12 A 1138 1079 26.26      
13 A 1070 1014 215.16      
14 A 928 879 3.46      
15 A 881 834 132.39      
16 A 484 458 44.92      
17 A 405 384 102.25      
18 A 298 282 59.51      

Unscaled Zero Point Vibrational Energy (zpe) 15630.9 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 14808.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
1.28378 0.32146 0.28833

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.229 -0.156 -0.020
C2 -0.036 0.537 0.047
O3 -1.193 -0.267 -0.115
H4 1.252 -0.721 -0.861
H5 1.321 -0.795 0.761
H6 -0.071 1.086 0.994
H7 -0.079 1.259 -0.770
H8 -1.265 -0.827 0.659

Atom - Atom Distances (Å)
  N1 C2 O3 H4 H5 H6 H7 H8
N11.44392.42571.01391.01372.06432.06782.6699
C21.44391.41802.01722.03211.09561.09091.9356
O32.42571.41802.59642.71442.07851.99900.9585
H41.01392.01722.59641.62572.90902.38812.9425
H51.01372.03212.71441.62572.35242.92022.5884
H62.06431.09562.07852.90902.35241.77322.2800
H72.06781.09091.99902.38812.92021.77322.7928
H82.66991.93560.95852.94252.58842.28002.7928

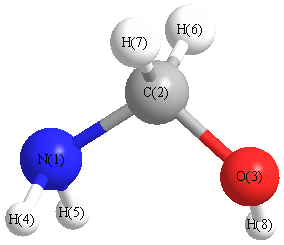
picture of aminomethanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 O3 115.896 N1 C2 H6 107.973
N1 C2 H7 108.526 C2 N1 H4 109.068
C2 N1 H5 110.326 C2 O3 H8 107.493
O3 C2 H6 110.913 O3 C2 H7 104.897
H4 N1 H5 106.602 H6 C2 H7 108.378
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability