return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5SC2H5 (Diethyl sulfide)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-555.664867
Energy at 298.15K-555.675474
HF Energy-554.834394
Nuclear repulsion energy234.262801
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3203 3026 16.12      
2 A1 3110 2938 2.95      
3 A1 3106 2933 34.08      
4 A1 1533 1448 3.65      
5 A1 1515 1431 0.20      
6 A1 1429 1350 2.25      
7 A1 1330 1256 8.56      
8 A1 1117 1055 0.14      
9 A1 1020 964 2.94      
10 A1 728 687 0.00      
11 A1 336 317 0.79      
12 A1 139 131 0.35      
13 A2 3213 3035 0.00      
14 A2 3169 2993 0.00      
15 A2 1523 1439 0.00      
16 A2 1277 1207 0.00      
17 A2 1050 992 0.00      
18 A2 790 747 0.00      
19 A2 255 241 0.00      
20 A2 86 82 0.00      
21 B1 3213 3034 23.40      
22 B1 3163 2988 11.57      
23 B1 1523 1438 14.15      
24 B1 1284 1212 0.02      
25 B1 1073 1014 0.01      
26 B1 808 763 7.75      
27 B1 250 236 0.29      
28 B1 62 58 0.92      
29 B2 3204 3026 7.95      
30 B2 3110 2938 22.87      
31 B2 3108 2935 7.96      
32 B2 1532 1447 0.46      
33 B2 1508 1424 9.24      
34 B2 1427 1348 2.15      
35 B2 1294 1222 41.79      
36 B2 1070 1011 3.17      
37 B2 1015 959 4.96      
38 B2 734 693 0.61      
39 B2 337 319 0.70      

Unscaled Zero Point Vibrational Energy (zpe) 29823.0 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 28167.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.47872 0.05816 0.05395

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 0.555
C2 0.000 1.377 -0.622
C3 0.000 -1.377 -0.622
C4 0.000 2.699 0.138
C5 0.000 -2.699 0.138
H6 0.885 1.299 -1.258
H7 -0.885 1.299 -1.258
H8 0.885 -1.299 -1.258
H9 -0.885 -1.299 -1.258
H10 0.000 3.537 -0.560
H11 0.000 -3.537 -0.560
H12 -0.883 2.781 0.772
H13 0.883 2.781 0.772
H14 0.883 -2.781 0.772
H15 -0.883 -2.781 0.772

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.81191.81192.73072.73072.39972.39972.39972.39973.70893.70892.92652.92652.92652.9265
C21.81192.75491.52464.14661.09251.09252.89032.89032.16094.91532.16722.16724.47464.4746
C31.81192.75494.14661.52462.89032.89031.09251.09254.91532.16094.47464.47462.16722.1672
C42.73071.52464.14665.39752.16592.16594.32654.32651.09126.27511.09051.09055.58705.5870
C52.73074.14661.52465.39754.32654.32652.16592.16596.27511.09125.58705.58701.09051.0905
H62.39971.09252.89032.16594.32651.77012.59893.14452.50594.96663.07352.51374.55814.8892
H72.39971.09252.89032.16594.32651.77013.14452.59892.50594.96662.51373.07354.88924.5581
H82.39972.89031.09254.32652.16592.59893.14451.77014.96662.50594.88924.55812.51373.0735
H92.39972.89031.09254.32652.16593.14452.59891.77014.96662.50594.55814.88923.07352.5137
H103.70892.16094.91531.09126.27512.50592.50594.96664.96667.07491.76811.76816.51796.5179
H113.70894.91532.16096.27511.09124.96664.96662.50592.50597.07496.51796.51791.76811.7681
H122.92652.16724.47461.09055.58703.07352.51374.88924.55811.76816.51791.76705.83685.5629
H132.92652.16724.47461.09055.58702.51373.07354.55814.88921.76816.51791.76705.56295.8368
H142.92654.47462.16725.58701.09054.55814.88922.51373.07356.51791.76815.83685.56291.7670
H152.92654.47462.16725.58701.09054.88924.55813.07352.51376.51791.76815.56295.83681.7670

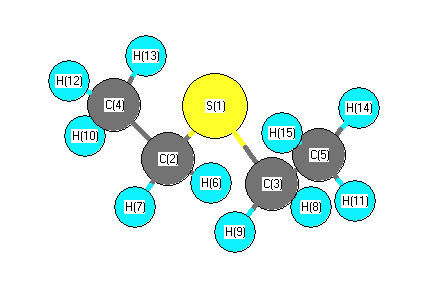
picture of Diethyl sulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C4 109.557 S1 C2 H6 108.893
S1 C2 H7 108.893 S1 C3 C5 109.557
S1 C3 H8 108.893 S1 C3 H9 108.893
C2 S1 C3 98.969 C2 C4 H10 110.302
C2 C4 H12 110.840 C2 C4 H13 110.840
C3 C5 H11 110.302 C3 C5 H14 110.840
C3 C5 H15 110.840 C4 C2 H6 110.623
C4 C2 H7 110.623 C5 C3 H8 110.623
C5 C3 H9 110.623 H6 C2 H7 108.206
H8 C3 H9 108.206 H10 C4 H12 108.276
H10 C4 H13 108.276 H11 C5 H14 108.276
H11 C5 H15 108.276 H12 C4 H13 108.216
H14 C5 H15 108.216
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability