return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (orthodifluorobenzene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-429.767706
Energy at 298.15K-429.772457
Nuclear repulsion energy348.194383
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3267 3086 1.35      
2 A1 3253 3072 3.48      
3 A1 1659 1567 33.39      
4 A1 1560 1473 192.04      
5 A1 1465 1384 5.29      
6 A1 1327 1254 80.30      
7 A1 1178 1113 0.79      
8 A1 1050 992 5.25      
9 A1 779 736 25.12      
10 A1 573 541 2.38      
11 A1 281 266 0.30      
12 A2 947 894 0.00      
13 A2 858 811 0.00      
14 A2 662 625 0.00      
15 A2 561 530 0.00      
16 A2 191 180 0.00      
17 B1 924 873 5.32      
18 B1 775 732 67.21      
19 B1 466 440 1.55      
20 B1 300 284 0.01      
21 B2 3260 3079 0.44      
22 B2 3243 3063 0.54      
23 B2 1658 1566 8.37      
24 B2 1488 1406 13.17      
25 B2 1283 1212 14.62      
26 B2 1234 1166 21.15      
27 B2 1124 1061 18.54      
28 B2 862 814 13.33      
29 B2 553 522 2.68      
30 B2 438 414 0.25      

Unscaled Zero Point Vibrational Energy (zpe) 18609.1 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 17576.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.10956 0.07443 0.04432

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.694 -0.537
C2 0.000 -0.694 -0.537
C3 0.000 -1.395 0.658
C4 0.000 -0.697 1.862
C5 0.000 0.697 1.862
C6 0.000 1.395 0.658
F7 0.000 1.344 -1.702
F8 0.000 -1.344 -1.702
H9 0.000 -2.476 0.623
H10 0.000 -1.243 2.796
H11 0.000 1.243 2.796
H12 0.000 2.476 0.623

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C11.38832.40652.77332.39941.38501.33352.34713.37623.85563.37852.1266
C21.38831.38502.39942.77332.40652.34711.33352.12663.37853.85563.3762
C32.40651.38501.39232.41362.78963.61462.35991.08222.14423.39603.8714
C42.77332.39941.39231.39342.41364.10673.62232.16881.08232.15303.4066
C52.39942.77332.41361.39341.39233.62234.10673.40662.15301.08232.1688
C61.38502.40652.78962.41361.39232.35993.61463.87143.39602.14421.0822
F71.33352.34713.61464.10673.62232.35992.68724.47195.18894.49942.5861
F82.34711.33352.35993.62234.10673.61462.68722.58614.49945.18894.4719
H93.37622.12661.08222.16883.40663.87144.47192.58612.49914.30804.9530
H103.85563.37852.14421.08232.15303.39605.18894.49942.49912.48614.3080
H113.37853.85563.39602.15301.08232.14424.49945.18894.30802.48612.4991
H122.12663.37623.87143.40662.16881.08222.58614.47194.95304.30802.4991

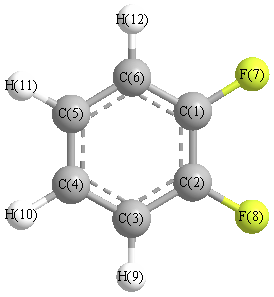
picture of orthodifluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.391 C1 C2 F8 119.146
C1 C6 C5 119.516 C1 C6 H12 118.557
C2 C1 C6 120.391 C2 C1 F7 119.146
C2 C3 C4 119.516 C2 C3 H9 118.557
C3 C2 F8 120.463 C3 C4 C5 120.093
C3 C4 H10 119.585 C4 C3 H9 121.927
C4 C5 C6 120.093 C4 C5 H11 120.322
C5 C4 H10 120.322 C5 C6 H12 121.927
C6 C1 F7 120.463 C6 C5 H11 119.585
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability