return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H11NO (Diethylhydroxylamine)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-288.239259
Energy at 298.15K-288.252774
HF Energy-287.150923
Nuclear repulsion energy265.193214
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3635 3433 0.71      
2 A 3231 3052 17.16      
3 A 3213 3035 17.95      
4 A 3149 2975 26.25      
5 A 3119 2946 1.73      
6 A 2990 2824 103.73      
7 A 1542 1456 8.95      
8 A 1518 1434 0.50      
9 A 1514 1430 19.62      
10 A 1452 1371 44.97      
11 A 1437 1357 17.03      
12 A 1407 1329 0.08      
13 A 1320 1247 4.21      
14 A 1204 1137 1.35      
15 A 1187 1121 11.52      
16 A 1112 1051 2.13      
17 A 973 919 13.51      
18 A 838 791 2.72      
19 A 798 754 12.81      
20 A 475 449 3.50      
21 A 432 408 0.06      
22 A 219 207 0.50      
23 A 210 198 1.71      
24 A 114 108 0.37      
25 A 3231 3051 0.11      
26 A 3213 3035 8.23      
27 A 3139 2965 0.29      
28 A 3119 2946 27.46      
29 A 2983 2817 2.61      
30 A 1539 1454 0.52      
31 A 1514 1430 0.84      
32 A 1504 1421 2.32      
33 A 1423 1344 21.36      
34 A 1391 1314 21.46      
35 A 1314 1241 1.24      
36 A 1223 1155 1.22      
37 A 1116 1054 3.03      
38 A 1093 1032 17.75      
39 A 975 921 0.37      
40 A 814 769 0.01      
41 A 550 519 22.47      
42 A 429 406 89.04      
43 A 352 332 5.92      
44 A 256 241 0.03      
45 A 138 130 0.19      

Unscaled Zero Point Vibrational Energy (zpe) 34202.5 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 32304.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.26117 0.06827 0.05983

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.253 0.000
O2 -1.322 -0.270 0.000
H3 -1.213 -1.239 0.000
C4 0.000 0.327 2.453
C5 0.000 0.327 -2.453
C6 0.667 -0.223 1.204
C7 0.667 -0.223 -1.204
H8 -1.038 0.006 2.501
H9 -1.038 0.006 -2.501
H10 0.682 -1.329 -1.226
H11 0.682 -1.329 1.226
H12 1.705 0.113 -1.144
H13 1.705 0.113 1.144
H14 0.021 1.417 2.436
H15 0.021 1.417 -2.436
H16 0.518 -0.020 3.348
H17 0.518 -0.020 -3.348

Atom - Atom Distances (Å)
  N1 O2 H3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
N11.42141.92302.45432.45431.45691.45692.71922.71922.11462.11462.05862.05862.69942.69943.39933.3993
O21.42140.97472.85012.85012.32542.32542.53212.53212.57662.57663.25873.25873.25273.25273.82863.8286
H31.92300.97473.15343.15342.45292.45292.79922.79922.25902.25903.41393.41393.80883.80883.96143.9614
C42.45432.85013.15344.90641.51893.75841.08845.07214.09192.17113.98692.16011.08995.00881.09105.8350
C52.45432.85013.15344.90643.75841.51895.07211.08842.17114.09192.16013.98695.00881.08995.83501.0910
C61.45692.32542.45291.51893.75842.40872.15464.08542.67001.10592.58991.09312.15044.04452.15874.5597
C71.45692.32542.45293.75841.51892.40874.08542.15461.10592.67001.09312.58994.04452.15044.55972.1587
H82.71922.53212.79921.08845.07212.15464.08545.00194.31662.52364.56383.06281.76515.24221.77246.0529
H92.71922.53212.79925.07211.08844.08542.15465.00192.52364.31663.06284.56385.24221.76516.05291.7724
H102.11462.57662.25904.09192.17112.67001.10594.31662.52362.45191.76992.95704.62433.07254.76062.4987
H112.11462.57662.25902.17114.09191.10592.67002.52364.31662.45192.95701.76993.07254.62432.49874.7606
H122.05863.25873.41393.98692.16012.58991.09314.56383.06281.76992.95702.28864.16602.49144.64892.5071
H132.05863.25873.41392.16013.98691.09312.58993.06284.56382.95701.76992.28862.49144.16602.50714.6489
H142.69943.25273.80881.08995.00882.15044.04451.76515.24224.62433.07254.16602.49144.87131.77395.9807
H152.69943.25273.80885.00881.08994.04452.15045.24221.76513.07254.62432.49144.16604.87135.98071.7739
H163.39933.82863.96141.09105.83502.15874.55971.77246.05294.76062.49874.64892.50711.77395.98076.6968
H173.39933.82863.96145.83501.09104.55972.15876.05291.77242.49874.76062.50714.64895.98071.77396.6968

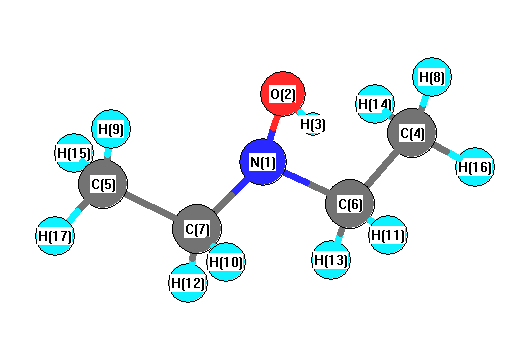
picture of Diethylhydroxylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 O2 H3 105.218 N1 C6 C4 111.113
N1 C6 H11 110.450 N1 C6 H13 106.795
N1 C7 C5 111.113 N1 C7 H10 110.450
N1 C7 H12 106.795 O2 N1 C6 107.773
O2 N1 C7 107.773 C4 C6 H11 110.632
C4 C6 H13 110.523 C5 C7 H10 110.632
C5 C7 H12 110.523 C6 N1 C7 111.517
C6 C4 H8 110.365 C6 C4 H14 109.940
C6 C4 H16 110.542 C7 C5 H9 110.365
C7 C5 H15 109.940 C7 C5 H17 110.542
H8 C4 H14 108.251 H8 C4 H16 108.830
H9 C5 H15 108.251 H9 C5 H17 108.830
H10 C7 H12 107.187 H11 C6 H13 107.187
H14 C4 H16 108.860 H15 C5 H17 108.860
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability