return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for H2NCHCHNH2 (diaminoethylene)

using model chemistry: CCD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at CCD/6-31G(2df,p)
 hartrees
Energy at 0K-188.853512
Energy at 298.15K 
HF Energy-188.112518
Nuclear repulsion energy117.590871
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3668 3475 1.74      
2 A 3573 3384 2.58      
3 A 3211 3041 0.04      
4 A 1809 1714 0.00      
5 A 1680 1592 24.38      
6 A 1364 1292 0.71      
7 A 1314 1244 0.29      
8 A 1061 1005 1.87      
9 A 978 927 52.37      
10 A 870 824 113.00      
11 A 560 531 0.62      
12 A 320 304 0.32      
13 A 253 240 15.00      
14 B 3668 3474 5.28      
15 B 3572 3383 1.89      
16 B 3211 3042 35.12      
17 B 1688 1599 45.38      
18 B 1424 1349 7.79      
19 B 1202 1139 62.33      
20 B 1152 1091 0.24      
21 B 864 818 196.80      
22 B 794 752 196.25      
23 B 341 323 33.77      
24 B 254 240 86.05      

Unscaled Zero Point Vibrational Energy (zpe) 19416.1 cm-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 18390.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G(2df,p)
ABC
1.43836 0.13253 0.12326

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G(2df,p)

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.314 0.588 0.024
C2 -0.314 -0.588 0.024
N3 -0.314 1.854 -0.107
N4 0.314 -1.854 -0.107
H5 1.399 0.632 0.024
H6 -1.399 -0.632 0.024
H7 -1.320 1.771 -0.034
H8 1.320 -1.771 -0.034
H9 -0.004 2.491 0.616
H10 0.004 -2.491 0.616

Atom - Atom Distances (Å)
  C1 C2 N3 N4 H5 H6 H7 H8 H9 H10
C11.33271.41922.44531.08612.10282.01802.56532.01823.1503
C21.33272.44531.41922.10281.08612.56532.01803.15032.0182
N31.41922.44533.76072.10832.71551.01183.97701.01184.4159
N42.44531.41923.76072.71552.10833.97701.01184.41591.0118
H51.08612.10282.10832.71553.07022.94842.40522.40323.4708
H62.10281.08612.71552.10833.07022.40522.94843.47082.4032
H72.01802.56531.01183.97702.94842.40524.41781.63404.5101
H82.56532.01803.97701.01182.40522.94844.41784.51011.6340
H92.01823.15031.01184.41592.40323.47081.63404.51014.9816
H103.15032.01824.41591.01183.47082.40324.51011.63404.9816

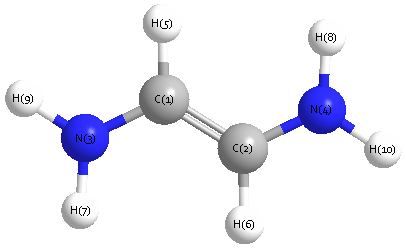
picture of diaminoethylene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N4 125.360 C1 C2 H6 120.420
C1 N3 H7 111.112 C1 N3 H9 111.133
C2 C1 N3 125.360 C2 C1 H5 120.420
C2 N4 H8 111.112 C2 N4 H10 111.133
N3 C1 H5 113.942 N4 C2 H6 113.942
H7 N3 H9 107.697 H8 N4 H10 107.697
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability