return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (Ethyl propyl sulfide)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-594.868188
Energy at 298.15K-594.880980
HF Energy-593.873834
Nuclear repulsion energy300.754043
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3203 3026 12.00      
2 A' 3199 3022 18.67      
3 A' 3114 2941 16.76      
4 A' 3110 2938 16.36      
5 A' 3107 2935 17.73      
6 A' 3104 2931 16.95      
7 A' 3093 2921 14.83      
8 A' 1537 1451 3.61      
9 A' 1533 1448 2.13      
10 A' 1521 1437 0.50      
11 A' 1513 1429 1.24      
12 A' 1504 1421 9.01      
13 A' 1431 1351 1.64      
14 A' 1428 1349 2.62      
15 A' 1384 1307 0.82      
16 A' 1316 1243 16.07      
17 A' 1268 1198 28.36      
18 A' 1139 1075 2.83      
19 A' 1086 1026 4.18      
20 A' 1078 1018 0.56      
21 A' 1018 961 4.25      
22 A' 930 879 2.40      
23 A' 801 757 1.08      
24 A' 730 689 0.30      
25 A' 386 365 0.22      
26 A' 308 291 0.61      
27 A' 273 258 0.63      
28 A' 105 100 0.34      
29 A" 3213 3034 11.16      
30 A" 3199 3021 27.26      
31 A" 3172 2996 2.95      
32 A" 3166 2990 8.70      
33 A" 3150 2975 2.80      
34 A" 1530 1445 6.75      
35 A" 1523 1439 7.23      
36 A" 1336 1262 0.17      
37 A" 1281 1210 0.01      
38 A" 1258 1188 0.00      
39 A" 1085 1025 0.86      
40 A" 1057 998 0.16      
41 A" 879 830 0.03      
42 A" 799 755 4.66      
43 A" 757 715 2.98      
44 A" 255 241 0.07      
45 A" 250 236 0.07      
46 A" 124 117 0.99      
47 A" 74 70 0.48      
48 A" 53 50 0.28      

Unscaled Zero Point Vibrational Energy (zpe) 36188.8 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 34180.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.38371 0.03448 0.03260

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.361 2.260 0.000
H2 3.452 2.244 0.000
H3 2.027 2.805 0.883
H4 2.027 2.805 -0.883
C5 1.812 0.837 0.000
H6 2.150 0.294 -0.885
H7 2.150 0.294 0.885
S8 0.000 0.887 0.000
C9 -0.331 -0.894 0.000
H10 0.129 -1.342 -0.886
H11 0.129 -1.342 0.886
C12 -1.834 -1.148 0.000
H13 -2.279 -0.672 0.878
H14 -2.279 -0.672 -0.878
C15 -2.146 -2.643 0.000
H16 -1.726 -3.130 -0.882
H17 -1.726 -3.130 0.882
H18 -3.222 -2.821 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 S8 C9 H10 H11 C12 H13 H14 C15 H16 H17 H18
C11.09121.09061.09061.52452.16602.16602.73094.14614.32914.32915.40495.55865.55866.65936.82126.82127.5487
H21.09121.76821.76822.16092.50612.50613.70924.91494.96904.96906.28126.49036.49037.43107.51477.51478.3786
H31.09061.76821.76702.16713.07352.51382.92654.47414.89184.56075.59565.53445.80796.91857.24017.02157.7445
H41.09061.76821.76702.16712.51383.07352.92654.47414.56074.89185.59565.80795.53446.91857.02157.24017.7445
C51.52452.16092.16712.16711.09251.09251.81232.75452.89282.89284.15164.44814.44815.27025.38835.38836.2228
H62.16602.50613.07352.51381.09251.77012.40002.88972.60093.14644.32924.86464.53395.27895.17205.46576.2731
H72.16602.50612.51383.07351.09251.77012.40002.88973.14642.60094.32924.53394.86465.27895.46575.17206.2731
S82.73093.70922.92652.92651.81232.40002.40001.81162.40252.40252.73982.89782.89784.13114.46034.46034.9124
C94.14614.91494.47414.47412.75452.88972.88971.81161.09381.09381.52512.14882.14882.52072.77932.77933.4748
H104.32914.96904.89184.56072.89282.60093.14642.40251.09381.77122.16253.05912.49962.76592.57603.12433.7683
H114.32914.96904.56074.89182.89283.14642.60092.40251.09381.77122.16252.49963.05912.76593.12432.57603.7683
C125.40496.28125.59565.59564.15164.32924.32922.73981.52512.16252.16251.09311.09311.52712.17212.17212.1736
H135.55866.49035.53445.80794.44814.86464.53392.89782.14883.05912.49961.09311.75572.16153.07312.51912.5053
H145.55866.49035.80795.53444.44814.53394.86462.89782.14882.49963.05911.09311.75572.16152.51913.07312.5053
C156.65937.43106.91856.91855.27025.27895.27894.13112.52072.76592.76591.52712.16152.16151.09181.09181.0909
H166.82127.51477.24017.02155.38835.17205.46574.46032.77932.57603.12432.17213.07312.51911.09181.76481.7642
H176.82127.51477.02157.24015.38835.46575.17204.46032.77933.12432.57602.17212.51913.07311.09181.76481.7642
H187.54878.37867.74457.74456.22286.27316.27314.91243.47483.76833.76832.17362.50532.50531.09091.76421.7642

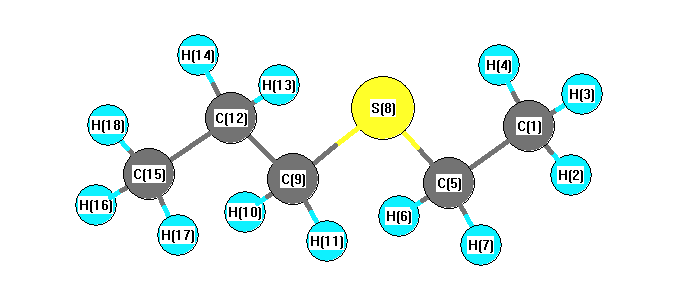
picture of Ethyl propyl sulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.633 C1 C5 H7 110.633
C1 C5 S8 109.550 H2 C1 H3 108.276
H2 C1 H4 108.276 H2 C1 C5 110.306
H3 C1 H4 108.216 H3 C1 C5 110.838
H4 C1 C5 110.838 C5 S8 C9 98.943
H6 C5 H7 108.208 H6 C5 S8 108.886
H7 C5 S8 108.886 S8 C9 H10 109.050
S8 C9 H11 109.050 S8 C9 C12 110.096
C9 C12 H13 109.199 C9 C12 H14 109.199
C9 C12 C15 111.352 H10 C9 H11 108.125
H10 C9 C12 110.239 H11 C9 C12 110.239
C12 C15 H16 110.989 C12 C15 H17 110.989
C12 C15 H18 111.158 H13 C12 H14 106.857
H13 C12 C15 110.060 H14 C12 C15 110.060
H16 C15 H17 107.847 H16 C15 H18 107.855
H17 C15 H18 107.855
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability