return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCHCHCH3 (2-Butenal)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-230.659613
Energy at 298.15K-230.665457
HF Energy-229.827113
Nuclear repulsion energy153.130079
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3242 3062 1.82      
2 A' 3213 3034 11.25      
3 A' 3191 3014 3.72      
4 A' 3099 2927 7.63      
5 A' 2949 2786 87.55      
6 A' 1772 1673 179.63      
7 A' 1710 1615 12.65      
8 A' 1513 1429 14.31      
9 A' 1440 1360 0.28      
10 A' 1427 1348 3.12      
11 A' 1336 1262 3.43      
12 A' 1280 1209 1.04      
13 A' 1180 1115 46.88      
14 A' 1115 1053 15.40      
15 A' 951 898 21.31      
16 A' 540 510 7.05      
17 A' 459 433 2.38      
18 A' 205 193 6.01      
19 A" 3176 3000 5.67      
20 A" 1507 1424 7.33      
21 A" 1078 1018 1.59      
22 A" 1045 987 6.11      
23 A" 1024 967 22.65      
24 A" 797 752 0.03      
25 A" 298 281 6.57      
26 A" 204 192 0.36      
27 A" 129 122 1.67      

Unscaled Zero Point Vibrational Energy (zpe) 19938.1 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 18831.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
1.10244 0.07277 0.06913

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.383 -0.473 0.000
C2 0.901 -0.649 0.000
C3 0.000 0.344 0.000
C4 -1.437 0.043 0.000
O5 -2.318 0.881 0.000
H6 -1.676 -1.041 0.000
H7 0.289 1.390 0.000
H8 0.530 -1.673 0.000
H9 2.660 0.581 0.000
H10 2.828 -0.950 0.877
H11 2.828 -0.950 -0.877

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.49282.51953.85494.89284.09912.80232.20831.08981.09291.0929
C21.49281.34102.43823.56462.60682.12851.08892.14692.13812.1381
C32.51951.34101.46802.37972.17451.08492.08522.67103.23113.2311
C43.85492.43821.46801.21661.11042.18942.60974.13244.46564.4656
O54.89283.56462.37971.21662.02732.65673.82544.98775.53215.5321
H64.09912.60682.17451.11042.02733.12612.29424.63014.58934.5893
H72.80232.12851.08492.18942.65673.12613.07172.50503.56183.5618
H82.20831.08892.08522.60973.82542.29423.07173.10182.56372.5637
H91.08982.14692.67104.13244.98774.63012.50503.10181.77271.7727
H101.09292.13813.23114.46565.53214.58933.56182.56371.77271.7539
H111.09292.13813.23114.46565.53214.58933.56182.56371.77271.7539

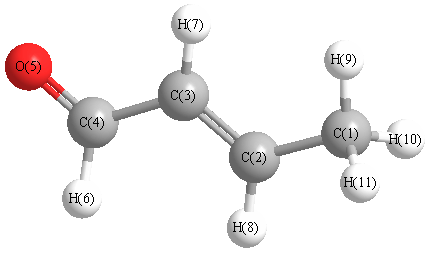
picture of 2-Butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.432 C1 C2 H8 116.730
C2 C1 H9 111.510 C2 C1 H10 110.615
C2 C1 H11 110.615 C2 C3 C4 120.381
C2 C3 H7 122.309 C3 C2 H8 117.837
C3 C4 O5 124.592 C3 C4 H6 114.278
C4 C3 H7 117.310 O5 C4 H6 121.130
H9 C1 H10 108.613 H9 C1 H11 108.613
H10 C1 H11 106.725
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability