Jump to
S1C2
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -148.462080 |
| Energy at 298.15K | -148.464412 |
| Nuclear repulsion energy | 59.127928 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3608 |
3418 |
41.14 |
|
|
|
| 2 |
A' |
2363 |
2239 |
65.47 |
|
|
|
| 3 |
A' |
1665 |
1577 |
38.26 |
|
|
|
| 4 |
A' |
1081 |
1024 |
5.57 |
|
|
|
| 5 |
A' |
685 |
649 |
213.38 |
|
|
|
| 6 |
A' |
495 |
469 |
25.60 |
|
|
|
| 7 |
A" |
3705 |
3510 |
55.08 |
|
|
|
| 8 |
A" |
1227 |
1163 |
0.00 |
|
|
|
| 9 |
A" |
413 |
391 |
1.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7620.8 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 7219.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.221 |
0.000 |
| N2 |
-0.047 |
1.382 |
0.000 |
| N3 |
0.120 |
-1.128 |
0.000 |
| H4 |
-0.255 |
-1.556 |
0.834 |
| H5 |
-0.255 |
-1.556 |
-0.834 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1623 | 1.3540 | 1.9791 | 1.9791 |
N2 | 1.1623 | | 2.5155 | 3.0611 | 3.0611 | N3 | 1.3540 | 2.5155 | | 1.0097 | 1.0097 | H4 | 1.9791 | 3.0611 | 1.0097 | | 1.6676 | H5 | 1.9791 | 3.0611 | 1.0097 | 1.6676 | |
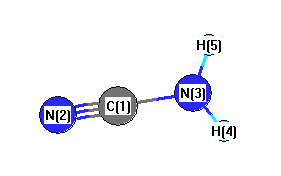 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
112.902 |
|
C1 |
N3 |
H5 |
112.902 |
| N2 |
C1 |
N3 |
177.237 |
|
H4 |
N3 |
H5 |
111.333 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -148.459671 |
| Energy at 298.15K | |
| HF Energy | -147.924660 |
| Nuclear repulsion energy | 59.373468 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3707 |
3512 |
81.26 |
|
|
|
| 2 |
A1 |
2364 |
2240 |
101.86 |
|
|
|
| 3 |
A1 |
1642 |
1556 |
47.82 |
|
|
|
| 4 |
A1 |
1128 |
1068 |
13.11 |
|
|
|
| 5 |
B1 |
534 |
506 |
1.09 |
|
|
|
| 6 |
B1 |
501i |
475i |
274.01 |
|
|
|
| 7 |
B2 |
3832 |
3631 |
99.24 |
|
|
|
| 8 |
B2 |
1148 |
1087 |
2.53 |
|
|
|
| 9 |
B2 |
407 |
386 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7130.2 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 6755.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.221 |
| N2 |
0.000 |
0.000 |
1.385 |
| N3 |
0.000 |
0.000 |
-1.112 |
| H4 |
0.000 |
0.864 |
-1.617 |
| H5 |
0.000 |
-0.864 |
-1.617 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1641 | 1.3322 | 2.0303 | 2.0303 |
N2 | 1.1641 | | 2.4963 | 3.1233 | 3.1233 | N3 | 1.3322 | 2.4963 | | 1.0006 | 1.0006 | H4 | 2.0303 | 3.1233 | 1.0006 | | 1.7274 | H5 | 2.0303 | 3.1233 | 1.0006 | 1.7274 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
120.328 |
|
C1 |
N3 |
H5 |
120.328 |
| N2 |
C1 |
N3 |
180.000 |
|
H4 |
N3 |
H5 |
119.345 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
