Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -376.791598 |
| Energy at 298.15K | |
| HF Energy | -375.807764 |
| Nuclear repulsion energy | 192.033797 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3196 |
3019 |
20.47 |
35.27 |
0.71 |
0.83 |
| 2 |
A |
3143 |
2969 |
30.13 |
67.90 |
0.32 |
0.49 |
| 3 |
A |
3123 |
2949 |
13.05 |
92.07 |
0.09 |
0.16 |
| 4 |
A |
1532 |
1447 |
3.03 |
6.70 |
0.73 |
0.85 |
| 5 |
A |
1494 |
1412 |
16.90 |
2.13 |
0.61 |
0.76 |
| 6 |
A |
1450 |
1369 |
21.85 |
3.36 |
0.74 |
0.85 |
| 7 |
A |
1378 |
1301 |
19.91 |
2.40 |
0.74 |
0.85 |
| 8 |
A |
1293 |
1221 |
12.67 |
6.81 |
0.72 |
0.83 |
| 9 |
A |
1207 |
1140 |
94.57 |
4.01 |
0.68 |
0.81 |
| 10 |
A |
1175 |
1110 |
146.05 |
2.21 |
0.73 |
0.84 |
| 11 |
A |
1167 |
1102 |
42.46 |
1.56 |
0.31 |
0.47 |
| 12 |
A |
1139 |
1076 |
26.90 |
5.07 |
0.35 |
0.52 |
| 13 |
A |
936 |
884 |
24.44 |
4.72 |
0.29 |
0.44 |
| 14 |
A |
588 |
555 |
3.75 |
1.62 |
0.32 |
0.49 |
| 15 |
A |
481 |
455 |
15.14 |
0.59 |
0.72 |
0.83 |
| 16 |
A |
433 |
409 |
5.54 |
1.19 |
0.49 |
0.66 |
| 17 |
A |
236 |
223 |
7.05 |
0.03 |
0.56 |
0.72 |
| 18 |
A |
126 |
119 |
6.42 |
0.01 |
0.69 |
0.81 |
Unscaled Zero Point Vibrational Energy (zpe) 12048.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 11379.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.763 |
-0.610 |
-0.267 |
| C2 |
0.471 |
0.021 |
0.330 |
| F3 |
-1.858 |
0.104 |
0.141 |
| F4 |
1.539 |
-0.733 |
-0.015 |
| F5 |
0.637 |
1.257 |
-0.182 |
| H6 |
-0.682 |
-0.578 |
-1.356 |
| H7 |
-0.848 |
-1.644 |
0.070 |
| H8 |
0.421 |
0.099 |
1.420 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5095 | 1.3697 | 2.3190 | 2.3349 | 1.0922 | 1.0916 | 2.1794 |
C2 | 1.5095 | | 2.3387 | 1.3525 | 1.3480 | 2.1292 | 2.1410 | 1.0930 | F3 | 1.3697 | 2.3387 | | 3.5025 | 2.7681 | 2.0226 | 2.0203 | 2.6135 | F4 | 2.3190 | 1.3525 | 3.5025 | | 2.1912 | 2.5988 | 2.5569 | 2.0005 | F5 | 2.3349 | 1.3480 | 2.7681 | 2.1912 | | 2.5466 | 3.2693 | 1.9884 | H6 | 1.0922 | 2.1292 | 2.0226 | 2.5988 | 2.5466 | | 1.7882 | 3.0626 | H7 | 1.0916 | 2.1410 | 2.0203 | 2.5569 | 3.2693 | 1.7882 | | 2.5443 | H8 | 2.1794 | 1.0930 | 2.6135 | 2.0005 | 1.9884 | 3.0626 | 2.5443 | |
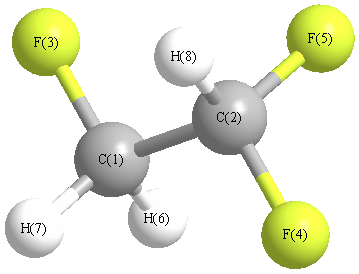 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.120 |
|
C1 |
C2 |
F5 |
109.468 |
| C1 |
C2 |
H8 |
112.759 |
|
C2 |
C1 |
F3 |
108.541 |
| C2 |
C1 |
H6 |
108.787 |
|
C2 |
C1 |
H7 |
109.758 |
| F3 |
C1 |
H6 |
109.969 |
|
F3 |
C1 |
H7 |
109.820 |
| F4 |
C2 |
F5 |
108.461 |
|
F4 |
C2 |
H8 |
109.312 |
| F5 |
C2 |
H8 |
108.635 |
|
H6 |
C1 |
H7 |
109.939 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -376.789837 |
| Energy at 298.15K | |
| HF Energy | -375.805432 |
| Nuclear repulsion energy | 195.178706 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3124 |
2951 |
56.62 |
103.55 |
0.04 |
0.09 |
| 2 |
A' |
3110 |
2937 |
5.12 |
48.66 |
0.49 |
0.66 |
| 3 |
A' |
1528 |
1443 |
4.90 |
5.93 |
0.73 |
0.85 |
| 4 |
A' |
1473 |
1391 |
26.65 |
2.13 |
0.43 |
0.61 |
| 5 |
A' |
1449 |
1369 |
19.16 |
2.90 |
0.72 |
0.84 |
| 6 |
A' |
1221 |
1154 |
102.93 |
4.91 |
0.35 |
0.52 |
| 7 |
A' |
1144 |
1081 |
15.02 |
2.43 |
0.75 |
0.86 |
| 8 |
A' |
908 |
858 |
26.78 |
7.17 |
0.16 |
0.28 |
| 9 |
A' |
777 |
734 |
43.52 |
2.29 |
0.47 |
0.64 |
| 10 |
A' |
525 |
496 |
11.10 |
1.19 |
0.75 |
0.85 |
| 11 |
A' |
237 |
223 |
1.41 |
0.08 |
0.37 |
0.53 |
| 12 |
A" |
3182 |
3005 |
17.26 |
45.01 |
0.75 |
0.86 |
| 13 |
A" |
1446 |
1366 |
29.62 |
1.46 |
0.75 |
0.86 |
| 14 |
A" |
1310 |
1237 |
18.21 |
11.74 |
0.75 |
0.86 |
| 15 |
A" |
1191 |
1125 |
93.66 |
0.84 |
0.75 |
0.86 |
| 16 |
A" |
990 |
935 |
43.50 |
2.92 |
0.75 |
0.86 |
| 17 |
A" |
373 |
352 |
0.18 |
0.11 |
0.75 |
0.86 |
| 18 |
A" |
121 |
114 |
1.71 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12053.8 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 11384.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.747 |
-0.824 |
0.000 |
| C2 |
0.357 |
0.633 |
0.000 |
| F3 |
-0.390 |
-1.583 |
0.000 |
| F4 |
-0.390 |
0.899 |
1.092 |
| F5 |
-0.390 |
0.899 |
-1.092 |
| H6 |
1.337 |
-1.043 |
-0.893 |
| H7 |
1.337 |
-1.043 |
0.893 |
| H8 |
1.233 |
1.290 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5089 | 1.3668 | 2.3361 | 2.3361 | 1.0927 | 1.0927 | 2.1696 |
C2 | 1.5089 | | 2.3384 | 1.3495 | 1.3495 | 2.1374 | 2.1374 | 1.0949 | F3 | 1.3668 | 2.3384 | | 2.7118 | 2.7118 | 2.0179 | 2.0179 | 3.2995 | F4 | 2.3361 | 1.3495 | 2.7118 | | 2.1838 | 3.2707 | 2.6068 | 1.9946 | F5 | 2.3361 | 1.3495 | 2.7118 | 2.1838 | | 2.6068 | 3.2707 | 1.9946 | H6 | 1.0927 | 2.1374 | 2.0179 | 3.2707 | 2.6068 | | 1.7869 | 2.5005 | H7 | 1.0927 | 2.1374 | 2.0179 | 2.6068 | 3.2707 | 1.7869 | | 2.5005 | H8 | 2.1696 | 1.0949 | 3.2995 | 1.9946 | 1.9946 | 2.5005 | 2.5005 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
109.501 |
|
C1 |
C2 |
F5 |
109.501 |
| C1 |
C2 |
H8 |
111.879 |
|
C2 |
C1 |
F3 |
108.709 |
| C2 |
C1 |
H6 |
109.443 |
|
C2 |
C1 |
H7 |
109.443 |
| F3 |
C1 |
H6 |
109.762 |
|
F3 |
C1 |
H7 |
109.762 |
| F4 |
C2 |
F5 |
108.026 |
|
F4 |
C2 |
H8 |
108.926 |
| F5 |
C2 |
H8 |
108.926 |
|
H6 |
C1 |
H7 |
109.704 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
