Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -187.584978 |
| Energy at 298.15K | -187.589044 |
| HF Energy | -186.899329 |
| Nuclear repulsion energy | 101.755295 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3687 |
3482 |
27.65 |
|
|
|
| 2 |
A |
3588 |
3389 |
4.51 |
|
|
|
| 3 |
A |
2412 |
2278 |
0.10 |
|
|
|
| 4 |
A |
1665 |
1573 |
12.25 |
|
|
|
| 5 |
A |
1218 |
1150 |
0.00 |
|
|
|
| 6 |
A |
827 |
781 |
12.96 |
|
|
|
| 7 |
A |
673 |
636 |
96.93 |
|
|
|
| 8 |
A |
398 |
376 |
2.19 |
|
|
|
| 9 |
A |
328 |
309 |
31.98 |
|
|
|
| 10 |
A |
216 |
204 |
14.07 |
|
|
|
| 11 |
B |
3687 |
3482 |
26.35 |
|
|
|
| 12 |
B |
3591 |
3391 |
21.97 |
|
|
|
| 13 |
B |
1664 |
1572 |
26.56 |
|
|
|
| 14 |
B |
1371 |
1295 |
81.27 |
|
|
|
| 15 |
B |
1218 |
1150 |
0.00 |
|
|
|
| 16 |
B |
728 |
688 |
338.36 |
|
|
|
| 17 |
B |
399 |
377 |
14.97 |
|
|
|
| 18 |
B |
217 |
205 |
16.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13942.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 13168.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.003 |
0.606 |
0.054 |
| C2 |
-0.003 |
-0.606 |
0.054 |
| N3 |
-0.003 |
1.956 |
-0.088 |
| N4 |
0.003 |
-1.956 |
-0.088 |
| H5 |
-0.314 |
2.455 |
0.731 |
| H6 |
0.862 |
2.339 |
-0.437 |
| H7 |
0.314 |
-2.455 |
0.731 |
| H8 |
-0.862 |
-2.339 |
-0.437 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2125 | 1.3573 | 2.5663 | 1.9945 | 1.9952 | 3.1510 | 3.1085 |
C2 | 1.2125 | | 2.5663 | 1.3573 | 3.1510 | 3.1085 | 1.9945 | 1.9952 | N3 | 1.3573 | 2.5663 | | 3.9123 | 1.0083 | 1.0084 | 4.4981 | 4.3938 | N4 | 2.5663 | 1.3573 | 3.9123 | | 4.4981 | 4.3938 | 1.0083 | 1.0084 | H5 | 1.9945 | 3.1510 | 1.0083 | 4.4981 | | 1.6620 | 4.9508 | 4.9646 | H6 | 1.9952 | 3.1085 | 1.0084 | 4.3938 | 1.6620 | | 4.9646 | 4.9852 | H7 | 3.1510 | 1.9945 | 4.4981 | 1.0083 | 4.9508 | 4.9646 | | 1.6620 | H8 | 3.1085 | 1.9952 | 4.3938 | 1.0084 | 4.9646 | 4.9852 | 1.6620 | |
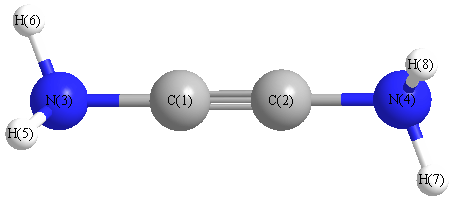 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
173.975 |
|
C1 |
N3 |
H5 |
114.135 |
| C1 |
N3 |
H6 |
114.182 |
|
C2 |
C1 |
N3 |
173.975 |
| C2 |
N4 |
H7 |
114.135 |
|
C2 |
N4 |
H8 |
114.182 |
| H5 |
N3 |
H6 |
110.996 |
|
H7 |
N4 |
H8 |
110.996 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -187.579690 |
| Energy at 298.15K | |
| HF Energy | -186.896346 |
| Nuclear repulsion energy | 102.357665 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3684 |
3479 |
0.00 |
|
|
|
| 2 |
A1 |
2430 |
2295 |
0.00 |
|
|
|
| 3 |
A1 |
1650 |
1558 |
0.00 |
|
|
|
| 4 |
A1 |
845 |
799 |
0.00 |
|
|
|
| 5 |
B1 |
490 |
463 |
0.00 |
|
|
|
| 6 |
B2 |
3689 |
3484 |
69.73 |
|
|
|
| 7 |
B2 |
1657 |
1565 |
53.04 |
|
|
|
| 8 |
B2 |
1445 |
1365 |
134.29 |
|
|
|
| 9 |
E |
3806 |
3595 |
52.82 |
|
|
|
| 9 |
E |
3806 |
3595 |
52.82 |
|
|
|
| 10 |
E |
1143 |
1079 |
1.47 |
|
|
|
| 10 |
E |
1143 |
1079 |
1.47 |
|
|
|
| 11 |
E |
418 |
395 |
0.93 |
|
|
|
| 11 |
E |
418 |
395 |
0.93 |
|
|
|
| 12 |
E |
190 |
179 |
24.60 |
|
|
|
| 12 |
E |
190 |
179 |
24.60 |
|
|
|
| 13 |
E |
514i |
485i |
243.37 |
|
|
|
| 13 |
E |
514i |
485i |
243.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12988.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12267.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.607 |
| C2 |
0.000 |
0.000 |
-0.607 |
| N3 |
0.000 |
0.000 |
1.944 |
| N4 |
0.000 |
0.000 |
-1.944 |
| H5 |
0.000 |
0.857 |
2.463 |
| H6 |
0.000 |
-0.857 |
2.463 |
| H7 |
0.857 |
0.000 |
-2.463 |
| H8 |
-0.857 |
0.000 |
-2.463 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2142 | 1.3371 | 2.5513 | 2.0438 | 2.0438 | 3.1870 | 3.1870 |
C2 | 1.2142 | | 2.5513 | 1.3371 | 3.1870 | 3.1870 | 2.0438 | 2.0438 | N3 | 1.3371 | 2.5513 | | 3.8884 | 1.0013 | 1.0013 | 4.4893 | 4.4893 | N4 | 2.5513 | 1.3371 | 3.8884 | | 4.4893 | 4.4893 | 1.0013 | 1.0013 | H5 | 2.0438 | 3.1870 | 1.0013 | 4.4893 | | 1.7133 | 5.0721 | 5.0721 | H6 | 2.0438 | 3.1870 | 1.0013 | 4.4893 | 1.7133 | | 5.0721 | 5.0721 | H7 | 3.1870 | 2.0438 | 4.4893 | 1.0013 | 5.0721 | 5.0721 | | 1.7133 | H8 | 3.1870 | 2.0438 | 4.4893 | 1.0013 | 5.0721 | 5.0721 | 1.7133 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.184 |
| C1 |
N3 |
H6 |
121.184 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.184 |
|
C2 |
N4 |
H8 |
121.184 |
| H5 |
N3 |
H6 |
117.632 |
|
H7 |
N4 |
H8 |
117.632 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
