Jump to
S1C2
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -187.987230 |
| Energy at 298.15K | -187.991377 |
| HF Energy | -187.987230 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3681 |
3681 |
22.44 |
|
|
|
| 2 |
A |
3589 |
3589 |
3.35 |
|
|
|
| 3 |
A |
2428 |
2428 |
0.07 |
|
|
|
| 4 |
A |
1657 |
1657 |
9.00 |
|
|
|
| 5 |
A |
1203 |
1203 |
0.00 |
|
|
|
| 6 |
A |
840 |
840 |
2.92 |
|
|
|
| 7 |
A |
567 |
567 |
125.02 |
|
|
|
| 8 |
A |
454 |
454 |
14.56 |
|
|
|
| 9 |
A |
406 |
406 |
18.00 |
|
|
|
| 10 |
A |
217 |
217 |
20.48 |
|
|
|
| 11 |
B |
3681 |
3681 |
21.79 |
|
|
|
| 12 |
B |
3591 |
3591 |
11.47 |
|
|
|
| 13 |
B |
1658 |
1658 |
28.95 |
|
|
|
| 14 |
B |
1390 |
1390 |
107.19 |
|
|
|
| 15 |
B |
1203 |
1203 |
0.02 |
|
|
|
| 16 |
B |
621 |
621 |
315.24 |
|
|
|
| 17 |
B |
459 |
459 |
31.39 |
|
|
|
| 18 |
B |
218 |
218 |
21.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13930.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13930.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.003 |
0.602 |
0.043 |
| C2 |
-0.003 |
-0.602 |
0.043 |
| N3 |
-0.003 |
1.951 |
-0.078 |
| N4 |
0.003 |
-1.951 |
-0.078 |
| H5 |
-0.338 |
2.452 |
0.731 |
| H6 |
0.848 |
2.351 |
-0.444 |
| H7 |
0.338 |
-2.452 |
0.731 |
| H8 |
-0.848 |
-2.351 |
-0.444 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2043 | 1.3543 | 2.5560 | 2.0031 | 2.0032 | 3.1489 | 3.1122 |
C2 | 1.2043 | | 2.5560 | 1.3543 | 3.1489 | 3.1122 | 2.0031 | 2.0032 | N3 | 1.3543 | 2.5560 | | 3.9020 | 1.0091 | 1.0093 | 4.4900 | 4.4000 | N4 | 2.5560 | 1.3543 | 3.9020 | | 4.4900 | 4.4000 | 1.0091 | 1.0093 | H5 | 2.0031 | 3.1489 | 1.0091 | 4.4900 | | 1.6729 | 4.9513 | 4.9716 | H6 | 2.0032 | 3.1122 | 1.0093 | 4.4000 | 1.6729 | | 4.9716 | 4.9997 | H7 | 3.1489 | 2.0031 | 4.4900 | 1.0091 | 4.9513 | 4.9716 | | 1.6729 | H8 | 3.1122 | 2.0032 | 4.4000 | 1.0093 | 4.9716 | 4.9997 | 1.6729 | |
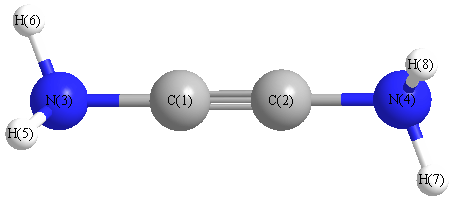 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
174.829 |
|
C1 |
N3 |
H5 |
115.107 |
| C1 |
N3 |
H6 |
115.110 |
|
C2 |
C1 |
N3 |
174.829 |
| C2 |
N4 |
H7 |
115.107 |
|
C2 |
N4 |
H8 |
115.110 |
| H5 |
N3 |
H6 |
111.956 |
|
H7 |
N4 |
H8 |
111.956 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.113 |
|
|
|
| 2 |
C |
0.113 |
|
|
|
| 3 |
N |
-0.671 |
|
|
|
| 4 |
N |
-0.671 |
|
|
|
| 5 |
H |
0.278 |
|
|
|
| 6 |
H |
0.280 |
|
|
|
| 7 |
H |
0.278 |
|
|
|
| 8 |
H |
0.280 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.441 |
1.441 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.940 |
4.056 |
0.000 |
| y |
4.056 |
-13.476 |
0.000 |
| z |
0.000 |
0.000 |
-23.066 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.668 |
4.056 |
0.000 |
| y |
4.056 |
9.527 |
0.000 |
| z |
0.000 |
0.000 |
-4.858 |
|
| Polar |
|---|
| 3z2-r2 | -9.717 |
|---|
| x2-y2 | -9.463 |
|---|
| xy | 4.056 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.181 |
0.235 |
0.000 |
| y |
0.235 |
8.886 |
0.000 |
| z |
0.000 |
0.000 |
3.179 |
<r2> (average value of r
2) Å
2
| <r2> |
96.313 |
| (<r2>)1/2 |
9.814 |
Jump to
S1C1
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -187.984840 |
| Energy at 298.15K | |
| HF Energy | -187.984840 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3649 |
3649 |
0.00 |
|
|
|
| 2 |
A1 |
2402 |
2402 |
0.00 |
|
|
|
| 3 |
A1 |
1643 |
1643 |
0.00 |
|
|
|
| 4 |
A1 |
861 |
861 |
0.00 |
|
|
|
| 5 |
B1 |
537 |
537 |
0.00 |
|
|
|
| 6 |
B2 |
3655 |
3655 |
28.87 |
|
|
|
| 7 |
B2 |
1653 |
1653 |
51.62 |
|
|
|
| 8 |
B2 |
1443 |
1443 |
143.02 |
|
|
|
| 9 |
E |
3760 |
3760 |
35.87 |
|
|
|
| 9 |
E |
3760 |
3760 |
35.87 |
|
|
|
| 10 |
E |
1143 |
1143 |
0.91 |
|
|
|
| 10 |
E |
1143 |
1143 |
0.91 |
|
|
|
| 11 |
E |
474 |
474 |
0.11 |
|
|
|
| 11 |
E |
474 |
474 |
0.11 |
|
|
|
| 12 |
E |
193 |
193 |
29.87 |
|
|
|
| 12 |
E |
193 |
193 |
29.87 |
|
|
|
| 13 |
E |
397i |
397i |
234.51 |
|
|
|
| 13 |
E |
397i |
397i |
234.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13094.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13094.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.604 |
| C2 |
0.000 |
0.000 |
-0.604 |
| N3 |
0.000 |
0.000 |
1.937 |
| N4 |
0.000 |
0.000 |
-1.937 |
| H5 |
0.000 |
0.857 |
2.459 |
| H6 |
0.000 |
-0.857 |
2.459 |
| H7 |
0.857 |
0.000 |
-2.459 |
| H8 |
-0.857 |
0.000 |
-2.459 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2087 | 1.3331 | 2.5417 | 2.0427 | 2.0427 | 3.1806 | 3.1806 |
C2 | 1.2087 | | 2.5417 | 1.3331 | 3.1806 | 3.1806 | 2.0427 | 2.0427 | N3 | 1.3331 | 2.5417 | | 3.8748 | 1.0028 | 1.0028 | 4.4788 | 4.4788 | N4 | 2.5417 | 1.3331 | 3.8748 | | 4.4788 | 4.4788 | 1.0028 | 1.0028 | H5 | 2.0427 | 3.1806 | 1.0028 | 4.4788 | | 1.7134 | 5.0644 | 5.0644 | H6 | 2.0427 | 3.1806 | 1.0028 | 4.4788 | 1.7134 | | 5.0644 | 5.0644 | H7 | 3.1806 | 2.0427 | 4.4788 | 1.0028 | 5.0644 | 5.0644 | | 1.7134 | H8 | 3.1806 | 2.0427 | 4.4788 | 1.0028 | 5.0644 | 5.0644 | 1.7134 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.319 |
| C1 |
N3 |
H6 |
121.319 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.319 |
|
C2 |
N4 |
H8 |
121.319 |
| H5 |
N3 |
H6 |
117.361 |
|
H7 |
N4 |
H8 |
117.361 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.129 |
|
|
|
| 2 |
C |
0.129 |
|
|
|
| 3 |
N |
-0.690 |
|
|
|
| 4 |
N |
-0.690 |
|
|
|
| 5 |
H |
0.281 |
|
|
|
| 6 |
H |
0.281 |
|
|
|
| 7 |
H |
0.281 |
|
|
|
| 8 |
H |
0.281 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.159 |
0.000 |
0.000 |
| y |
0.000 |
-23.159 |
0.000 |
| z |
0.000 |
0.000 |
-10.508 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.325 |
0.000 |
0.000 |
| y |
0.000 |
-6.325 |
0.000 |
| z |
0.000 |
0.000 |
12.651 |
|
| Polar |
|---|
| 3z2-r2 | 25.301 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.088 |
0.000 |
0.000 |
| y |
0.000 |
3.088 |
0.000 |
| z |
0.000 |
0.000 |
9.117 |
<r2> (average value of r
2) Å
2
| <r2> |
95.880 |
| (<r2>)1/2 |
9.792 |
