Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -153.983780 |
| Energy at 298.15K | |
| HF Energy | -153.463014 |
| Nuclear repulsion energy | 74.906776 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3873 |
3658 |
23.32 |
|
|
|
| 2 |
A' |
3242 |
3062 |
4.51 |
|
|
|
| 3 |
A' |
3066 |
2895 |
48.19 |
|
|
|
| 4 |
A' |
1556 |
1469 |
0.77 |
|
|
|
| 5 |
A' |
1508 |
1425 |
4.18 |
|
|
|
| 6 |
A' |
1457 |
1376 |
7.46 |
|
|
|
| 7 |
A' |
1255 |
1185 |
49.59 |
|
|
|
| 8 |
A' |
1087 |
1027 |
88.05 |
|
|
|
| 9 |
A' |
1027 |
970 |
22.02 |
|
|
|
| 10 |
A' |
606 |
572 |
14.27 |
|
|
|
| 11 |
A' |
373 |
352 |
32.00 |
|
|
|
| 12 |
A" |
3362 |
3175 |
4.39 |
|
|
|
| 13 |
A" |
3113 |
2941 |
42.91 |
|
|
|
| 14 |
A" |
1310 |
1237 |
0.27 |
|
|
|
| 15 |
A" |
1193 |
1127 |
0.98 |
|
|
|
| 16 |
A" |
817 |
772 |
0.19 |
|
|
|
| 17 |
A" |
268 |
253 |
116.84 |
|
|
|
| 18 |
A" |
93i |
88i |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14510.0 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 13704.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.107 |
-0.365 |
0.000 |
| C2 |
0.000 |
0.540 |
0.000 |
| C3 |
1.256 |
-0.258 |
0.000 |
| H4 |
-1.903 |
0.172 |
0.000 |
| H5 |
-0.032 |
1.186 |
0.885 |
| H6 |
-0.032 |
1.186 |
-0.885 |
| H7 |
1.643 |
-0.657 |
-0.925 |
| H8 |
1.643 |
-0.657 |
0.925 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4300 | 2.3654 | 0.9608 | 2.0848 | 2.0848 | 2.9163 | 2.9163 |
C2 | 1.4300 | | 1.4878 | 1.9387 | 1.0963 | 1.0963 | 2.2334 | 2.2334 | C3 | 2.3654 | 1.4878 | | 3.1885 | 2.1274 | 2.1274 | 1.0794 | 1.0794 | H4 | 0.9608 | 1.9387 | 3.1885 | | 2.3055 | 2.3055 | 3.7579 | 3.7579 | H5 | 2.0848 | 1.0963 | 2.1274 | 2.3055 | | 1.7701 | 3.0786 | 2.4905 | H6 | 2.0848 | 1.0963 | 2.1274 | 2.3055 | 1.7701 | | 2.4905 | 3.0786 | H7 | 2.9163 | 2.2334 | 1.0794 | 3.7579 | 3.0786 | 2.4905 | | 1.8504 | H8 | 2.9163 | 2.2334 | 1.0794 | 3.7579 | 2.4905 | 3.0786 | 1.8504 | |
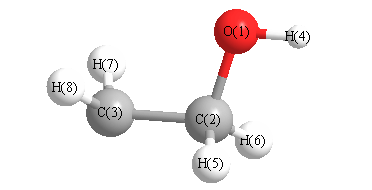 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.302 |
|
O1 |
C2 |
H5 |
110.533 |
| O1 |
C2 |
H6 |
110.533 |
|
C2 |
O1 |
H4 |
106.720 |
| C2 |
C3 |
H7 |
120.067 |
|
C2 |
C3 |
H8 |
120.067 |
| C3 |
C2 |
H5 |
109.900 |
|
C3 |
C2 |
H6 |
109.900 |
| H5 |
C2 |
H6 |
107.675 |
|
H7 |
C3 |
H8 |
117.996 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -153.985235 |
| Energy at 298.15K | -153.989936 |
| HF Energy | -153.464155 |
| Nuclear repulsion energy | 75.208989 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3895 |
3679 |
31.86 |
|
|
|
| 2 |
A |
3373 |
3186 |
1.78 |
|
|
|
| 3 |
A |
3248 |
3068 |
2.97 |
|
|
|
| 4 |
A |
3059 |
2889 |
47.45 |
|
|
|
| 5 |
A |
3001 |
2835 |
49.00 |
|
|
|
| 6 |
A |
1539 |
1454 |
3.81 |
|
|
|
| 7 |
A |
1496 |
1413 |
7.54 |
|
|
|
| 8 |
A |
1449 |
1369 |
1.46 |
|
|
|
| 9 |
A |
1293 |
1221 |
55.25 |
|
|
|
| 10 |
A |
1250 |
1181 |
9.78 |
|
|
|
| 11 |
A |
1150 |
1087 |
38.87 |
|
|
|
| 12 |
A |
1093 |
1032 |
29.05 |
|
|
|
| 13 |
A |
992 |
936 |
8.51 |
|
|
|
| 14 |
A |
885 |
836 |
13.87 |
|
|
|
| 15 |
A |
454 |
429 |
22.16 |
|
|
|
| 16 |
A |
398 |
376 |
35.04 |
|
|
|
| 17 |
A |
285 |
269 |
93.81 |
|
|
|
| 18 |
A |
165 |
155 |
17.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14513.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 13707.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.097 |
-0.384 |
-0.057 |
| C2 |
-0.014 |
0.528 |
0.035 |
| C3 |
1.241 |
-0.255 |
-0.020 |
| H4 |
-1.893 |
0.124 |
0.106 |
| H5 |
-0.068 |
1.102 |
0.974 |
| H6 |
-0.038 |
1.263 |
-0.781 |
| H7 |
2.179 |
0.238 |
-0.213 |
| H8 |
1.233 |
-1.289 |
0.284 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4184 | 2.3410 | 0.9589 | 2.0813 | 2.0880 | 3.3382 | 2.5227 |
C2 | 1.4184 | | 1.4801 | 1.9231 | 1.1026 | 1.0988 | 2.2265 | 2.2181 | C3 | 2.3410 | 1.4801 | | 3.1591 | 2.1319 | 2.1258 | 1.0780 | 1.0778 | H4 | 0.9589 | 1.9231 | 3.1591 | | 2.2453 | 2.3512 | 4.0866 | 3.4356 | H5 | 2.0813 | 1.1026 | 2.1319 | 2.2453 | | 1.7633 | 2.6851 | 2.8090 | H6 | 2.0880 | 1.0988 | 2.1258 | 2.3512 | 1.7633 | | 2.5076 | 3.0439 | H7 | 3.3382 | 2.2265 | 1.0780 | 4.0866 | 2.6851 | 2.5076 | | 1.8642 | H8 | 2.5227 | 2.2181 | 1.0778 | 3.4356 | 2.8090 | 3.0439 | 1.8642 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
107.721 |
|
O1 |
C2 |
H5 |
110.667 |
| O1 |
C2 |
H6 |
111.464 |
|
C2 |
O1 |
H4 |
106.383 |
| C2 |
C3 |
H7 |
120.190 |
|
C2 |
C3 |
H8 |
119.431 |
| C3 |
C2 |
H5 |
110.414 |
|
C3 |
C2 |
H6 |
110.159 |
| H5 |
C2 |
H6 |
106.442 |
|
H7 |
C3 |
H8 |
119.705 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
