return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H12N2 (1,2-Butanediamine)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-268.444613
Energy at 298.15K-268.459563
HF Energy-267.377281
Nuclear repulsion energy264.366925
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3654 3451 8.03      
2 A 3645 3443 2.17      
3 A 3552 3355 4.56      
4 A 3547 3350 0.66      
5 A 3198 3020 26.82      
6 A 3193 3016 19.92      
7 A 3178 3002 2.22      
8 A 3130 2956 31.06      
9 A 3113 2941 11.56      
10 A 3099 2927 17.67      
11 A 3021 2853 98.98      
12 A 3008 2841 30.54      
13 A 1672 1579 38.25      
14 A 1656 1564 45.54      
15 A 1542 1456 6.60      
16 A 1540 1454 4.78      
17 A 1529 1445 3.84      
18 A 1506 1422 1.71      
19 A 1449 1368 1.61      
20 A 1441 1361 9.27      
21 A 1430 1350 1.28      
22 A 1407 1329 11.40      
23 A 1355 1280 1.21      
24 A 1318 1245 1.25      
25 A 1304 1232 1.94      
26 A 1280 1209 1.57      
27 A 1208 1141 5.59      
28 A 1180 1114 6.05      
29 A 1128 1065 3.14      
30 A 1092 1031 4.11      
31 A 1090 1030 2.42      
32 A 1032 974 0.39      
33 A 1001 946 26.61      
34 A 956 902 25.77      
35 A 940 888 53.02      
36 A 886 837 126.26      
37 A 839 793 14.13      
38 A 778 735 0.83      
39 A 625 591 16.55      
40 A 451 426 5.55      
41 A 386 364 3.54      
42 A 342 323 32.54      
43 A 303 286 17.00      
44 A 285 269 47.14      
45 A 245 231 9.73      
46 A 226 214 0.86      
47 A 158 149 3.49      
48 A 85 80 0.98      

Unscaled Zero Point Vibrational Energy (zpe) 37498.6 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 35417.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.18890 0.07456 0.06467

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.921 -0.621 -0.662
H2 1.780 0.166 -1.288
H3 2.919 -0.788 -0.622
C4 -2.388 -0.322 -0.125
H5 -3.031 -1.147 -0.435
H6 -2.746 0.030 0.846
H7 -2.531 0.480 -0.851
N8 0.020 1.474 -0.335
H9 0.532 2.217 0.131
H10 -0.921 1.829 -0.455
C11 -0.929 -0.768 -0.049
H12 -0.854 -1.660 0.580
H13 -0.559 -1.046 -1.037
C14 1.433 -0.215 0.653
H15 2.036 0.575 1.130
H16 1.453 -1.078 1.325
C17 -0.002 0.293 0.537
H18 -0.353 0.527 1.555

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 H7 N8 H9 H10 C11 H12 H13 C14 H15 H16 C17 H18
N11.01511.01314.35244.98484.94784.58972.84713.25723.75802.91903.21352.54421.46052.15732.09242.44383.3777
H21.01511.62924.35405.05865.00534.34362.38992.78793.27903.12143.70972.64612.00802.46512.91242.55363.5724
H31.01311.62925.35065.96375.90955.60043.68833.91114.65073.89074.05523.51282.04052.38902.45453.32384.1450
C44.35244.35405.35061.09111.09301.09173.01123.87802.62451.52722.15362.16753.90024.68454.17392.55082.7715
H54.98485.05865.96371.09111.76311.75254.02324.93273.64792.17042.45602.54544.68785.57534.81713.49153.7328
H64.94785.00535.90951.09301.76311.76863.33654.00542.87402.17722.55083.07954.19054.82124.36902.77372.5446
H74.58974.34365.60041.09171.75251.76862.78563.65512.13662.18343.07242.49994.29574.97824.79892.89033.2450
N82.84712.38993.68833.01124.02323.33652.78561.01561.01362.45113.37972.67862.41302.64863.36441.46822.1467
H93.25722.78793.91113.87804.93274.00543.65511.01561.61443.32834.14203.63302.64552.44033.62392.03812.3811
H103.75803.27904.65072.62453.64792.87402.13661.01361.61442.62883.64022.95523.30873.58174.15422.04692.4615
C112.91903.12143.89071.52722.17042.17722.18342.45113.32832.62881.09411.09062.52503.46152.76691.52562.1406
H123.21353.70974.05522.15362.45602.55083.07243.37974.14203.64021.09411.75472.70623.69452.49302.13102.4466
H132.54422.64613.51282.16752.54543.07952.49992.67863.63302.95521.09061.75472.74103.74893.10272.13973.0388
C141.46052.00802.04053.90024.68784.19054.29572.41302.64553.30872.52502.70622.74101.10231.09411.52652.1343
H152.15732.46512.38904.68455.57534.82124.97822.64862.44033.58173.46153.69453.74891.10231.76382.14112.4272
H162.09242.91242.45454.17394.81714.36904.79893.36443.62394.15422.76692.49303.10271.09411.76382.14872.4275
C172.44382.55363.32382.55083.49152.77372.89031.46822.03812.04691.52562.13102.13971.52652.14112.14871.1022
H183.37773.57244.14502.77153.73282.54463.24502.14672.38112.46152.14062.44663.03882.13432.42722.42751.1022

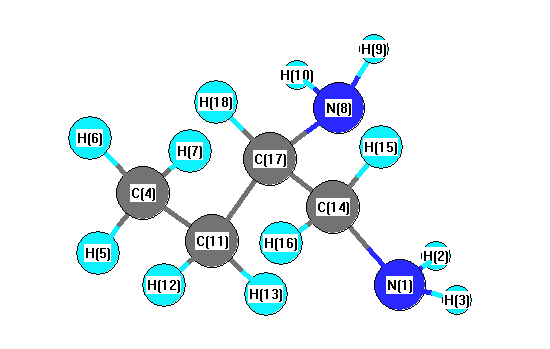
picture of 1,2-Butanediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C14 H15 113.932 N1 C14 H16 109.145
N1 C14 C17 109.776 H2 N1 H3 106.887
H2 N1 C14 107.034 H3 N1 C14 109.839
C4 C11 H12 109.374 C4 C11 H13 110.675
C4 C11 C17 113.344 H5 C4 H6 107.659
H5 C4 H7 106.809 H5 C4 C11 110.875
H6 C4 H7 108.100 H6 C4 C11 111.303
H7 C4 C11 111.882 N8 C17 C11 109.901
N8 C17 C14 107.349 N8 C17 H18 112.485
H9 N8 H10 105.422 H9 N8 C17 108.912
H10 N8 C17 109.771 C11 C17 C14 111.644
C11 C17 H18 108.010 H12 C11 H13 106.866
H12 C11 C17 107.733 H13 C11 C17 108.605
C14 C17 H18 107.471 H15 C14 H16 106.841
H15 C14 C17 107.979 H16 C14 C17 109.039
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability