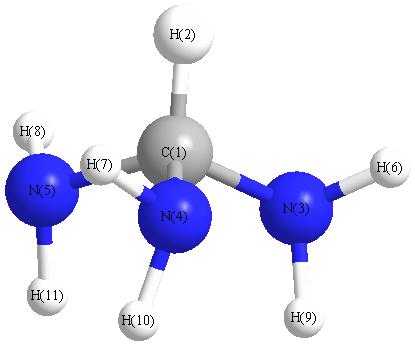
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3606 |
3606 |
0.41 |
|
|
|
| 2 |
A |
3514 |
3514 |
2.30 |
|
|
|
| 3 |
A |
3060 |
3060 |
38.73 |
|
|
|
| 4 |
A |
1651 |
1651 |
47.43 |
|
|
|
| 5 |
A |
1369 |
1369 |
2.22 |
|
|
|
| 6 |
A |
943 |
943 |
12.79 |
|
|
|
| 7 |
A |
878 |
878 |
0.53 |
|
|
|
| 8 |
A |
569 |
569 |
17.06 |
|
|
|
| 9 |
A |
300 |
300 |
61.92 |
|
|
|
| 10 |
E |
3610 |
3610 |
0.63 |
|
|
|
| 10 |
E |
3610 |
3610 |
0.31 |
|
|
|
| 11 |
E |
3519 |
3519 |
0.07 |
|
|
|
| 11 |
E |
3518 |
3518 |
0.19 |
|
|
|
| 12 |
E |
1662 |
1662 |
26.70 |
|
|
|
| 12 |
E |
1660 |
1660 |
26.59 |
|
|
|
| 13 |
E |
1420 |
1420 |
25.39 |
|
|
|
| 13 |
E |
1420 |
1420 |
24.95 |
|
|
|
| 14 |
E |
1237 |
1237 |
54.75 |
|
|
|
| 14 |
E |
1237 |
1237 |
54.55 |
|
|
|
| 15 |
E |
1065 |
1065 |
38.12 |
|
|
|
| 15 |
E |
1064 |
1064 |
37.07 |
|
|
|
| 16 |
E |
911 |
911 |
223.25 |
|
|
|
| 16 |
E |
911 |
911 |
223.15 |
|
|
|
| 17 |
E |
448 |
448 |
35.34 |
|
|
|
| 17 |
E |
446 |
446 |
35.76 |
|
|
|
| 18 |
E |
277 |
277 |
23.43 |
|
|
|
| 18 |
E |
269 |
269 |
20.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22085.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22085.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.084 |
|
|
|
| 2 |
H |
0.121 |
|
|
|
| 3 |
N |
-0.551 |
|
|
|
| 4 |
N |
-0.551 |
|
|
|
| 5 |
N |
-0.551 |
|
|
|
| 6 |
H |
0.255 |
|
|
|
| 7 |
H |
0.255 |
|
|
|
| 8 |
H |
0.255 |
|
|
|
| 9 |
H |
0.228 |
|
|
|
| 10 |
H |
0.228 |
|
|
|
| 11 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.000 |
-0.000 |
-1.646 |
1.646 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.790 |
0.000 |
0.000 |
| y |
0.000 |
-26.790 |
0.000 |
| z |
0.000 |
0.000 |
-21.854 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.469 |
0.000 |
0.000 |
| y |
0.000 |
-2.468 |
0.000 |
| z |
0.000 |
0.000 |
4.936 |
|
| Polar |
|---|
| 3z2-r2 | 9.873 |
|---|
| x2-y2 | -0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.114 |
-0.000 |
-0.000 |
| y |
-0.000 |
5.115 |
-0.001 |
| z |
-0.000 |
-0.001 |
4.984 |
<r2> (average value of r
2) Å
2
| <r2> |
81.519 |
| (<r2>)1/2 |
9.029 |