return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2FCH2CH2F (1,3-difluoropropane)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-316.916759
Energy at 298.15K 
HF Energy-315.984313
Nuclear repulsion energy189.819862
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3185 3008 23.07 43.27 0.74 0.85
2 A 3121 2948 6.12 200.67 0.00 0.01
3 A 3114 2941 0.89 33.52 0.64 0.78
4 A 1547 1461 1.14 12.82 0.72 0.84
5 A 1484 1402 7.34 7.80 0.75 0.85
6 A 1466 1385 9.60 1.56 0.60 0.75
7 A 1333 1259 0.55 20.58 0.71 0.83
8 A 1269 1198 0.53 6.13 0.62 0.76
9 A 1152 1088 28.36 1.28 0.73 0.84
10 A 1041 983 29.78 4.44 0.65 0.79
11 A 904 854 0.00 9.90 0.11 0.20
12 A 548 518 2.88 0.52 0.67 0.80
13 A 252 238 3.47 0.17 0.24 0.39
14 A 86 81 2.75 0.02 0.67 0.80
15 B 3190 3013 25.14 12.13 0.75 0.86
16 B 3174 2998 8.36 84.64 0.75 0.86
17 B 3117 2944 57.02 33.47 0.75 0.86
18 B 1551 1465 4.07 0.01 0.75 0.86
19 B 1436 1356 13.33 1.65 0.75 0.86
20 B 1407 1329 0.84 0.31 0.75 0.86
21 B 1287 1216 8.34 0.63 0.75 0.86
22 B 1146 1083 29.10 0.49 0.75 0.86
23 B 1131 1069 67.52 3.89 0.75 0.86
24 B 987 932 14.82 3.32 0.75 0.86
25 B 789 745 0.88 0.59 0.75 0.86
26 B 416 393 6.19 0.21 0.75 0.86
27 B 206 194 9.96 0.06 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 20168.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 19049.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.29828 0.10421 0.09634

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.003
C2 0.000 1.247 0.143
C3 0.000 -1.247 0.143
F4 1.140 1.251 -0.638
F5 -1.140 -1.251 -0.638
H6 0.884 -0.004 1.645
H7 -0.884 0.004 1.645
H8 -0.869 1.254 -0.518
H9 -0.004 2.153 0.752
H10 0.869 -1.254 -0.518
H11 0.004 -2.153 0.752

Atom - Atom Distances (Å)
  C1 C2 C3 F4 F5 H6 H7 H8 H9 H10 H11
C11.51431.51432.35752.35751.09301.09302.15362.16792.15362.1679
C21.51432.49361.38212.85482.14502.14091.09171.09232.72823.4544
C31.51432.49362.85481.38212.14092.14502.72823.45441.09171.0923
F42.35751.38212.85483.38512.61803.29672.01222.01402.52223.8492
F52.35752.85481.38213.38513.29672.61802.52223.84922.01222.0140
H61.09302.14502.14092.61803.29671.76843.05482.49762.49832.4889
H71.09302.14092.14503.29672.61801.76842.49832.48893.05482.4976
H82.15361.09172.72822.01222.52223.05482.49831.78103.05023.7393
H92.16791.09233.45442.01403.84922.49762.48891.78103.73934.3069
H102.15362.72821.09172.52222.01222.49833.05483.05023.73931.7810
H112.16793.45441.09233.84922.01402.48892.49763.73934.30691.7810

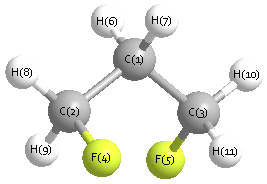
picture of 1,3-difluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 F4 108.878 C1 C2 H8 110.412
C1 C2 H9 111.518 C1 C3 F5 108.878
C1 C3 H10 110.412 C1 C3 H11 111.518
C2 C1 C3 110.836 C2 C1 H6 109.652
C2 C1 H7 109.329 C3 C1 H6 109.329
C3 C1 H7 109.652 F4 C2 H8 108.294
F4 C2 H9 108.393 F5 C3 H10 108.294
F5 C3 H11 108.393 H6 C1 H7 107.993
H8 C2 H9 109.266 H10 C3 H11 109.266
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability