Jump to
S1C2
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -244.650658 |
| Energy at 298.15K | |
| HF Energy | -243.851997 |
| Nuclear repulsion energy | 123.971754 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3881 |
3677 |
83.94 |
|
|
|
| 2 |
A' |
3848 |
3646 |
74.21 |
|
|
|
| 3 |
A' |
3708 |
3513 |
69.41 |
|
|
|
| 4 |
A' |
1891 |
1792 |
474.63 |
|
|
|
| 5 |
A' |
1642 |
1555 |
137.44 |
|
|
|
| 6 |
A' |
1479 |
1401 |
114.80 |
|
|
|
| 7 |
A' |
1268 |
1202 |
201.07 |
|
|
|
| 8 |
A' |
1100 |
1042 |
33.28 |
|
|
|
| 9 |
A' |
986 |
934 |
29.12 |
|
|
|
| 10 |
A' |
595 |
564 |
38.35 |
|
|
|
| 11 |
A' |
494 |
468 |
5.70 |
|
|
|
| 12 |
A" |
811 |
768 |
30.71 |
|
|
|
| 13 |
A" |
607 |
575 |
59.66 |
|
|
|
| 14 |
A" |
497 |
471 |
59.60 |
|
|
|
| 15 |
A" |
273i |
259i |
231.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11266.2 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 10673.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.132 |
0.000 |
| O2 |
-0.048 |
1.337 |
0.000 |
| N3 |
1.127 |
-0.619 |
0.000 |
| O4 |
-1.095 |
-0.666 |
0.000 |
| H5 |
2.011 |
-0.150 |
0.000 |
| H6 |
1.081 |
-1.619 |
0.000 |
| H7 |
-1.841 |
-0.059 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2068 | 1.3543 | 1.3542 | 2.0301 | 2.0577 | 1.8509 |
O2 | 1.2068 | | 2.2824 | 2.2600 | 2.5394 | 3.1648 | 2.2728 | N3 | 1.3543 | 2.2824 | | 2.2221 | 1.0006 | 1.0007 | 3.0205 | O4 | 1.3542 | 2.2600 | 2.2221 | | 3.1477 | 2.3758 | 0.9618 | H5 | 2.0301 | 2.5394 | 1.0006 | 3.1477 | | 1.7384 | 3.8527 | H6 | 2.0577 | 3.1648 | 1.0007 | 2.3758 | 1.7384 | | 3.3128 | H7 | 1.8509 | 2.2728 | 3.0205 | 0.9618 | 3.8527 | 3.3128 | |
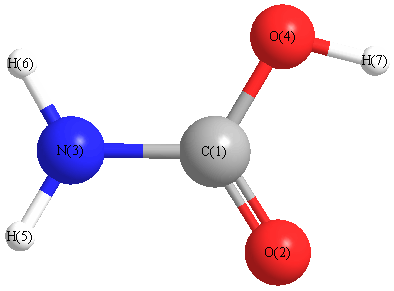 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.334 |
|
C1 |
N3 |
H6 |
121.066 |
| C1 |
O4 |
H7 |
104.834 |
|
O2 |
C1 |
N3 |
125.949 |
| O2 |
C1 |
O4 |
123.792 |
|
N3 |
C1 |
O4 |
110.259 |
| H5 |
N3 |
H6 |
120.600 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -244.650924 |
| Energy at 298.15K | -244.655807 |
| HF Energy | -243.851890 |
| Nuclear repulsion energy | 123.886323 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3879 |
3675 |
84.15 |
|
|
|
| 2 |
A |
3807 |
3607 |
58.77 |
|
|
|
| 3 |
A |
3678 |
3484 |
52.11 |
|
|
|
| 4 |
A |
1894 |
1794 |
447.35 |
|
|
|
| 5 |
A |
1651 |
1564 |
114.56 |
|
|
|
| 6 |
A |
1477 |
1399 |
119.18 |
|
|
|
| 7 |
A |
1274 |
1207 |
168.60 |
|
|
|
| 8 |
A |
1117 |
1058 |
65.26 |
|
|
|
| 9 |
A |
987 |
935 |
26.56 |
|
|
|
| 10 |
A |
809 |
767 |
44.56 |
|
|
|
| 11 |
A |
601 |
569 |
63.86 |
|
|
|
| 12 |
A |
587 |
557 |
63.97 |
|
|
|
| 13 |
A |
509 |
482 |
18.79 |
|
|
|
| 14 |
A |
472 |
447 |
17.57 |
|
|
|
| 15 |
A |
374 |
354 |
262.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11557.1 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 10949.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.044 |
0.124 |
-0.003 |
| O2 |
-0.480 |
1.248 |
0.008 |
| N3 |
1.274 |
-0.221 |
-0.060 |
| O4 |
-0.818 |
-0.985 |
0.003 |
| H5 |
1.924 |
0.512 |
0.158 |
| H6 |
1.529 |
-1.158 |
0.191 |
| H7 |
-1.722 |
-0.658 |
0.007 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2056 | 1.3634 | 1.3529 | 2.0118 | 2.0388 | 1.8517 |
O2 | 1.2056 | | 2.2888 | 2.2586 | 2.5184 | 3.1403 | 2.2746 | N3 | 1.3634 | 2.2888 | | 2.2283 | 1.0034 | 1.0036 | 3.0289 | O4 | 1.3529 | 2.2586 | 2.2283 | | 3.1278 | 2.3612 | 0.9620 | H5 | 2.0118 | 2.5184 | 1.0034 | 3.1278 | | 1.7166 | 3.8321 | H6 | 2.0388 | 3.1403 | 1.0036 | 2.3612 | 1.7166 | | 3.2952 | H7 | 1.8517 | 2.2746 | 3.0289 | 0.9620 | 3.8321 | 3.2952 | |
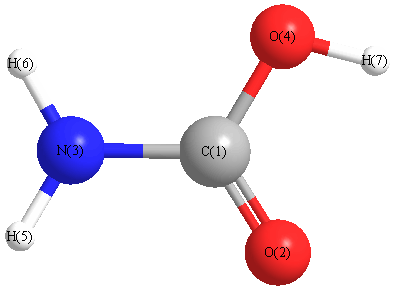 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
115.588 |
|
C1 |
N3 |
H6 |
118.139 |
| C1 |
O4 |
H7 |
104.989 |
|
O2 |
C1 |
N3 |
125.876 |
| O2 |
C1 |
O4 |
123.861 |
|
N3 |
C1 |
O4 |
110.237 |
| H5 |
N3 |
H6 |
117.590 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
