Jump to
S1C2
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -244.642721 |
| Energy at 298.15K | |
| HF Energy | -243.852536 |
| Nuclear repulsion energy | 124.226888 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3903 |
3697 |
88.09 |
|
|
|
| 2 |
A' |
3860 |
3656 |
77.60 |
|
|
|
| 3 |
A' |
3719 |
3523 |
72.82 |
|
|
|
| 4 |
A' |
1923 |
1821 |
511.94 |
|
|
|
| 5 |
A' |
1649 |
1561 |
151.46 |
|
|
|
| 6 |
A' |
1489 |
1410 |
116.59 |
|
|
|
| 7 |
A' |
1277 |
1210 |
207.17 |
|
|
|
| 8 |
A' |
1107 |
1049 |
31.95 |
|
|
|
| 9 |
A' |
997 |
944 |
28.70 |
|
|
|
| 10 |
A' |
603 |
571 |
39.42 |
|
|
|
| 11 |
A' |
498 |
472 |
5.79 |
|
|
|
| 12 |
A" |
821 |
778 |
31.86 |
|
|
|
| 13 |
A" |
609 |
577 |
61.19 |
|
|
|
| 14 |
A" |
501 |
475 |
59.72 |
|
|
|
| 15 |
A" |
257i |
243i |
233.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11350.0 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 10750.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.132 |
0.000 |
| O2 |
-0.045 |
1.333 |
0.000 |
| N3 |
1.124 |
-0.621 |
0.000 |
| O4 |
-1.094 |
-0.661 |
0.000 |
| H5 |
2.009 |
-0.153 |
0.000 |
| H6 |
1.076 |
-1.619 |
0.000 |
| H7 |
-1.841 |
-0.056 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2025 | 1.3528 | 1.3510 | 2.0286 | 2.0556 | 1.8502 |
O2 | 1.2025 | | 2.2772 | 2.2532 | 2.5353 | 3.1587 | 2.2703 | N3 | 1.3528 | 2.2772 | | 2.2189 | 1.0001 | 1.0001 | 3.0183 | O4 | 1.3510 | 2.2532 | 2.2189 | | 3.1439 | 2.3729 | 0.9607 | H5 | 2.0286 | 2.5353 | 1.0001 | 3.1439 | | 1.7374 | 3.8504 | H6 | 2.0556 | 3.1587 | 1.0001 | 2.3729 | 1.7374 | | 3.3097 | H7 | 1.8502 | 2.2703 | 3.0183 | 0.9607 | 3.8504 | 3.3097 | |
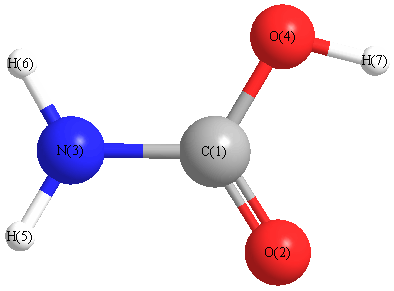 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.356 |
|
C1 |
N3 |
H6 |
121.041 |
| C1 |
O4 |
H7 |
105.072 |
|
O2 |
C1 |
N3 |
125.939 |
| O2 |
C1 |
O4 |
123.762 |
|
N3 |
C1 |
O4 |
110.299 |
| H5 |
N3 |
H6 |
120.604 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -244.642933 |
| Energy at 298.15K | -244.647817 |
| HF Energy | -243.852474 |
| Nuclear repulsion energy | 124.149840 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3901 |
3695 |
88.12 |
|
|
|
| 2 |
A |
3823 |
3621 |
63.23 |
|
|
|
| 3 |
A |
3693 |
3498 |
56.76 |
|
|
|
| 4 |
A |
1926 |
1824 |
484.65 |
|
|
|
| 5 |
A |
1656 |
1569 |
127.87 |
|
|
|
| 6 |
A |
1487 |
1408 |
122.61 |
|
|
|
| 7 |
A |
1282 |
1215 |
178.19 |
|
|
|
| 8 |
A |
1123 |
1063 |
59.75 |
|
|
|
| 9 |
A |
998 |
945 |
26.78 |
|
|
|
| 10 |
A |
820 |
777 |
43.92 |
|
|
|
| 11 |
A |
607 |
575 |
57.83 |
|
|
|
| 12 |
A |
592 |
560 |
70.39 |
|
|
|
| 13 |
A |
513 |
486 |
20.81 |
|
|
|
| 14 |
A |
477 |
452 |
18.25 |
|
|
|
| 15 |
A |
354 |
336 |
264.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11626.2 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 11012.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.044 |
0.124 |
-0.003 |
| O2 |
-0.475 |
1.246 |
0.007 |
| N3 |
1.271 |
-0.224 |
-0.057 |
| O4 |
-0.820 |
-0.980 |
0.003 |
| H5 |
1.926 |
0.507 |
0.149 |
| H6 |
1.526 |
-1.163 |
0.182 |
| H7 |
-1.724 |
-0.653 |
0.007 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2014 | 1.3609 | 1.3500 | 2.0123 | 2.0387 | 1.8511 |
O2 | 1.2014 | | 2.2828 | 2.2520 | 2.5163 | 3.1365 | 2.2722 | N3 | 1.3609 | 2.2828 | | 2.2246 | 1.0026 | 1.0027 | 3.0260 | O4 | 1.3500 | 2.2520 | 2.2246 | | 3.1264 | 2.3604 | 0.9608 | H5 | 2.0123 | 2.5163 | 1.0026 | 3.1264 | | 1.7176 | 3.8322 | H6 | 2.0387 | 3.1365 | 1.0027 | 2.3604 | 1.7176 | | 3.2944 | H7 | 1.8511 | 2.2722 | 3.0260 | 0.9608 | 3.8322 | 3.2944 | |
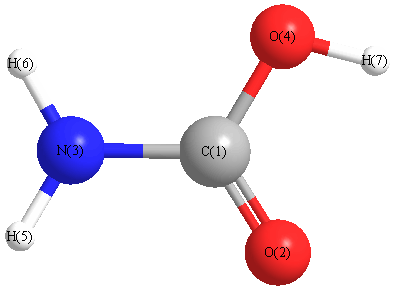 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
115.897 |
|
C1 |
N3 |
H6 |
118.417 |
| C1 |
O4 |
H7 |
105.222 |
|
O2 |
C1 |
N3 |
125.860 |
| O2 |
C1 |
O4 |
123.827 |
|
N3 |
C1 |
O4 |
110.290 |
| H5 |
N3 |
H6 |
117.861 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
