return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H2O2 (Propiolic acid)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-265.330098
Energy at 298.15K-265.331734
HF Energy-264.457128
Nuclear repulsion energy144.466003
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3781 3572 100.05      
2 A' 3529 3334 65.14      
3 A' 2177 2056 65.40      
4 A' 1820 1719 251.24      
5 A' 1388 1311 100.21      
6 A' 1193 1127 335.61      
7 A' 833 786 27.79      
8 A' 660 624 33.44      
9 A' 592 559 4.51      
10 A' 529 500 25.97      
11 A' 188 178 4.20      
12 A" 785 741 55.54      
13 A" 655 619 6.72      
14 A" 627 592 98.66      
15 A" 248 234 5.00      

Unscaled Zero Point Vibrational Energy (zpe) 9503.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 8976.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.40273 0.13808 0.10282

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.493 0.000
C2 -0.223 -0.935 0.000
C3 -0.478 -2.122 0.000
O4 1.316 0.775 0.000
O5 -0.875 1.325 0.000
H6 -0.691 -3.163 0.000
H7 1.368 1.743 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7
C11.44602.65901.34601.20753.72131.8529
C21.44601.21402.30092.35292.27653.1151
C32.65901.21403.40803.47021.06254.2834
O41.34602.30093.40802.25964.42030.9691
O51.20752.35293.47022.25964.49212.2822
H63.72132.27651.06254.42034.49215.3207
H71.85293.11514.28340.96912.28225.3207

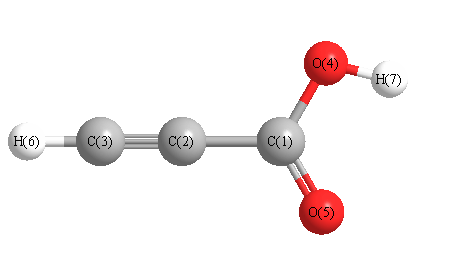
picture of Propiolic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 176.709 C1 O4 H7 105.157
C2 C1 O4 110.939 C2 C1 O5 124.678
C2 C3 H6 179.407 O4 C1 O5 124.384
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability