Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -357.727842 |
| Energy at 298.15K | -357.732877 |
| HF Energy | -356.583380 |
| Nuclear repulsion energy | 232.785847 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3808 |
3597 |
100.69 |
|
|
|
| 2 |
A' |
3781 |
3571 |
90.95 |
|
|
|
| 3 |
A' |
3655 |
3453 |
83.24 |
|
|
|
| 4 |
A' |
1839 |
1737 |
142.15 |
|
|
|
| 5 |
A' |
1819 |
1718 |
324.34 |
|
|
|
| 6 |
A' |
1603 |
1514 |
112.09 |
|
|
|
| 7 |
A' |
1456 |
1376 |
23.18 |
|
|
|
| 8 |
A' |
1336 |
1261 |
56.02 |
|
|
|
| 9 |
A' |
1197 |
1130 |
270.17 |
|
|
|
| 10 |
A' |
1098 |
1037 |
2.81 |
|
|
|
| 11 |
A' |
792 |
748 |
7.28 |
|
|
|
| 12 |
A' |
609 |
575 |
69.96 |
|
|
|
| 13 |
A' |
533 |
504 |
0.13 |
|
|
|
| 14 |
A' |
419 |
395 |
4.08 |
|
|
|
| 15 |
A' |
272 |
257 |
14.08 |
|
|
|
| 16 |
A" |
844 |
797 |
4.73 |
|
|
|
| 17 |
A" |
695 |
656 |
118.92 |
|
|
|
| 18 |
A" |
652 |
616 |
17.05 |
|
|
|
| 19 |
A" |
441 |
417 |
4.25 |
|
|
|
| 20 |
A" |
293 |
277 |
210.44 |
|
|
|
| 21 |
A" |
70 |
66 |
1.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13606.1 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12850.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.749 |
0.000 |
| C2 |
-0.054 |
-0.785 |
0.000 |
| O3 |
-1.095 |
-1.411 |
0.000 |
| O4 |
1.034 |
1.382 |
0.000 |
| O5 |
-1.223 |
1.277 |
0.000 |
| N6 |
1.198 |
-1.294 |
0.000 |
| H7 |
1.325 |
-2.290 |
0.000 |
| H8 |
1.984 |
-0.668 |
0.000 |
| H9 |
-1.092 |
2.237 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5344 | 2.4212 | 1.2121 | 1.3324 | 2.3678 | 3.3152 | 2.4377 | 1.8460 |
C2 | 1.5344 | | 1.2148 | 2.4243 | 2.3702 | 1.3511 | 2.0413 | 2.0411 | 3.1952 | O3 | 2.4212 | 1.2148 | | 3.5115 | 2.6907 | 2.2955 | 2.5744 | 3.1672 | 3.6479 | O4 | 1.2121 | 2.4243 | 3.5115 | | 2.2592 | 2.6810 | 3.6839 | 2.2595 | 2.2911 | O5 | 1.3324 | 2.3702 | 2.6907 | 2.2592 | | 3.5312 | 4.3836 | 3.7507 | 0.9692 | N6 | 2.3678 | 1.3511 | 2.2955 | 2.6810 | 3.5312 | | 1.0046 | 1.0050 | 4.2084 | H7 | 3.3152 | 2.0413 | 2.5744 | 3.6839 | 4.3836 | 1.0046 | | 1.7513 | 5.1321 | H8 | 2.4377 | 2.0411 | 3.1672 | 2.2595 | 3.7507 | 1.0050 | 1.7513 | | 4.2309 | H9 | 1.8460 | 3.1952 | 3.6479 | 2.2911 | 0.9692 | 4.2084 | 5.1321 | 4.2309 | |
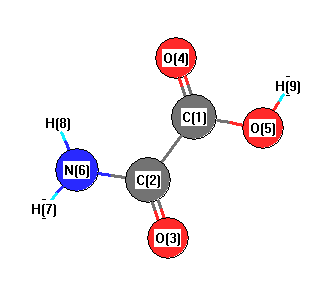 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
123.031 |
|
C1 |
C2 |
N6 |
110.126 |
| C1 |
O5 |
H9 |
105.571 |
|
C2 |
C1 |
O4 |
123.511 |
| C2 |
C1 |
O5 |
111.342 |
|
C2 |
N6 |
H7 |
119.397 |
| C2 |
N6 |
H8 |
119.339 |
|
O3 |
C2 |
N6 |
126.843 |
| O4 |
C1 |
O5 |
125.147 |
|
H7 |
N6 |
H8 |
121.264 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -357.734955 |
| Energy at 298.15K | -357.740282 |
| Nuclear repulsion energy | 234.345068 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3796 |
3585 |
108.87 |
|
|
|
| 2 |
A' |
3644 |
3442 |
103.99 |
|
|
|
| 3 |
A' |
3615 |
3415 |
144.10 |
|
|
|
| 4 |
A' |
1866 |
1762 |
209.93 |
|
|
|
| 5 |
A' |
1824 |
1723 |
222.77 |
|
|
|
| 6 |
A' |
1606 |
1517 |
67.32 |
|
|
|
| 7 |
A' |
1474 |
1392 |
217.44 |
|
|
|
| 8 |
A' |
1366 |
1290 |
324.42 |
|
|
|
| 9 |
A' |
1235 |
1166 |
6.57 |
|
|
|
| 10 |
A' |
1105 |
1043 |
3.38 |
|
|
|
| 11 |
A' |
818 |
772 |
10.11 |
|
|
|
| 12 |
A' |
631 |
596 |
11.64 |
|
|
|
| 13 |
A' |
551 |
520 |
1.97 |
|
|
|
| 14 |
A' |
412 |
390 |
7.76 |
|
|
|
| 15 |
A' |
282 |
266 |
41.10 |
|
|
|
| 16 |
A" |
840 |
793 |
0.40 |
|
|
|
| 17 |
A" |
779 |
736 |
92.23 |
|
|
|
| 18 |
A" |
683 |
645 |
2.44 |
|
|
|
| 19 |
A" |
472 |
446 |
82.47 |
|
|
|
| 20 |
A" |
365 |
345 |
152.31 |
|
|
|
| 21 |
A" |
121 |
115 |
3.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13741.6 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12979.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.006 |
-0.790 |
0.000 |
| C2 |
0.000 |
0.745 |
0.000 |
| O3 |
-1.068 |
1.346 |
0.000 |
| O4 |
1.016 |
-1.451 |
0.000 |
| O5 |
-1.230 |
-1.271 |
0.000 |
| N6 |
1.233 |
1.265 |
0.000 |
| H7 |
1.367 |
2.261 |
0.000 |
| H8 |
2.015 |
0.631 |
0.000 |
| H9 |
-1.796 |
-0.473 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5346 | 2.3904 | 1.2075 | 1.3270 | 2.3936 | 3.3409 | 2.4607 | 1.8300 |
C2 | 1.5346 | | 1.2254 | 2.4199 | 2.3621 | 1.3387 | 2.0419 | 2.0184 | 2.1707 | O3 | 2.3904 | 1.2254 | | 3.4882 | 2.6222 | 2.3028 | 2.6019 | 3.1649 | 1.9598 | O4 | 1.2075 | 2.4199 | 3.4882 | | 2.2539 | 2.7251 | 3.7290 | 2.3095 | 2.9777 | O5 | 1.3270 | 2.3621 | 2.6222 | 2.2539 | | 3.5362 | 4.3850 | 3.7621 | 0.9781 | N6 | 2.3936 | 1.3387 | 2.3028 | 2.7251 | 3.5362 | | 1.0049 | 1.0066 | 3.4933 | H7 | 3.3409 | 2.0419 | 2.6019 | 3.7290 | 4.3850 | 1.0049 | | 1.7540 | 4.1820 | H8 | 2.4607 | 2.0184 | 3.1649 | 2.3095 | 3.7621 | 1.0066 | 1.7540 | | 3.9684 | H9 | 1.8300 | 2.1707 | 1.9598 | 2.9777 | 0.9781 | 3.4933 | 4.1820 | 3.9684 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.591 |
|
C1 |
C2 |
N6 |
112.645 |
| C1 |
O5 |
H9 |
104.072 |
|
C2 |
C1 |
O4 |
123.447 |
| C2 |
C1 |
O5 |
111.052 |
|
C2 |
N6 |
H7 |
120.543 |
| C2 |
N6 |
H8 |
118.078 |
|
O3 |
C2 |
N6 |
127.764 |
| O4 |
C1 |
O5 |
125.501 |
|
H7 |
N6 |
H8 |
121.379 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
