Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -132.936914 |
| Energy at 298.15K | -132.941149 |
| HF Energy | -132.481390 |
| Nuclear repulsion energy | 65.382433 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3549 |
3352 |
0.86 |
|
|
|
| 2 |
A' |
3366 |
3180 |
0.44 |
|
|
|
| 3 |
A' |
3242 |
3062 |
2.39 |
|
|
|
| 4 |
A' |
3136 |
2962 |
33.92 |
|
|
|
| 5 |
A' |
1525 |
1440 |
14.63 |
|
|
|
| 6 |
A' |
1475 |
1393 |
16.40 |
|
|
|
| 7 |
A' |
1399 |
1321 |
15.42 |
|
|
|
| 8 |
A' |
1254 |
1185 |
30.34 |
|
|
|
| 9 |
A' |
1058 |
1000 |
8.17 |
|
|
|
| 10 |
A' |
1017 |
960 |
10.04 |
|
|
|
| 11 |
A' |
492 |
465 |
14.52 |
|
|
|
| 12 |
A" |
1156 |
1092 |
1.23 |
|
|
|
| 13 |
A" |
844 |
797 |
2.48 |
|
|
|
| 14 |
A" |
741 |
700 |
104.63 |
|
|
|
| 15 |
A" |
541 |
511 |
1.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12398.1 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 11710.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| C2 |
1.137 |
-0.402 |
0.000 |
| N3 |
-1.179 |
-0.107 |
0.000 |
| H4 |
0.168 |
1.496 |
0.000 |
| H5 |
2.134 |
0.014 |
0.000 |
| H6 |
1.009 |
-1.474 |
0.000 |
| H7 |
-1.883 |
0.629 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4010 | 1.2898 | 1.0932 | 2.1712 | 2.1427 | 1.8953 |
C2 | 1.4010 | | 2.3349 | 2.1316 | 1.0793 | 1.0801 | 3.1917 | N3 | 1.2898 | 2.3349 | | 2.0940 | 3.3145 | 2.5795 | 1.0188 | H4 | 1.0932 | 2.1316 | 2.0940 | | 2.4622 | 3.0874 | 2.2272 | H5 | 2.1712 | 1.0793 | 3.3145 | 2.4622 | | 1.8652 | 4.0637 | H6 | 2.1427 | 1.0801 | 2.5795 | 3.0874 | 1.8652 | | 3.5758 | H7 | 1.8953 | 3.1917 | 1.0188 | 2.2272 | 4.0637 | 3.5758 | |
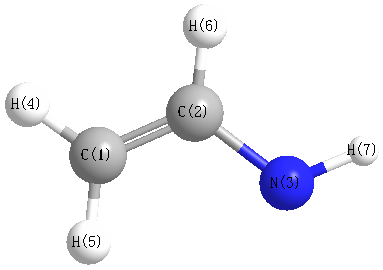 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.634 |
|
C1 |
C2 |
H6 |
118.878 |
| C1 |
N3 |
H7 |
109.816 |
|
C2 |
C1 |
N3 |
120.337 |
| C2 |
C1 |
H4 |
116.894 |
|
N3 |
C1 |
H4 |
122.769 |
| H5 |
C2 |
H6 |
119.488 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -132.936603 |
| Energy at 298.15K | -132.940835 |
| HF Energy | -132.481014 |
| Nuclear repulsion energy | 65.375461 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3500 |
3306 |
1.50 |
|
|
|
| 2 |
A' |
3347 |
3161 |
1.81 |
|
|
|
| 3 |
A' |
3228 |
3049 |
1.52 |
|
|
|
| 4 |
A' |
3196 |
3018 |
15.66 |
|
|
|
| 5 |
A' |
1526 |
1441 |
8.10 |
|
|
|
| 6 |
A' |
1465 |
1384 |
0.60 |
|
|
|
| 7 |
A' |
1411 |
1332 |
22.23 |
|
|
|
| 8 |
A' |
1243 |
1174 |
44.25 |
|
|
|
| 9 |
A' |
1088 |
1028 |
32.85 |
|
|
|
| 10 |
A' |
1005 |
949 |
0.04 |
|
|
|
| 11 |
A' |
501 |
473 |
8.78 |
|
|
|
| 12 |
A" |
1182 |
1116 |
40.29 |
|
|
|
| 13 |
A" |
867 |
819 |
71.57 |
|
|
|
| 14 |
A" |
743 |
701 |
0.34 |
|
|
|
| 15 |
A" |
511 |
483 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12405.2 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 11716.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.447 |
0.000 |
| C2 |
1.114 |
-0.404 |
0.000 |
| N3 |
-1.241 |
0.092 |
0.000 |
| H4 |
0.194 |
1.519 |
0.000 |
| H5 |
2.121 |
-0.014 |
0.000 |
| H6 |
0.983 |
-1.478 |
0.000 |
| H7 |
-1.297 |
-0.929 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4015 | 1.2905 | 1.0892 | 2.1705 | 2.1607 | 1.8910 |
C2 | 1.4015 | | 2.4062 | 2.1307 | 1.0800 | 1.0819 | 2.4675 | N3 | 1.2905 | 2.4062 | | 2.0238 | 3.3635 | 2.7213 | 1.0225 | H4 | 1.0892 | 2.1307 | 2.0238 | | 2.4616 | 3.0980 | 2.8663 | H5 | 2.1705 | 1.0800 | 3.3635 | 2.4616 | | 1.8545 | 3.5385 | H6 | 2.1607 | 1.0819 | 2.7213 | 3.0980 | 1.8545 | | 2.3445 | H7 | 1.8910 | 2.4675 | 1.0225 | 2.8663 | 3.5385 | 2.3445 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.474 |
|
C1 |
C2 |
H6 |
120.385 |
| C1 |
N3 |
H7 |
109.126 |
|
C2 |
C1 |
N3 |
126.665 |
| C2 |
C1 |
H4 |
117.075 |
|
N3 |
C1 |
H4 |
116.260 |
| H5 |
C2 |
H6 |
118.141 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
