Jump to
S1C2
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -132.970507 |
| Energy at 298.15K | -132.974743 |
| HF Energy | -132.481837 |
| Nuclear repulsion energy | 65.195876 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3536 |
3349 |
0.13 |
|
|
|
| 2 |
A' |
3340 |
3163 |
1.37 |
|
|
|
| 3 |
A' |
3225 |
3055 |
2.62 |
|
|
|
| 4 |
A' |
3138 |
2972 |
30.93 |
|
|
|
| 5 |
A' |
1530 |
1450 |
16.18 |
|
|
|
| 6 |
A' |
1477 |
1399 |
15.77 |
|
|
|
| 7 |
A' |
1389 |
1316 |
14.80 |
|
|
|
| 8 |
A' |
1245 |
1179 |
28.20 |
|
|
|
| 9 |
A' |
1077 |
1020 |
4.51 |
|
|
|
| 10 |
A' |
1020 |
966 |
9.43 |
|
|
|
| 11 |
A' |
492 |
466 |
13.13 |
|
|
|
| 12 |
A" |
1123 |
1064 |
0.45 |
|
|
|
| 13 |
A" |
843 |
798 |
7.05 |
|
|
|
| 14 |
A" |
729 |
691 |
99.62 |
|
|
|
| 15 |
A" |
553 |
524 |
3.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12357.7 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 11705.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| C2 |
1.144 |
-0.387 |
0.000 |
| N3 |
-1.184 |
-0.123 |
0.000 |
| H4 |
0.147 |
1.499 |
0.000 |
| H5 |
2.136 |
0.042 |
0.000 |
| H6 |
1.035 |
-1.463 |
0.000 |
| H7 |
-1.894 |
0.609 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3981 | 1.3011 | 1.0924 | 2.1684 | 2.1452 | 1.9037 |
C2 | 1.3981 | | 2.3435 | 2.1330 | 1.0806 | 1.0812 | 3.1972 | N3 | 1.3011 | 2.3435 | | 2.0981 | 3.3243 | 2.5925 | 1.0191 | H4 | 1.0924 | 2.1330 | 2.0981 | | 2.4648 | 3.0916 | 2.2269 | H5 | 2.1684 | 1.0806 | 3.3243 | 2.4648 | | 1.8647 | 4.0695 | H6 | 2.1452 | 1.0812 | 2.5925 | 3.0916 | 1.8647 | | 3.5875 | H7 | 1.9037 | 3.1972 | 1.0191 | 2.2269 | 4.0695 | 3.5875 | |
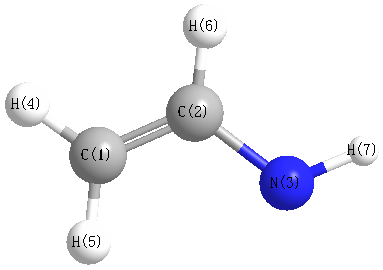 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.518 |
|
C1 |
C2 |
H6 |
119.271 |
| C1 |
N3 |
H7 |
109.673 |
|
C2 |
C1 |
N3 |
120.465 |
| C2 |
C1 |
H4 |
117.314 |
|
N3 |
C1 |
H4 |
122.222 |
| H5 |
C2 |
H6 |
119.211 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -132.970156 |
| Energy at 298.15K | -132.974395 |
| HF Energy | -132.481431 |
| Nuclear repulsion energy | 65.204915 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3491 |
3307 |
2.36 |
|
|
|
| 2 |
A' |
3322 |
3146 |
3.30 |
|
|
|
| 3 |
A' |
3212 |
3043 |
1.66 |
|
|
|
| 4 |
A' |
3190 |
3022 |
15.87 |
|
|
|
| 5 |
A' |
1528 |
1447 |
6.67 |
|
|
|
| 6 |
A' |
1473 |
1395 |
1.21 |
|
|
|
| 7 |
A' |
1397 |
1323 |
30.89 |
|
|
|
| 8 |
A' |
1234 |
1169 |
32.77 |
|
|
|
| 9 |
A' |
1104 |
1046 |
29.23 |
|
|
|
| 10 |
A' |
1011 |
957 |
0.28 |
|
|
|
| 11 |
A' |
503 |
476 |
8.85 |
|
|
|
| 12 |
A" |
1149 |
1088 |
41.15 |
|
|
|
| 13 |
A" |
856 |
811 |
68.38 |
|
|
|
| 14 |
A" |
738 |
699 |
2.96 |
|
|
|
| 15 |
A" |
528 |
500 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12367.3 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 11714.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.449 |
0.000 |
| C2 |
1.119 |
-0.390 |
0.000 |
| N3 |
-1.247 |
0.075 |
0.000 |
| H4 |
0.177 |
1.523 |
0.000 |
| H5 |
2.123 |
0.012 |
0.000 |
| H6 |
1.003 |
-1.466 |
0.000 |
| H7 |
-1.289 |
-0.947 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3984 | 1.3020 | 1.0889 | 2.1677 | 2.1618 | 1.9002 |
C2 | 1.3984 | | 2.4114 | 2.1322 | 1.0811 | 1.0831 | 2.4721 | N3 | 1.3020 | 2.4114 | | 2.0315 | 3.3708 | 2.7268 | 1.0225 | H4 | 1.0889 | 2.1322 | 2.0315 | | 2.4641 | 3.1014 | 2.8727 | H5 | 2.1677 | 1.0811 | 3.3708 | 2.4641 | | 1.8548 | 3.5445 | H6 | 2.1618 | 1.0831 | 2.7268 | 3.1014 | 1.8548 | | 2.3499 | H7 | 1.9002 | 2.4721 | 1.0225 | 2.8727 | 3.5445 | 2.3499 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.376 |
|
C1 |
C2 |
H6 |
120.657 |
| C1 |
N3 |
H7 |
109.070 |
|
C2 |
C1 |
N3 |
126.455 |
| C2 |
C1 |
H4 |
117.475 |
|
N3 |
C1 |
H4 |
116.069 |
| H5 |
C2 |
H6 |
117.967 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
