return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CCH (phenylacetylene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-307.597239
Energy at 298.15K-307.601902
HF Energy-306.405727
Nuclear repulsion energy298.860238
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3534 3337 87.10      
2 A 3258 3077 5.02      
3 A 3245 3065 4.42      
4 A 3229 3050 0.37      
5 A 2154 2034 0.41      
6 A 1643 1552 3.30      
7 A 1520 1435 17.15      
8 A 1230 1162 1.21      
9 A 1205 1138 0.03      
10 A 1052 994 3.25      
11 A 1010 954 0.66      
12 A 770 728 1.53      
13 A 464 438 0.40      
14 A 948 895 0.00      
15 A 862 814 0.00      
16 A 404 381 0.00      
17 A 972 918 0.08      
18 A 920 869 2.98      
19 A 766 724 61.22      
20 A 683 645 5.52      
21 A 567 535 12.76      
22 A 502 474 26.50      
23 A 349 329 0.30      
24 A 137 130 1.10      
25 A 3251 3071 9.28      
26 A 3236 3057 0.89      
27 A 1616 1527 1.91      
28 A 1479 1397 4.73      
29 A 1446 1365 1.06      
30 A 1331 1257 0.09      
31 A 1188 1122 0.00      
32 A 1102 1041 3.80      
33 A 623 588 0.39      
34 A 596 563 39.10      
35 A 500 472 0.01      
36 A 144 136 0.60      

Unscaled Zero Point Vibrational Energy (zpe) 23967.5 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 22637.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.18999 0.05105 0.04024

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.589
C2 0.000 0.000 2.015
C3 0.000 1.211 -0.118
C4 0.000 -1.211 -0.118
C5 0.000 1.207 -1.510
C6 0.000 -1.207 -1.510
C7 0.000 0.000 -2.209
C8 0.000 0.000 3.233
H9 0.000 0.000 4.294
H10 0.000 2.145 0.432
H11 0.000 -2.145 0.432
H12 0.000 2.147 -2.049
H13 0.000 -2.147 -2.049
H14 0.000 0.000 -3.293

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.42561.40281.40282.42152.42152.79862.64313.70472.15062.15063.40163.40163.8824
C21.42562.45322.45323.72583.72584.22421.21752.27912.66592.66594.59634.59635.3080
C31.40282.45322.42231.39152.78992.41633.56304.57571.08373.40082.14553.87373.3978
C41.40282.45322.42232.78991.39152.41633.56304.57573.40081.08373.87372.14553.3978
C52.42153.72581.39152.78992.41381.39494.89365.92812.15633.87351.08393.39722.1532
C62.42153.72582.78991.39152.41381.39494.89365.92813.87352.15633.39721.08392.1532
C72.79864.22422.41632.41631.39491.39495.44176.50333.40233.40232.15322.15321.0838
C82.64311.21753.56303.56304.89364.89365.44171.06163.52763.52765.70125.70126.5255
H93.70472.27914.57574.57575.92815.92816.50331.06164.41794.41796.69666.69667.5871
H102.15062.66591.08373.40082.15633.87353.40233.52764.41794.28972.48074.95744.2982
H112.15062.66593.40081.08373.87352.15633.40233.52764.41794.28974.95742.48074.2982
H123.40164.59632.14553.87371.08393.39722.15325.70126.69662.48074.95744.29452.4816
H133.40164.59633.87372.14553.39721.08392.15325.70126.69664.95742.48074.29452.4816
H143.88245.30803.39783.39782.15322.15321.08386.52557.58714.29824.29822.48162.4816

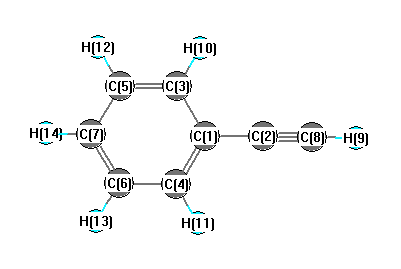
picture of phenylacetylene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C8 180.000 C1 C3 C5 120.126
C1 C3 H10 119.192 C1 C4 C6 120.126
C1 C4 H11 119.192 C2 C1 C3 120.300
C2 C1 C4 120.300 C2 C8 H9 180.000
C3 C1 C4 119.400 C3 C5 C7 120.264
C3 C5 H12 119.648 C4 C6 C7 120.264
C4 C6 H13 119.648 C5 C3 H10 120.682
C5 C7 C6 119.819 C5 C7 H14 120.090
C6 C4 H11 120.682 C6 C7 H14 120.090
C7 C5 H12 120.088 C7 C6 H13 120.088
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability