Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -322.897598 |
| Energy at 298.15K | |
| HF Energy | -321.770419 |
| Nuclear repulsion energy | 234.869913 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3207 |
3029 |
14.50 |
|
|
|
| 2 |
A' |
3124 |
2950 |
19.88 |
|
|
|
| 3 |
A' |
3108 |
2936 |
14.60 |
|
|
|
| 4 |
A' |
3100 |
2928 |
15.97 |
|
|
|
| 5 |
A' |
1639 |
1548 |
109.89 |
|
|
|
| 6 |
A' |
1547 |
1461 |
16.03 |
|
|
|
| 7 |
A' |
1536 |
1450 |
1.38 |
|
|
|
| 8 |
A' |
1521 |
1437 |
1.47 |
|
|
|
| 9 |
A' |
1435 |
1356 |
7.43 |
|
|
|
| 10 |
A' |
1429 |
1350 |
17.51 |
|
|
|
| 11 |
A' |
1341 |
1266 |
8.50 |
|
|
|
| 12 |
A' |
1178 |
1113 |
5.95 |
|
|
|
| 13 |
A' |
1104 |
1043 |
31.87 |
|
|
|
| 14 |
A' |
1073 |
1013 |
8.11 |
|
|
|
| 15 |
A' |
945 |
893 |
128.43 |
|
|
|
| 16 |
A' |
874 |
826 |
459.08 |
|
|
|
| 17 |
A' |
679 |
641 |
40.29 |
|
|
|
| 18 |
A' |
397 |
375 |
0.02 |
|
|
|
| 19 |
A' |
354 |
334 |
1.99 |
|
|
|
| 20 |
A' |
155 |
147 |
0.31 |
|
|
|
| 21 |
A" |
3203 |
3025 |
31.31 |
|
|
|
| 22 |
A" |
3181 |
3004 |
6.39 |
|
|
|
| 23 |
A" |
3159 |
2984 |
6.60 |
|
|
|
| 24 |
A" |
1533 |
1448 |
7.05 |
|
|
|
| 25 |
A" |
1334 |
1260 |
0.00 |
|
|
|
| 26 |
A" |
1284 |
1213 |
0.33 |
|
|
|
| 27 |
A" |
1203 |
1136 |
0.54 |
|
|
|
| 28 |
A" |
905 |
855 |
1.19 |
|
|
|
| 29 |
A" |
770 |
728 |
2.28 |
|
|
|
| 30 |
A" |
243 |
230 |
0.03 |
|
|
|
| 31 |
A" |
217 |
205 |
0.87 |
|
|
|
| 32 |
A" |
106 |
100 |
1.10 |
|
|
|
| 33 |
A" |
48i |
45i |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23417.7 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 22118.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.914 |
2.192 |
0.000 |
| C2 |
-1.507 |
0.721 |
0.000 |
| C3 |
0.000 |
0.574 |
0.000 |
| O4 |
0.282 |
-0.836 |
0.000 |
| N5 |
1.669 |
-1.012 |
0.000 |
| O6 |
1.947 |
-2.168 |
0.000 |
| H7 |
-2.999 |
2.296 |
0.000 |
| H8 |
-1.530 |
2.707 |
0.882 |
| H9 |
-1.530 |
2.707 |
-0.882 |
| H10 |
-1.910 |
0.212 |
-0.879 |
| H11 |
-1.910 |
0.212 |
0.879 |
| H12 |
0.442 |
1.034 |
0.888 |
| H13 |
0.442 |
1.034 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5266 | 2.5066 | 3.7412 | 4.8066 | 5.8245 | 1.0900 | 1.0916 | 1.0916 | 2.1663 | 2.1663 | 2.7710 | 2.7710 |
C2 | 1.5266 | | 1.5141 | 2.3721 | 3.6178 | 4.5034 | 2.1695 | 2.1736 | 2.1736 | 1.0923 | 1.0923 | 2.1641 | 2.1641 | C3 | 2.5066 | 1.5141 | | 1.4380 | 2.3020 | 3.3632 | 3.4584 | 2.7695 | 2.7695 | 2.1331 | 2.1331 | 1.0933 | 1.0933 | O4 | 3.7412 | 2.3721 | 1.4380 | | 1.3975 | 2.1322 | 4.5364 | 4.0766 | 4.0766 | 2.5840 | 2.5840 | 2.0767 | 2.0767 | N5 | 4.8066 | 3.6178 | 2.3020 | 1.3975 | | 1.1895 | 5.7211 | 4.9841 | 4.9841 | 3.8829 | 3.8829 | 2.5458 | 2.5458 | O6 | 5.8245 | 4.5034 | 3.3632 | 2.1322 | 1.1895 | | 6.6631 | 6.0530 | 6.0530 | 4.6170 | 4.6170 | 3.6485 | 3.6485 | H7 | 1.0900 | 2.1695 | 3.4584 | 4.5364 | 5.7211 | 6.6631 | | 1.7627 | 1.7627 | 2.5100 | 2.5100 | 3.7708 | 3.7708 | H8 | 1.0916 | 2.1736 | 2.7695 | 4.0766 | 4.9841 | 6.0530 | 1.7627 | | 1.7648 | 3.0775 | 2.5239 | 2.5855 | 3.1335 | H9 | 1.0916 | 2.1736 | 2.7695 | 4.0766 | 4.9841 | 6.0530 | 1.7627 | 1.7648 | | 2.5239 | 3.0775 | 3.1335 | 2.5855 | H10 | 2.1663 | 1.0923 | 2.1331 | 2.5840 | 3.8829 | 4.6170 | 2.5100 | 3.0775 | 2.5239 | | 1.7570 | 3.0538 | 2.4912 | H11 | 2.1663 | 1.0923 | 2.1331 | 2.5840 | 3.8829 | 4.6170 | 2.5100 | 2.5239 | 3.0775 | 1.7570 | | 2.4912 | 3.0538 | H12 | 2.7710 | 2.1641 | 1.0933 | 2.0767 | 2.5458 | 3.6485 | 3.7708 | 2.5855 | 3.1335 | 3.0538 | 2.4912 | | 1.7756 | H13 | 2.7710 | 2.1641 | 1.0933 | 2.0767 | 2.5458 | 3.6485 | 3.7708 | 3.1335 | 2.5855 | 2.4912 | 3.0538 | 1.7756 | |
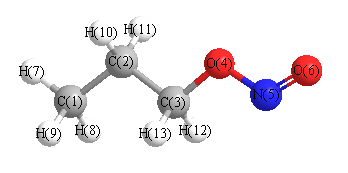 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.046 |
|
C1 |
C2 |
H10 |
110.526 |
| C1 |
C2 |
H11 |
110.526 |
|
C2 |
C1 |
H7 |
110.922 |
| C2 |
C1 |
H8 |
111.149 |
|
C2 |
C1 |
H9 |
111.149 |
| C2 |
C3 |
O4 |
106.907 |
|
C2 |
C3 |
H12 |
111.168 |
| C2 |
C3 |
H13 |
111.168 |
|
C3 |
C2 |
H10 |
108.775 |
| C3 |
C2 |
H11 |
108.775 |
|
C3 |
O4 |
N5 |
108.545 |
| O4 |
C3 |
H12 |
109.494 |
|
O4 |
C3 |
H13 |
109.494 |
| O4 |
N5 |
O6 |
110.763 |
|
H7 |
C1 |
H8 |
107.797 |
| H7 |
C1 |
H9 |
107.797 |
|
H8 |
C1 |
H9 |
107.873 |
| H10 |
C2 |
H11 |
107.084 |
|
H12 |
C3 |
H13 |
108.584 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -322.900271 |
| Energy at 298.15K | -322.909386 |
| HF Energy | -321.769740 |
| Nuclear repulsion energy | 242.969970 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3212 |
3034 |
14.62 |
|
|
|
| 2 |
A |
3202 |
3024 |
18.65 |
|
|
|
| 3 |
A |
3179 |
3003 |
13.22 |
|
|
|
| 4 |
A |
3164 |
2988 |
7.74 |
|
|
|
| 5 |
A |
3112 |
2939 |
35.52 |
|
|
|
| 6 |
A |
3110 |
2937 |
5.55 |
|
|
|
| 7 |
A |
3107 |
2935 |
8.34 |
|
|
|
| 8 |
A |
1645 |
1554 |
130.18 |
|
|
|
| 9 |
A |
1538 |
1453 |
8.20 |
|
|
|
| 10 |
A |
1530 |
1445 |
6.79 |
|
|
|
| 11 |
A |
1513 |
1429 |
1.52 |
|
|
|
| 12 |
A |
1504 |
1420 |
2.18 |
|
|
|
| 13 |
A |
1437 |
1358 |
6.24 |
|
|
|
| 14 |
A |
1417 |
1338 |
11.30 |
|
|
|
| 15 |
A |
1393 |
1316 |
0.95 |
|
|
|
| 16 |
A |
1320 |
1246 |
4.42 |
|
|
|
| 17 |
A |
1306 |
1233 |
10.65 |
|
|
|
| 18 |
A |
1201 |
1135 |
12.47 |
|
|
|
| 19 |
A |
1133 |
1070 |
9.02 |
|
|
|
| 20 |
A |
1114 |
1052 |
31.40 |
|
|
|
| 21 |
A |
1021 |
964 |
9.58 |
|
|
|
| 22 |
A |
943 |
891 |
90.83 |
|
|
|
| 23 |
A |
900 |
850 |
1.66 |
|
|
|
| 24 |
A |
853 |
805 |
379.17 |
|
|
|
| 25 |
A |
790 |
746 |
29.53 |
|
|
|
| 26 |
A |
626 |
591 |
42.01 |
|
|
|
| 27 |
A |
459 |
433 |
9.74 |
|
|
|
| 28 |
A |
395 |
373 |
4.22 |
|
|
|
| 29 |
A |
290 |
274 |
0.89 |
|
|
|
| 30 |
A |
260 |
245 |
0.93 |
|
|
|
| 31 |
A |
189 |
178 |
0.20 |
|
|
|
| 32 |
A |
111 |
105 |
0.73 |
|
|
|
| 33 |
A |
62 |
58 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23516.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 22211.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.147 |
-0.930 |
0.184 |
| C2 |
-1.752 |
0.458 |
-0.309 |
| C3 |
-0.406 |
0.900 |
0.222 |
| O4 |
0.556 |
-0.030 |
-0.303 |
| N5 |
1.768 |
0.212 |
0.316 |
| O6 |
2.559 |
-0.553 |
-0.140 |
| H7 |
-3.124 |
-1.219 |
-0.205 |
| H8 |
-2.199 |
-0.955 |
1.275 |
| H9 |
-1.418 |
-1.674 |
-0.136 |
| H10 |
-1.722 |
0.480 |
-1.401 |
| H11 |
-2.489 |
1.202 |
0.006 |
| H12 |
-0.148 |
1.908 |
-0.110 |
| H13 |
-0.375 |
0.864 |
1.314 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5256 | 2.5261 | 2.8903 | 4.0799 | 4.7324 | 1.0905 | 1.0919 | 1.0897 | 2.1644 | 2.1672 | 3.4842 | 2.7634 |
C2 | 1.5256 | | 1.5132 | 2.3594 | 3.5837 | 4.4318 | 2.1690 | 2.1693 | 2.1656 | 1.0928 | 1.0936 | 2.1717 | 2.1672 | C3 | 2.5261 | 1.5132 | | 1.4373 | 2.2820 | 3.3217 | 3.4723 | 2.7861 | 2.7890 | 2.1314 | 2.1165 | 1.0925 | 1.0931 | O4 | 2.8903 | 2.3594 | 1.4373 | | 1.3822 | 2.0768 | 3.8683 | 3.3066 | 2.5746 | 2.5798 | 3.2998 | 2.0709 | 2.0694 | N5 | 4.0799 | 3.5837 | 2.2820 | 1.3822 | | 1.1913 | 5.1230 | 4.2442 | 3.7298 | 3.8987 | 4.3820 | 2.5938 | 2.4522 | O6 | 4.7324 | 4.4318 | 3.3217 | 2.0768 | 1.1913 | | 5.7223 | 4.9800 | 4.1324 | 4.5812 | 5.3471 | 3.6585 | 3.5683 | H7 | 1.0905 | 2.1690 | 3.4723 | 3.8683 | 5.1230 | 5.7223 | | 1.7649 | 1.7668 | 2.5067 | 2.5115 | 4.3178 | 3.7686 | H8 | 1.0919 | 2.1693 | 2.7861 | 3.3066 | 4.2442 | 4.9800 | 1.7649 | | 1.7656 | 3.0739 | 2.5196 | 3.7843 | 2.5763 | H9 | 1.0897 | 2.1656 | 2.7890 | 2.5746 | 3.7298 | 4.1324 | 1.7668 | 1.7656 | | 2.5170 | 3.0728 | 3.8010 | 3.1042 | H10 | 2.1644 | 1.0928 | 2.1314 | 2.5798 | 3.8987 | 4.5812 | 2.5067 | 3.0739 | 2.5170 | | 1.7579 | 2.4869 | 3.0554 | H11 | 2.1672 | 1.0936 | 2.1165 | 3.2998 | 4.3820 | 5.3471 | 2.5115 | 2.5196 | 3.0728 | 1.7579 | | 2.4486 | 2.5094 | H12 | 3.4842 | 2.1717 | 1.0925 | 2.0709 | 2.5938 | 3.6585 | 4.3178 | 3.7843 | 3.8010 | 2.4869 | 2.4486 | | 1.7802 | H13 | 2.7634 | 2.1672 | 1.0931 | 2.0694 | 2.4522 | 3.5683 | 3.7686 | 2.5763 | 3.1042 | 3.0554 | 2.5094 | 1.7802 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.462 |
|
C1 |
C2 |
H10 |
110.413 |
| C1 |
C2 |
H11 |
110.582 |
|
C2 |
C1 |
H7 |
110.917 |
| C2 |
C1 |
H8 |
110.853 |
|
C2 |
C1 |
H9 |
110.690 |
| C2 |
C3 |
O4 |
106.168 |
|
C2 |
C3 |
H12 |
111.896 |
| C2 |
C3 |
H13 |
111.497 |
|
C3 |
C2 |
H10 |
108.680 |
| C3 |
C2 |
H11 |
107.475 |
|
C3 |
O4 |
N5 |
108.049 |
| O4 |
C3 |
H12 |
109.136 |
|
O4 |
C3 |
H13 |
108.976 |
| O4 |
N5 |
O6 |
107.377 |
|
H7 |
C1 |
H8 |
107.938 |
| H7 |
C1 |
H9 |
108.271 |
|
H8 |
C1 |
H9 |
108.060 |
| H10 |
C2 |
H11 |
107.031 |
|
H12 |
C3 |
H13 |
109.076 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
