Jump to
S1C2
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -93.743903 |
| Energy at 298.15K | -93.745210 |
| HF Energy | -93.405829 |
| Nuclear repulsion energy | 28.124458 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3411 |
3232 |
3.20 |
|
|
|
| 2 |
A' |
3064 |
2903 |
39.42 |
|
|
|
| 3 |
A' |
1851 |
1753 |
24.21 |
|
|
|
| 4 |
A' |
1059 |
1003 |
130.35 |
|
|
|
| 5 |
A' |
877 |
831 |
109.73 |
|
|
|
| 6 |
A" |
930 |
881 |
5.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5595.7 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 5301.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.112 |
0.640 |
0.000 |
| N2 |
0.112 |
-0.592 |
0.000 |
| H3 |
-0.673 |
1.404 |
0.000 |
| H4 |
-0.780 |
-1.095 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2319 | 1.0957 | 1.9503 |
N2 | 1.2319 | | 2.1451 | 1.0235 | H3 | 1.0957 | 2.1451 | | 2.5012 | H4 | 1.9503 | 1.0235 | 2.5012 | |
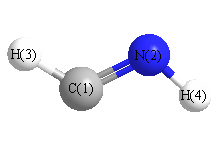 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
119.417 |
|
H3 |
C1 |
N2 |
134.233 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -93.751672 |
| Energy at 298.15K | -93.753027 |
| HF Energy | -93.411128 |
| Nuclear repulsion energy | 28.056157 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3518 |
3333 |
8.39 |
|
|
|
| 2 |
A' |
3108 |
2945 |
24.21 |
|
|
|
| 3 |
A' |
1806 |
1711 |
27.10 |
|
|
|
| 4 |
A' |
1230 |
1165 |
9.16 |
|
|
|
| 5 |
A' |
932 |
883 |
212.76 |
|
|
|
| 6 |
A" |
1008 |
955 |
93.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5800.8 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 5495.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.651 |
0.000 |
| N2 |
0.001 |
-0.590 |
0.000 |
| H3 |
0.901 |
1.274 |
0.000 |
| H4 |
-0.908 |
-1.050 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2414 | 1.0944 | 1.9290 |
N2 | 1.2414 | | 2.0700 | 1.0188 | H3 | 1.0944 | 2.0700 | | 2.9452 | H4 | 1.9290 | 1.0188 | 2.9452 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
116.838 |
|
H3 |
C1 |
N2 |
124.686 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
