return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-235.182860
Energy at 298.15K-235.195195
HF Energy-234.206867
Nuclear repulsion energy244.433325
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3295 3113 10.36      
2 A' 3202 3024 22.26      
3 A' 3200 3023 1.89      
4 A' 3196 3019 12.81      
5 A' 3190 3013 41.14      
6 A' 3100 2928 15.15      
7 A' 3098 2926 9.56      
8 A' 3096 2924 12.45      
9 A' 1714 1619 6.97      
10 A' 1541 1455 4.78      
11 A' 1535 1449 8.64      
12 A' 1520 1436 9.78      
13 A' 1472 1390 1.52      
14 A' 1433 1353 3.79      
15 A' 1424 1345 6.79      
16 A' 1354 1279 2.49      
17 A' 1329 1255 0.41      
18 A' 1205 1138 2.02      
19 A' 1121 1059 11.33      
20 A' 1014 958 1.62      
21 A' 992 937 0.67      
22 A' 924 872 1.57      
23 A' 749 708 1.23      
24 A' 520 491 1.19      
25 A' 439 414 0.80      
26 A' 328 310 0.24      
27 A' 285 269 0.18      
28 A' 269 254 0.28      
29 A" 3196 3018 7.00      
30 A" 3188 3011 3.41      
31 A" 3176 3000 12.98      
32 A" 3097 2925 16.78      
33 A" 1529 1444 0.36      
34 A" 1515 1431 0.89      
35 A" 1504 1421 7.65      
36 A" 1410 1331 5.43      
37 A" 1355 1280 1.89      
38 A" 1150 1086 1.86      
39 A" 1075 1015 0.14      
40 A" 982 927 0.26      
41 A" 936 884 13.89      
42 A" 930 878 21.34      
43 A" 755 713 0.47      
44 A" 549 518 5.95      
45 A" 277 261 0.01      
46 A" 211 199 0.30      
47 A" 174 164 0.17      
48 A" 29 27 0.09      

Unscaled Zero Point Vibrational Energy (zpe) 36789.6 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 34747.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.14850 0.08401 0.08300

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.752 -1.064 0.000
C2 -0.476 -0.661 0.000
C3 -0.126 0.808 0.000
C4 0.655 -1.654 0.000
C5 0.655 1.202 1.257
C6 0.655 1.202 -1.257
H7 -2.016 -2.115 0.000
H8 -2.568 -0.351 0.000
H9 -1.070 1.364 0.000
H10 0.274 -2.676 0.000
H11 1.293 -1.532 -0.878
H12 1.293 -1.532 0.878
H13 0.812 2.283 1.285
H14 0.117 0.912 2.162
H15 1.638 0.727 1.277
H16 0.812 2.283 -1.285
H17 0.117 0.912 -2.162
H18 1.638 0.727 -1.277

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33812.47922.47803.53683.53681.08351.08352.52172.58903.20393.20394.40773.47414.04074.40773.47414.0407
C21.33811.51021.50482.51642.51642.11762.11522.11042.14982.15882.15883.46102.73852.83293.46102.73852.8329
C32.47921.51022.58311.53191.53193.48032.70321.09533.50702.87472.87472.16982.17792.17932.16982.17792.1793
C42.47801.50482.58313.12113.12112.71003.47633.47621.09051.09261.09264.14463.39812.87534.14463.39812.8753
C53.53682.51641.53193.12112.51484.44043.79252.14064.09503.52812.83391.09231.09161.09202.76713.47342.7598
C63.53682.51641.53193.12112.51484.44043.79252.14064.09502.83393.52812.76713.47342.75981.09231.09161.0920
H71.08352.11763.48032.71004.44044.44041.84783.60492.35763.47303.47305.38424.28754.80155.38424.28754.8015
H81.08352.11522.70323.47633.79253.79251.84782.27743.67174.13264.13264.47433.67144.52584.47433.67144.5258
H92.52172.11041.09533.47622.14062.14063.60492.27744.25763.84003.84002.45762.50733.06082.45762.50733.0608
H102.58902.14983.50701.09054.09504.09502.35763.67174.25761.76601.76605.15104.19183.88225.15104.19183.8822
H113.20392.15882.87471.09263.52812.83393.47304.13263.84001.76601.75624.41224.07423.14143.86693.00102.3200
H123.20392.15882.87471.09262.83393.52813.47304.13263.84001.76601.75623.86693.00102.32004.41224.07423.1414
H134.40773.46102.16984.14461.09232.76715.38424.47432.45765.15104.41223.86691.76981.76122.57023.77423.1093
H143.47412.73852.17793.39811.09163.47344.28753.67142.50734.19184.07423.00101.76981.76893.77424.32363.7648
H154.04072.83292.17932.87531.09202.75984.80154.52583.06083.88223.14142.32001.76121.76893.10933.76482.5545
H164.40773.46102.16984.14462.76711.09235.38424.47432.45765.15103.86694.41222.57023.77423.10931.76981.7612
H173.47412.73852.17793.39813.47341.09164.28753.67142.50734.19183.00104.07423.77424.32363.76481.76981.7689
H184.04072.83292.17932.87532.75981.09204.80154.52583.06083.88222.32003.14143.10933.76482.55451.76121.7689

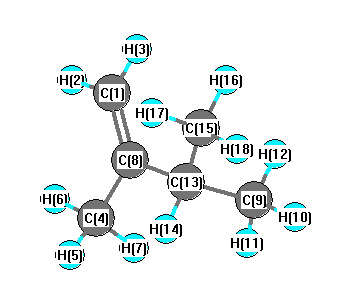
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.899 C1 C2 C4 121.192
C2 C1 H7 121.617 C2 C1 H8 121.370
C2 C3 C5 111.623 C2 C3 C6 111.623
C2 C3 H9 107.113 C2 C4 H10 110.852
C2 C4 H11 111.443 C2 C4 H12 111.443
C3 C2 C4 117.909 C3 C5 H13 110.432
C3 C5 H14 111.119 C3 C5 H15 111.206
C3 C6 H16 110.432 C3 C6 H17 111.119
C3 C6 H18 111.206 C5 C3 C6 110.334
C5 C3 H9 107.978 C6 C3 H9 107.978
H7 C1 H8 117.013 H10 C4 H11 107.979
H10 C4 H12 107.979 H11 C4 H12 106.961
H13 C5 H14 108.266 H13 C5 H15 107.477
H14 C5 H15 108.209 H16 C6 H17 108.266
H16 C6 H18 107.477 H17 C6 H18 108.209
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability