return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-398.138553
Energy at 298.15K-398.148676
HF Energy-396.766277
Nuclear repulsion energy326.638324
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3875 3671 61.46      
2 A 3828 3626 79.56      
3 A 3636 3445 4.92      
4 A 3546 3359 1.72      
5 A 3159 2993 18.23      
6 A 3105 2942 10.26      
7 A 3045 2885 38.90      
8 A 1845 1748 227.25      
9 A 1697 1608 33.01      
10 A 1536 1455 2.71      
11 A 1473 1395 42.60      
12 A 1442 1366 22.90      
13 A 1430 1355 50.47      
14 A 1408 1334 3.89      
15 A 1351 1280 9.76      
16 A 1269 1202 15.26      
17 A 1218 1154 17.69      
18 A 1199 1136 199.87      
19 A 1167 1106 17.02      
20 A 1131 1072 95.00      
21 A 1057 1002 37.51      
22 A 1018 965 1.70      
23 A 926 877 173.97      
24 A 834 790 16.23      
25 A 759 719 26.74      
26 A 637 603 92.61      
27 A 590 559 10.11      
28 A 552 523 190.99      
29 A 523 495 11.48      
30 A 449 426 6.71      
31 A 304 288 6.88      
32 A 293 277 20.94      
33 A 270 256 21.07      
34 A 222 211 2.69      
35 A 170 161 3.02      
36 A 38 36 1.00      

Unscaled Zero Point Vibrational Energy (zpe) 25501.0 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 24159.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
0.11948 0.07870 0.05208

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.782 -0.538 0.004
O2 2.065 -0.324 -0.339
O3 0.413 -1.569 0.515
C4 -0.094 0.667 -0.317
C5 -1.514 0.441 0.203
O6 -2.119 -0.693 -0.371
N7 0.431 1.924 0.210
H8 2.544 -1.128 -0.106
H9 -0.137 0.743 -1.408
H10 -1.490 0.380 1.300
H11 -2.115 1.310 -0.070
H12 -1.631 -1.447 -0.030
H13 1.361 2.085 -0.158
H14 0.537 1.851 1.217

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.34561.20861.52412.50392.92872.49541.86212.11662.77203.43682.57852.69152.6906
O21.34562.23862.37593.69994.20012.83270.96522.66993.97784.49593.87532.51623.0800
O31.20862.23862.43932.80232.82143.50692.26383.05732.83513.87582.11843.83543.4944
C41.52412.37592.43931.52862.43931.46093.19861.09522.15542.13472.62922.03872.0384
C52.50393.69992.80231.52861.40742.44614.36242.14111.09941.09101.90613.33202.6883
O62.92874.20012.82142.43931.40743.69974.69092.65822.08272.02510.96084.45824.0063
N72.49542.83273.50691.46092.44613.69973.72602.08292.69542.63433.95911.01341.0153
H81.86210.96522.26383.19864.36244.69093.72603.51934.53125.25894.18823.42483.8284
H92.11662.66993.05731.09522.14112.65822.08293.51933.04972.45472.98822.36872.9288
H102.77203.97782.83512.15541.09942.08272.69544.53123.04971.77022.26393.62872.5069
H113.43684.49593.87582.13471.09102.02512.63435.25892.45471.77022.79943.56302.9979
H122.57853.87532.11842.62921.90610.96083.95914.18822.98822.26392.79944.63134.1396
H132.69152.51623.83542.03873.33204.45821.01343.42482.36873.62873.56304.63131.6202
H142.69063.08003.49442.03842.68834.00631.01533.82842.92882.50692.99794.13961.6202

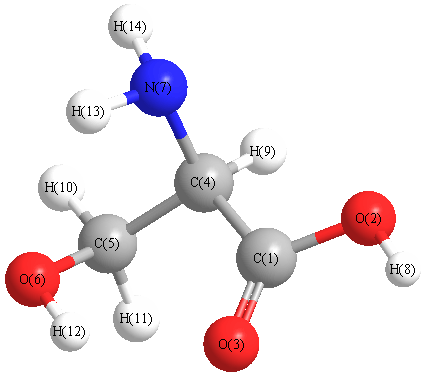
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 106.221 C1 C4 C5 110.215
C1 C4 N7 113.424 C1 C4 H9 106.677
O2 C1 O3 122.343 O2 C1 C4 111.626
O3 C1 C4 126.027 C4 C5 O6 112.302
C4 C5 H10 109.112 C4 C5 H11 107.991
C4 N7 H13 109.635 C4 N7 H14 109.498
C5 C4 N7 109.792 C5 C4 H9 108.242
C5 O6 H12 105.648 O6 C5 H10 111.783
O6 C5 H11 107.633 N7 C4 H9 108.303
H10 C5 H11 107.842 H13 N7 H14 106.002
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability