return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: CCD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/6-31G(2df,p)
 hartrees
Energy at 0K-398.125991
Energy at 298.15K-398.136138
HF Energy-396.767075
Nuclear repulsion energy326.969262
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3900 3694 62.27      
2 A 3854 3650 84.00      
3 A 3652 3459 6.05      
4 A 3561 3373 2.29      
5 A 3173 3005 16.68      
6 A 3116 2952 9.54      
7 A 3060 2898 36.18      
8 A 1889 1789 242.19      
9 A 1700 1611 34.38      
10 A 1540 1459 3.01      
11 A 1477 1399 38.45      
12 A 1451 1374 36.75      
13 A 1438 1362 46.37      
14 A 1413 1339 3.39      
15 A 1358 1286 9.73      
16 A 1275 1208 20.64      
17 A 1226 1161 38.30      
18 A 1209 1146 184.55      
19 A 1173 1111 12.48      
20 A 1141 1081 98.45      
21 A 1063 1007 35.63      
22 A 1022 968 1.87      
23 A 927 878 174.47      
24 A 840 796 17.69      
25 A 766 726 26.78      
26 A 639 605 93.88      
27 A 595 563 13.20      
28 A 548 519 195.48      
29 A 525 497 7.43      
30 A 452 428 6.54      
31 A 305 289 6.82      
32 A 294 278 20.11      
33 A 271 257 22.29      
34 A 222 211 2.56      
35 A 170 161 3.12      
36 A 40 38 1.01      

Unscaled Zero Point Vibrational Energy (zpe) 25642.7 cm-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 24288.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G(2df,p)
ABC
0.11981 0.07868 0.05218

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.777 -0.542 0.008
O2 2.048 -0.345 -0.365
O3 0.396 -1.548 0.549
C4 -0.087 0.671 -0.314
C5 -1.509 0.443 0.192
O6 -2.099 -0.692 -0.385
N7 0.439 1.917 0.226
H8 2.531 -1.142 -0.128
H9 -0.121 0.754 -1.405
H10 -1.494 0.387 1.289
H11 -2.107 1.310 -0.091
H12 -1.633 -1.445 -0.021
H13 1.347 2.118 -0.169
H14 0.560 1.843 1.229

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.33971.20341.52352.49552.90612.49171.85922.11722.76733.42842.57322.72622.6880
O21.33972.23872.36463.68544.16152.83790.96232.64423.97694.48053.85732.56773.0883
O31.20342.23872.42902.77822.79813.47972.27603.06322.80383.85232.10983.85423.4620
C41.52352.36462.42901.52652.43151.45593.19001.09482.15142.13032.63742.04182.0423
C52.49553.68542.77821.52651.40302.44304.35162.13841.09851.09051.90413.33052.7055
O62.90614.16152.79812.43151.40303.69054.65892.65422.08102.02380.95804.45144.0129
N72.49172.83793.47971.45592.44303.69053.72312.07972.68442.63593.95691.01071.0123
H81.85920.96232.27603.19004.35164.65893.72313.50134.53275.24644.17623.46873.8255
H92.11722.64423.06321.09482.13842.65422.07973.50133.04552.44543.00682.35352.9298
H102.76733.97692.80382.15141.09852.08102.68444.53273.04551.76922.25613.63222.5193
H113.42844.48053.85232.13031.09052.02382.63595.24642.44541.76922.79703.54763.0230
H122.57323.85732.10982.63741.90410.95803.95694.17623.00682.25612.79704.64724.1456
H132.72622.56773.85422.04183.33054.45141.01073.46872.35353.63223.54764.64721.6272
H142.68803.08833.46202.04232.70554.01291.01233.82552.92982.51933.02304.14561.6272

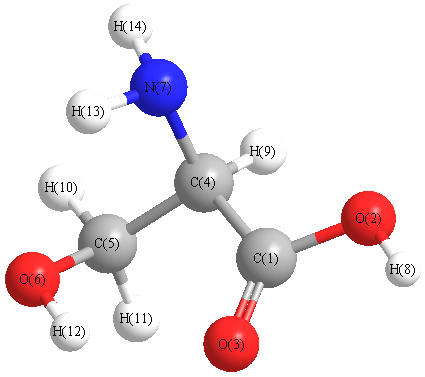
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 106.583 C1 C4 C5 109.808
C1 C4 N7 113.487 C1 C4 H9 106.775
O2 C1 O3 123.265 O2 C1 C4 111.188
O3 C1 C4 125.533 C4 C5 O6 112.128
C4 C5 H10 108.999 C4 C5 H11 107.815
C4 N7 H13 110.439 C4 N7 H14 110.382
C5 C4 N7 109.972 C5 C4 H9 108.202
C5 O6 H12 105.969 O6 C5 H10 112.019
O6 C5 H11 107.854 N7 C4 H9 108.413
H10 C5 H11 107.844 H13 N7 H14 107.090
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability