Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -550.428874 |
| Energy at 298.15K | |
| HF Energy | -548.971893 |
| Nuclear repulsion energy | 354.894361 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is D4h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
1.014 |
0.000 |
| C2 |
1.014 |
0.000 |
0.000 |
| C3 |
0.000 |
-1.014 |
0.000 |
| C4 |
-1.014 |
0.000 |
0.000 |
| F5 |
0.000 |
2.319 |
0.000 |
| F6 |
2.319 |
0.000 |
0.000 |
| F7 |
0.000 |
-2.319 |
0.000 |
| F8 |
-2.319 |
0.000 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4343 | 2.0284 | 1.4343 | 1.3048 | 2.5310 | 3.3331 | 2.5310 |
C2 | 1.4343 | | 1.4343 | 2.0284 | 2.5310 | 1.3048 | 2.5310 | 3.3331 | C3 | 2.0284 | 1.4343 | | 1.4343 | 3.3331 | 2.5310 | 1.3048 | 2.5310 | C4 | 1.4343 | 2.0284 | 1.4343 | | 2.5310 | 3.3331 | 2.5310 | 1.3048 | F5 | 1.3048 | 2.5310 | 3.3331 | 2.5310 | | 3.2795 | 4.6379 | 3.2795 | F6 | 2.5310 | 1.3048 | 2.5310 | 3.3331 | 3.2795 | | 3.2795 | 4.6379 | F7 | 3.3331 | 2.5310 | 1.3048 | 2.5310 | 4.6379 | 3.2795 | | 3.2795 | F8 | 2.5310 | 3.3331 | 2.5310 | 1.3048 | 3.2795 | 4.6379 | 3.2795 | |
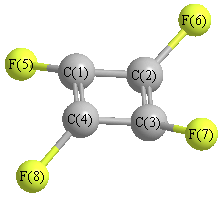 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
135.000 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.000 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
135.000 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.000 |
| C3 |
C2 |
F6 |
135.000 |
|
C3 |
C4 |
F8 |
135.000 |
| C4 |
C1 |
F5 |
135.000 |
|
C4 |
C3 |
F7 |
135.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -550.521183 |
| Energy at 298.15K | -550.521234 |
| HF Energy | -549.042146 |
| Nuclear repulsion energy | 355.050492 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1840 |
1738 |
0.00 |
|
|
|
| 2 |
Ag |
1227 |
1159 |
0.00 |
|
|
|
| 3 |
Ag |
714 |
674 |
0.00 |
|
|
|
| 4 |
Ag |
317 |
300 |
0.00 |
|
|
|
| 5 |
Ag |
186 |
176 |
0.00 |
|
|
|
| 6 |
Au |
1352 |
1277 |
468.09 |
|
|
|
| 7 |
Au |
936 |
884 |
36.41 |
|
|
|
| 8 |
Au |
581 |
549 |
58.28 |
|
|
|
| 9 |
Au |
243 |
230 |
0.03 |
|
|
|
| 10 |
Au |
157 |
148 |
0.05 |
|
|
|
| 11 |
Bg |
1386 |
1309 |
0.00 |
|
|
|
| 12 |
Bg |
767 |
725 |
0.00 |
|
|
|
| 13 |
Bg |
496 |
468 |
0.00 |
|
|
|
| 14 |
Bg |
442 |
417 |
0.00 |
|
|
|
| 15 |
Bu |
1796 |
1696 |
192.61 |
|
|
|
| 16 |
Bu |
999 |
944 |
154.96 |
|
|
|
| 17 |
Bu |
314 |
297 |
4.56 |
|
|
|
| 18 |
Bu |
186 |
176 |
2.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6968.8 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 6582.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.118 |
0.767 |
0.670 |
| C2 |
-0.118 |
-0.767 |
0.670 |
| C3 |
-0.118 |
-0.767 |
-0.670 |
| C4 |
0.118 |
0.767 |
-0.670 |
| F5 |
-0.118 |
1.654 |
1.604 |
| F6 |
0.118 |
-1.654 |
1.604 |
| F7 |
0.118 |
-1.654 |
-1.604 |
| F8 |
-0.118 |
1.654 |
-1.604 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5520 | 2.0503 | 1.3398 | 1.3097 | 2.5953 | 3.3217 | 2.4523 |
C2 | 1.5520 | | 1.3398 | 2.0503 | 2.5953 | 1.3097 | 2.4523 | 3.3217 | C3 | 2.0503 | 1.3398 | | 1.5520 | 3.3217 | 2.4523 | 1.3097 | 2.5953 | C4 | 1.3398 | 2.0503 | 1.5520 | | 2.4523 | 3.3217 | 2.5953 | 1.3097 | F5 | 1.3097 | 2.5953 | 3.3217 | 2.4523 | | 3.3168 | 4.6146 | 3.2083 | F6 | 2.5953 | 1.3097 | 2.4523 | 3.3217 | 3.3168 | | 3.2083 | 4.6146 | F7 | 3.3217 | 2.4523 | 1.3097 | 2.5953 | 4.6146 | 3.2083 | | 3.3168 | F8 | 2.4523 | 3.3217 | 2.5953 | 1.3097 | 3.2083 | 4.6146 | 3.3168 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
129.966 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.506 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
129.966 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.506 |
| C3 |
C2 |
F6 |
135.506 |
|
C3 |
C4 |
F8 |
129.966 |
| C4 |
C1 |
F5 |
135.506 |
|
C4 |
C3 |
F7 |
129.966 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
