return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (3-Pentanol)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-272.273898
Energy at 298.15K-272.287379
Nuclear repulsion energy259.742776
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3886 3671 26.30      
2 A 3221 3042 11.79      
3 A 3199 3021 18.58      
4 A 3194 3017 27.58      
5 A 3192 3015 23.92      
6 A 3163 2987 17.52      
7 A 3156 2980 0.49      
8 A 3108 2936 23.73      
9 A 3105 2932 30.92      
10 A 3101 2929 15.68      
11 A 3099 2927 0.56      
12 A 3026 2858 37.78      
13 A 1545 1460 5.70      
14 A 1540 1454 5.00      
15 A 1530 1445 5.76      
16 A 1527 1442 5.15      
17 A 1506 1422 2.72      
18 A 1496 1413 0.77      
19 A 1455 1374 6.84      
20 A 1429 1350 2.14      
21 A 1423 1344 4.08      
22 A 1416 1337 14.14      
23 A 1378 1301 0.33      
24 A 1345 1270 13.27      
25 A 1320 1246 1.95      
26 A 1297 1225 7.16      
27 A 1272 1201 24.97      
28 A 1196 1129 25.82      
29 A 1173 1108 3.05      
30 A 1118 1056 5.79      
31 A 1093 1033 23.57      
32 A 1057 998 3.20      
33 A 1039 982 1.22      
34 A 996 941 43.22      
35 A 956 903 9.13      
36 A 875 826 4.05      
37 A 793 749 1.75      
38 A 778 735 0.30      
39 A 497 470 2.72      
40 A 479 453 8.55      
41 A 403 380 4.25      
42 A 326 308 39.09      
43 A 304 287 32.50      
44 A 245 231 32.49      
45 A 224 212 10.25      
46 A 189 178 0.21      
47 A 114 108 1.57      
48 A 100 95 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 36940.3 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 34890.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.25081 0.06396 0.05563

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.008 0.014 0.223
C2 -1.271 -0.688 -0.248
C3 1.254 -0.730 -0.202
C4 -2.539 -0.001 0.250
C5 2.533 -0.014 0.223
O6 -0.050 1.322 -0.338
H7 -0.028 0.078 1.323
H8 -1.241 -1.728 0.089
H9 -1.256 -0.702 -1.342
H10 1.234 -0.845 -1.290
H11 1.228 -1.734 0.232
H12 -3.434 -0.501 -0.123
H13 -2.581 -0.010 1.342
H14 -2.558 1.036 -0.079
H15 3.418 -0.593 -0.041
H16 2.623 0.955 -0.271
H17 2.553 0.148 1.304
H18 0.637 1.839 0.088

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.51961.52522.53062.54111.42381.10202.13772.12552.13732.14123.48102.80612.76393.49002.83772.78291.9406
C21.51962.52571.52523.89182.35322.14481.09371.09432.71772.75222.17382.16882.15824.69534.22664.21063.1844
C31.52522.52573.88811.52622.43512.14922.70212.75641.09391.09434.69334.19644.20302.17522.17232.17402.6579
C42.53061.52523.88815.07152.87902.73162.16602.16184.16134.14661.09091.09231.08925.99365.27525.20133.6742
C52.54113.89181.52625.07152.96182.78804.14644.15642.16002.15865.99605.23515.20691.09061.09151.09302.6542
O61.42382.35322.43512.87902.96182.07513.30152.56092.69283.36163.84913.31692.53713.97362.69923.29380.9601
H71.10202.14482.14922.73162.78802.07512.50053.03623.04432.45953.74502.55543.04783.76633.21472.58132.2513
H82.13771.09372.70212.16604.14643.30152.50051.76102.96722.47332.52142.51343.06684.79704.71744.40264.0309
H92.12551.09432.75642.16184.15642.56093.03621.76102.49433.11712.50383.07252.51264.85334.35174.71463.4766
H102.13732.71771.09394.16132.16002.69283.04432.96722.49431.76294.82334.70954.40282.52902.49143.07433.0755
H112.14122.75221.09434.14662.15863.36162.45952.47333.11711.76294.83534.32654.70242.48443.07062.53843.6246
H123.48102.17384.69331.09095.99603.84913.74502.52142.50384.82334.83531.76441.77016.85316.23076.18804.7003
H132.80612.16884.19641.09235.23513.31692.55542.51343.07254.70954.32651.76441.76536.18475.53335.13663.9180
H142.76392.15824.20301.08925.20692.53713.04783.06682.51264.40284.70241.77011.76536.19495.18525.36873.2989
H153.49004.69532.17525.99361.09063.97363.76634.79704.85332.52902.48446.85316.18476.19491.75581.76273.6970
H162.83774.22662.17235.27521.09152.69923.21474.71744.35172.49143.07066.23075.53335.18521.75581.77092.2030
H172.78294.21062.17405.20131.09303.29382.58134.40264.71463.07432.53846.18805.13665.36871.76271.77092.8294
H181.94063.18442.65793.67422.65420.96012.25134.03093.47663.07553.62464.70033.91803.29893.69702.20302.8294

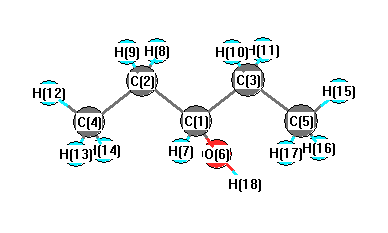
picture of 3-Pentanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.429 C1 C2 H8 108.671
C1 C2 H9 107.702 C1 C3 C5 112.768
C1 C3 H10 108.258 C1 C3 H11 108.536
C1 O6 H18 107.386 C2 C1 C3 112.096
C2 C1 O6 106.114 C2 C1 H7 108.752
C2 C4 H12 111.305 C2 C4 H13 110.824
C2 C4 H14 110.165 C3 C1 O6 111.281
C3 C1 H7 108.715 C3 C5 H15 111.358
C3 C5 H16 111.079 C3 C5 H17 111.116
C4 C2 H8 110.516 C4 C2 H9 110.150
C5 C3 H10 109.955 C5 C3 H11 109.818
O6 C1 H7 109.830 H8 C2 H9 107.191
H10 C3 H11 107.341 H12 C4 H13 107.832
H12 C4 H14 108.568 H13 C4 H14 108.041
H15 C5 H16 107.149 H15 C5 H17 107.647
H16 C5 H17 108.319
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability