Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -615.843447 |
| Energy at 298.15K | |
| HF Energy | -615.032648 |
| Nuclear repulsion energy | 206.196032 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.316 |
0.301 |
0.000 |
| C2 |
0.000 |
1.013 |
0.000 |
| C3 |
1.215 |
0.463 |
0.000 |
| C4 |
2.488 |
1.251 |
0.000 |
| Cl5 |
-1.189 |
-1.478 |
0.000 |
| H6 |
-1.901 |
0.569 |
0.882 |
| H7 |
-1.901 |
0.569 |
-0.882 |
| H8 |
-0.097 |
2.097 |
0.000 |
| H9 |
1.303 |
-0.619 |
0.000 |
| H10 |
2.295 |
2.324 |
0.000 |
| H11 |
3.094 |
1.012 |
0.877 |
| H12 |
3.094 |
1.012 |
-0.877 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4965 | 2.5366 | 3.9215 | 1.7837 | 1.0914 | 1.0914 | 2.1704 | 2.7760 | 4.1396 | 4.5529 | 4.5529 |
C2 | 1.4965 | | 1.3337 | 2.4997 | 2.7599 | 2.1421 | 2.1421 | 1.0884 | 2.0877 | 2.6434 | 3.2163 | 3.2163 | C3 | 2.5366 | 1.3337 | | 1.4975 | 3.0895 | 3.2402 | 3.2402 | 2.0958 | 1.0849 | 2.1521 | 2.1454 | 2.1454 | C4 | 3.9215 | 2.4997 | 1.4975 | | 4.5791 | 4.5288 | 4.5288 | 2.7206 | 2.2136 | 1.0907 | 1.0927 | 1.0927 | Cl5 | 1.7837 | 2.7599 | 3.0895 | 4.5791 | | 2.3398 | 2.3398 | 3.7377 | 2.6357 | 5.1570 | 5.0311 | 5.0311 | H6 | 1.0914 | 2.1421 | 3.2402 | 4.5288 | 2.3398 | | 1.7635 | 2.5230 | 3.5288 | 4.6332 | 5.0150 | 5.3145 | H7 | 1.0914 | 2.1421 | 3.2402 | 4.5288 | 2.3398 | 1.7635 | | 2.5230 | 3.5288 | 4.6332 | 5.3145 | 5.0150 | H8 | 2.1704 | 1.0884 | 2.0958 | 2.7206 | 3.7377 | 2.5230 | 2.5230 | | 3.0551 | 2.4032 | 3.4834 | 3.4834 | H9 | 2.7760 | 2.0877 | 1.0849 | 2.2136 | 2.6357 | 3.5288 | 3.5288 | 3.0551 | | 3.1056 | 2.5761 | 2.5761 | H10 | 4.1396 | 2.6434 | 2.1521 | 1.0907 | 5.1570 | 4.6332 | 4.6332 | 2.4032 | 3.1056 | | 1.7697 | 1.7697 | H11 | 4.5529 | 3.2163 | 2.1454 | 1.0927 | 5.0311 | 5.0150 | 5.3145 | 3.4834 | 2.5761 | 1.7697 | | 1.7547 | H12 | 4.5529 | 3.2163 | 2.1454 | 1.0927 | 5.0311 | 5.3145 | 5.0150 | 3.4834 | 2.5761 | 1.7697 | 1.7547 | |
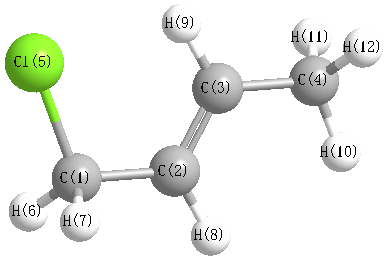 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.253 |
|
C1 |
C2 |
H8 |
113.263 |
| C2 |
C1 |
Cl5 |
114.291 |
|
C2 |
C1 |
H6 |
110.768 |
| C2 |
C1 |
H7 |
110.768 |
|
C2 |
C3 |
C4 |
123.896 |
| C2 |
C3 |
H9 |
118.996 |
|
C3 |
C2 |
H8 |
119.484 |
| C3 |
C4 |
H10 |
111.543 |
|
C3 |
C4 |
H11 |
110.873 |
| C3 |
C4 |
H12 |
110.873 |
|
C4 |
C3 |
H9 |
117.108 |
| Cl5 |
C1 |
H6 |
106.445 |
|
Cl5 |
C1 |
H7 |
106.445 |
| H6 |
C1 |
H7 |
107.794 |
|
H10 |
C4 |
H11 |
108.284 |
| H10 |
C4 |
H12 |
108.284 |
|
H11 |
C4 |
H12 |
106.811 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -615.845353 |
| Energy at 298.15K | -615.852443 |
| HF Energy | -615.035284 |
| Nuclear repulsion energy | 201.918158 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3227 |
3048 |
8.44 |
|
|
|
| 2 |
A |
3211 |
3033 |
5.89 |
|
|
|
| 3 |
A |
3211 |
3032 |
10.20 |
|
|
|
| 4 |
A |
3197 |
3019 |
2.37 |
|
|
|
| 5 |
A |
3176 |
3000 |
8.83 |
|
|
|
| 6 |
A |
3142 |
2968 |
10.65 |
|
|
|
| 7 |
A |
3098 |
2926 |
14.18 |
|
|
|
| 8 |
A |
1734 |
1638 |
7.38 |
|
|
|
| 9 |
A |
1520 |
1436 |
11.13 |
|
|
|
| 10 |
A |
1509 |
1425 |
6.55 |
|
|
|
| 11 |
A |
1505 |
1422 |
2.92 |
|
|
|
| 12 |
A |
1431 |
1352 |
2.32 |
|
|
|
| 13 |
A |
1354 |
1279 |
5.23 |
|
|
|
| 14 |
A |
1338 |
1264 |
1.33 |
|
|
|
| 15 |
A |
1294 |
1222 |
34.19 |
|
|
|
| 16 |
A |
1220 |
1152 |
1.83 |
|
|
|
| 17 |
A |
1136 |
1073 |
0.55 |
|
|
|
| 18 |
A |
1120 |
1058 |
2.04 |
|
|
|
| 19 |
A |
1075 |
1016 |
0.76 |
|
|
|
| 20 |
A |
1019 |
963 |
26.66 |
|
|
|
| 21 |
A |
962 |
909 |
9.44 |
|
|
|
| 22 |
A |
901 |
851 |
2.31 |
|
|
|
| 23 |
A |
813 |
768 |
19.03 |
|
|
|
| 24 |
A |
730 |
690 |
37.80 |
|
|
|
| 25 |
A |
494 |
467 |
1.40 |
|
|
|
| 26 |
A |
364 |
344 |
2.33 |
|
|
|
| 27 |
A |
277 |
262 |
1.90 |
|
|
|
| 28 |
A |
212 |
200 |
0.92 |
|
|
|
| 29 |
A |
160 |
152 |
2.00 |
|
|
|
| 30 |
A |
88 |
83 |
0.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22259.8 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 21024.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.783 |
0.831 |
0.105 |
| C2 |
0.510 |
0.186 |
0.457 |
| C3 |
1.588 |
0.238 |
-0.332 |
| C4 |
2.909 |
-0.385 |
-0.007 |
| Cl5 |
-2.078 |
-0.404 |
-0.102 |
| H6 |
-1.140 |
1.498 |
0.889 |
| H7 |
-0.717 |
1.380 |
-0.833 |
| H8 |
0.548 |
-0.346 |
1.403 |
| H9 |
1.515 |
0.767 |
-1.280 |
| H10 |
2.880 |
-0.894 |
0.957 |
| H11 |
3.193 |
-1.113 |
-0.771 |
| H12 |
3.700 |
0.367 |
0.024 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4873 | 2.4830 | 3.8886 | 1.8017 | 1.0897 | 1.0891 | 2.2002 | 2.6837 | 4.1373 | 4.5117 | 4.5080 |
C2 | 1.4873 | | 1.3369 | 2.5090 | 2.7125 | 2.1525 | 2.1439 | 1.0861 | 2.0887 | 2.6518 | 3.2238 | 3.2246 | C3 | 2.4830 | 1.3369 | | 1.4960 | 3.7291 | 3.2441 | 2.6211 | 2.1057 | 1.0878 | 2.1473 | 2.1431 | 2.1460 | C4 | 3.8886 | 2.5090 | 1.4960 | | 4.9877 | 4.5548 | 4.1166 | 2.7504 | 2.2113 | 1.0904 | 1.0928 | 1.0927 | Cl5 | 1.8017 | 2.7125 | 3.7291 | 4.9877 | | 2.3410 | 2.3600 | 3.0265 | 3.9585 | 5.0932 | 5.3604 | 5.8310 | H6 | 1.0897 | 2.1525 | 3.2441 | 4.5548 | 2.3410 | | 1.7776 | 2.5527 | 3.5054 | 4.6786 | 5.3247 | 5.0457 | H7 | 1.0891 | 2.1439 | 2.6211 | 4.1166 | 2.3600 | 1.7776 | | 3.0952 | 2.3575 | 4.6167 | 4.6376 | 4.6127 | H8 | 2.2002 | 1.0861 | 2.1057 | 2.7504 | 3.0265 | 2.5527 | 3.0952 | | 3.0612 | 2.4368 | 3.5089 | 3.5142 | H9 | 2.6837 | 2.0887 | 1.0878 | 2.2113 | 3.9585 | 3.5054 | 2.3575 | 3.0612 | | 3.1022 | 2.5707 | 2.5760 | H10 | 4.1373 | 2.6518 | 2.1473 | 1.0904 | 5.0932 | 4.6786 | 4.6167 | 2.4368 | 3.1022 | | 1.7700 | 1.7702 | H11 | 4.5117 | 3.2238 | 2.1431 | 1.0928 | 5.3604 | 5.3247 | 4.6376 | 3.5089 | 2.5707 | 1.7700 | | 1.7555 | H12 | 4.5080 | 3.2246 | 2.1460 | 1.0927 | 5.8310 | 5.0457 | 4.6127 | 3.5142 | 2.5760 | 1.7702 | 1.7555 | |
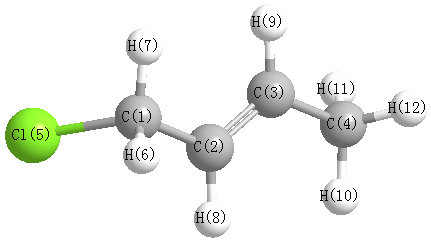 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.003 |
|
C1 |
C2 |
H8 |
116.654 |
| C2 |
C1 |
Cl5 |
110.759 |
|
C2 |
C1 |
H6 |
112.379 |
| C2 |
C1 |
H7 |
111.712 |
|
C2 |
C3 |
C4 |
124.577 |
| C2 |
C3 |
H9 |
118.600 |
|
C3 |
C2 |
H8 |
120.343 |
| C3 |
C4 |
H10 |
111.279 |
|
C3 |
C4 |
H11 |
110.797 |
| C3 |
C4 |
H12 |
111.027 |
|
C4 |
C3 |
H9 |
116.823 |
| Cl5 |
C1 |
H6 |
105.465 |
|
Cl5 |
C1 |
H7 |
106.847 |
| H6 |
C1 |
H7 |
109.350 |
|
H10 |
C4 |
H11 |
108.338 |
| H10 |
C4 |
H12 |
108.359 |
|
H11 |
C4 |
H12 |
106.884 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
