return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (hex-1-ene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-235.174557
Energy at 298.15K-235.186998
HF Energy-234.205431
Nuclear repulsion energy228.652817
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3306 3122 9.14      
2 A 3212 3034 3.27      
3 A 3200 3022 14.37      
4 A 3196 3019 22.74      
5 A 3195 3018 28.83      
6 A 3160 2985 35.20      
7 A 3144 2969 0.40      
8 A 3131 2957 0.94      
9 A 3103 2930 33.08      
10 A 3099 2927 32.88      
11 A 3089 2918 7.36      
12 A 3082 2911 4.50      
13 A 1708 1613 6.35      
14 A 1539 1454 4.52      
15 A 1531 1446 6.23      
16 A 1526 1441 1.45      
17 A 1514 1430 0.46      
18 A 1505 1422 2.03      
19 A 1468 1387 1.36      
20 A 1432 1352 2.23      
21 A 1407 1329 0.49      
22 A 1361 1286 0.84      
23 A 1347 1272 0.51      
24 A 1332 1258 0.17      
25 A 1327 1254 0.41      
26 A 1275 1204 2.49      
27 A 1262 1192 0.10      
28 A 1211 1144 0.29      
29 A 1142 1078 2.70      
30 A 1099 1038 0.19      
31 A 1064 1005 4.61      
32 A 1052 994 6.65      
33 A 1044 986 5.11      
34 A 957 904 0.65      
35 A 947 895 31.66      
36 A 932 881 1.88      
37 A 920 869 1.65      
38 A 804 759 0.70      
39 A 744 703 3.43      
40 A 661 624 8.68      
41 A 455 430 1.05      
42 A 361 341 0.05      
43 A 356 337 0.38      
44 A 256 242 0.00      
45 A 176 166 0.01      
46 A 131 124 0.05      
47 A 94 89 0.01      
48 A 75 71 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 36966.3 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 34914.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.43844 0.04218 0.04139

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 3.043 -0.235 -0.444
C2 2.027 -0.157 0.418
C3 0.770 0.624 0.186
C4 -0.464 -0.282 0.122
C5 -1.760 0.499 -0.075
C6 -2.982 -0.413 -0.142
H7 3.023 0.300 -1.386
H8 3.922 -0.831 -0.234
H9 2.081 -0.711 1.353
H10 0.630 1.351 0.994
H11 0.857 1.194 -0.745
H12 -0.337 -1.000 -0.695
H13 -0.530 -0.870 1.044
H14 -1.877 1.215 0.745
H15 -1.689 1.090 -0.993
H16 -3.902 0.157 -0.283
H17 -2.895 -1.119 -0.971
H18 -3.086 -0.993 0.777

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33502.51023.55254.87306.03541.08391.08222.09243.22582.62933.47473.92205.26574.94476.95786.02686.2950
C21.33501.49742.51093.87445.04632.11102.11491.08772.13442.13182.74522.72694.15074.16525.97795.20365.1927
C32.51021.49741.53202.54673.90672.76603.49682.20501.09611.09442.15362.15882.76972.76634.71864.22014.2230
C43.55252.51091.53201.52592.53563.84344.43432.85902.15012.16191.09491.09572.15082.15073.48952.79412.7947
C54.87303.87442.54671.52591.52624.96395.83784.27242.75342.78972.15762.15381.09461.09452.17872.16972.1699
C66.03545.04633.90672.53561.52626.17426.91735.28754.17744.20542.76502.76212.15812.15761.09121.09191.0919
H71.08392.11102.76603.84344.96396.17421.84763.06733.53472.42963.66894.46085.42164.79417.01386.10046.6085
H81.08222.11493.49684.43435.83786.91731.84762.43334.13603.70894.28734.63186.22715.97897.88606.86297.0821
H92.09241.08772.20502.85904.27245.28753.06732.43332.54673.08623.18162.63394.44364.79126.26275.50715.2063
H103.22582.13441.09612.15012.75344.17743.53474.13602.54671.76013.05192.50682.52353.06474.85744.73164.3990
H112.62932.13181.09442.16192.78974.20542.42963.70893.08621.76012.49843.06343.11392.56014.89234.41364.7588
H123.47472.74522.15361.09492.15762.76503.66894.28733.18163.05192.49841.75403.05772.50663.77012.57563.1178
H133.92202.72692.15881.09572.15382.76214.46084.63182.63392.50683.06341.75402.50073.05533.76653.11702.5727
H145.26574.15072.76972.15081.09462.15815.42166.22714.44362.52353.11393.05772.50071.75252.50493.07012.5173
H154.94474.16522.76632.15071.09452.15764.79415.97894.79123.06472.56012.50663.05531.75252.50422.51673.0698
H166.95785.97794.71863.48952.17871.09127.01387.88606.26274.85744.89233.77013.76652.50492.50421.76441.7644
H176.02685.20364.22012.79412.16971.09196.10046.86295.50714.73164.41362.57563.11703.07012.51671.76441.7628
H186.29505.19274.22302.79472.16991.09196.60857.08215.20634.39904.75883.11782.57272.51733.06981.76441.7628

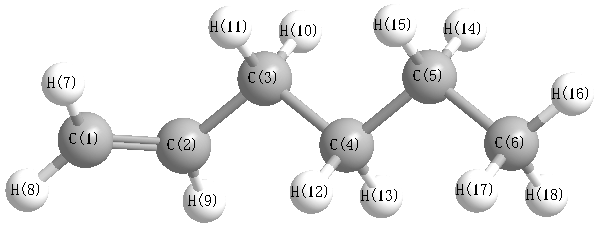
picture of hex-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.708 C1 C2 H9 119.110
C2 C1 H7 121.203 C2 C1 H8 121.720
C2 C3 C4 111.952 C2 C3 H10 109.797
C2 C3 H11 109.692 C3 C2 H9 116.165
C3 C4 C5 112.779 C3 C4 H12 109.003
C3 C4 H13 109.356 C4 C3 H10 108.658
C4 C3 H11 109.677 C4 C5 C6 112.355
C4 C5 H14 109.217 C4 C5 H15 109.211
C5 C4 H12 109.728 C5 C4 H13 109.384
C5 C6 H16 111.618 C5 C6 H17 110.855
C5 C6 H18 110.865 C6 C5 H14 109.773
C6 C5 H15 109.737 H7 C1 H8 117.077
H10 C3 H11 106.931 H12 C4 H13 106.389
H14 C5 H15 106.365 H16 C6 H17 107.850
H16 C6 H18 107.843 H17 C6 H18 107.647
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability