return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 ((Z)-hexa-2,3,4-triene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-232.720538
Energy at 298.15K-232.726830
HF Energy-231.796993
Nuclear repulsion energy190.687405
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3216 3038 11.11      
2 A1 3192 3015 4.64      
3 A1 3094 2922 9.17      
4 A1 2198 2076 0.75      
5 A1 1522 1438 5.57      
6 A1 1420 1342 4.82      
7 A1 1393 1316 0.00      
8 A1 1139 1076 1.42      
9 A1 1081 1021 1.14      
10 A1 767 725 0.37      
11 A1 472 446 6.33      
12 A1 93 88 1.10      
13 A2 3169 2993 0.00      
14 A2 1509 1425 0.00      
15 A2 1057 999 0.00      
16 A2 860 812 0.00      
17 A2 541 511 0.00      
18 A2 230 217 0.00      
19 A2 156 147 0.00      
20 B1 3169 2993 19.67      
21 B1 1510 1426 10.37      
22 B1 1054 996 0.39      
23 B1 708 668 34.77      
24 B1 239 226 1.63      
25 B1 135 128 2.13      
26 B2 3216 3038 5.87      
27 B2 3192 3015 0.21      
28 B2 3094 2922 30.58      
29 B2 1713 1618 6.87      
30 B2 1511 1427 11.10      
31 B2 1417 1339 9.60      
32 B2 1290 1219 14.48      
33 B2 1111 1049 1.77      
34 B2 953 900 28.66      
35 B2 543 513 2.94      
36 B2 197 186 0.35      

Unscaled Zero Point Vibrational Energy (zpe) 26079.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 24632.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.40996 0.04690 0.04275

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.000 2.147 1.629
H2 0.879 3.431 0.779
H3 -0.879 3.431 0.779
H4 0.000 -2.147 1.629
H5 0.879 -3.431 0.779
H6 -0.879 -3.431 0.779
C7 0.000 2.782 0.744
C8 0.000 -2.782 0.744
H9 0.000 2.487 -1.459
H10 0.000 -2.487 -1.459
C11 0.000 -0.634 -0.522
C12 0.000 0.634 -0.522
C13 0.000 1.955 -0.510
C14 0.000 -1.955 -0.510

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 H9 H10 C11 C12 C13 C14
H11.77301.77304.29415.71035.71031.08925.00773.10675.56883.51582.62972.14784.6265
H21.77301.75825.71036.86167.08331.09326.27472.58306.38784.35763.20732.14765.6075
H31.77301.75825.71037.08336.86161.09326.27472.58306.38784.35763.20732.14765.6075
H44.29415.71035.71031.77301.77305.00771.08925.56883.10672.62973.51584.62652.1478
H55.71036.86167.08331.77301.75826.27471.09326.38782.58303.20734.35765.60752.1476
H65.71037.08336.86161.77301.75826.27471.09326.38782.58303.20734.35765.60752.1476
C71.08921.09321.09325.00776.27476.27475.56372.22265.71103.64292.49301.50204.9003
C85.00776.27476.27471.08921.09321.09325.56375.71102.22262.49303.64294.90031.5020
H93.10672.58302.58305.56886.38786.38782.22265.71104.97443.25892.07661.08774.5426
H105.56886.38786.38783.10672.58302.58305.71102.22264.97442.07663.25894.54261.0877
C113.51584.35764.35762.62973.20733.20733.64292.49303.25892.07661.26822.58941.3212
C122.62973.20733.20733.51584.35764.35762.49303.64292.07663.25891.26821.32122.5894
C132.14782.14762.14764.62655.60755.60751.50204.90031.08774.54262.58941.32123.9105
C144.62655.60755.60752.14782.14762.14764.90031.50204.54261.08771.32122.58943.9105

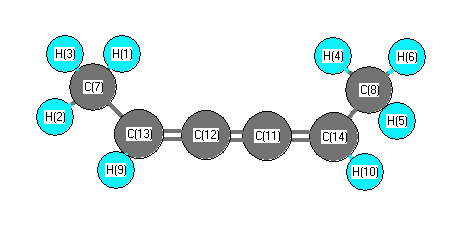
picture of (Z)-hexa-2,3,4-triene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C7 H2 108.657 H1 C7 H3 108.657
H1 C7 C13 110.961 H2 C7 H3 107.049
H2 C7 C13 110.700 H3 C7 C13 110.700
H4 C8 H5 108.657 H4 C8 H6 108.657
H4 C8 C14 110.961 H5 C8 H6 107.049
H5 C8 C14 110.700 H6 C8 C14 110.700
C7 C13 H9 117.334 C7 C13 C12 123.896
C8 C14 H10 117.334 C8 C14 C11 123.896
H9 C13 C12 118.770 H10 C14 C11 118.770
C11 C12 C13 179.494 C12 C11 C14 179.494
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability