Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -190.210403 |
| Energy at 298.15K | |
| HF Energy | -189.556057 |
| Nuclear repulsion energy | 88.241530 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3139 |
2965 |
20.37 |
175.48 |
0.11 |
0.20 |
| 2 |
A1 |
2351 |
2220 |
696.65 |
31.83 |
0.33 |
0.49 |
| 3 |
A1 |
1790 |
1691 |
8.46 |
10.33 |
0.36 |
0.53 |
| 4 |
A1 |
1501 |
1418 |
3.47 |
18.07 |
0.49 |
0.66 |
| 5 |
A1 |
922 |
871 |
2.44 |
36.07 |
0.25 |
0.40 |
| 6 |
B1 |
1021 |
964 |
20.86 |
0.01 |
0.75 |
0.86 |
| 7 |
B1 |
640 |
604 |
16.57 |
0.38 |
0.75 |
0.86 |
| 8 |
B1 |
219 |
207 |
0.21 |
2.78 |
0.75 |
0.86 |
| 9 |
B2 |
3219 |
3040 |
6.44 |
105.16 |
0.75 |
0.86 |
| 10 |
B2 |
1079 |
1019 |
1.28 |
0.28 |
0.75 |
0.86 |
| 11 |
B2 |
460 |
435 |
10.03 |
1.48 |
0.75 |
0.86 |
| 12 |
B2 |
188i |
177i |
15.24 |
0.23 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8076.1 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 7627.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.877 |
| C2 |
0.000 |
0.000 |
-0.557 |
| C3 |
0.000 |
0.000 |
0.720 |
| O4 |
0.000 |
0.000 |
1.899 |
| H5 |
0.000 |
0.922 |
-2.456 |
| H6 |
0.000 |
-0.922 |
-2.456 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3192 | 2.5963 | 3.7760 | 1.0890 | 1.0890 |
C2 | 1.3192 | | 1.2772 | 2.4568 | 2.1105 | 2.1105 | C3 | 2.5963 | 1.2772 | | 1.1797 | 3.3067 | 3.3067 | O4 | 3.7760 | 2.4568 | 1.1797 | | 4.4518 | 4.4518 | H5 | 1.0890 | 2.1105 | 3.3067 | 4.4518 | | 1.8443 | H6 | 1.0890 | 2.1105 | 3.3067 | 4.4518 | 1.8443 | |
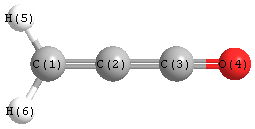 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.135 |
| C2 |
C1 |
H6 |
122.135 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.730 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -190.212395 |
| Energy at 298.15K | |
| HF Energy | -189.554635 |
| Nuclear repulsion energy | 88.498571 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3240 |
3060 |
5.33 |
91.79 |
0.69 |
0.81 |
| 2 |
A' |
3152 |
2977 |
18.58 |
129.92 |
0.11 |
0.21 |
| 3 |
A' |
2252 |
2127 |
663.55 |
42.95 |
0.35 |
0.52 |
| 4 |
A' |
1745 |
1648 |
4.99 |
3.55 |
0.50 |
0.67 |
| 5 |
A' |
1500 |
1417 |
1.14 |
15.97 |
0.53 |
0.69 |
| 6 |
A' |
1086 |
1026 |
19.04 |
4.15 |
0.09 |
0.16 |
| 7 |
A' |
949 |
896 |
1.64 |
32.58 |
0.20 |
0.33 |
| 8 |
A' |
530 |
501 |
12.60 |
3.58 |
0.69 |
0.81 |
| 9 |
A' |
178 |
168 |
18.22 |
3.88 |
0.75 |
0.86 |
| 10 |
A" |
1028 |
971 |
21.13 |
0.08 |
0.75 |
0.86 |
| 11 |
A" |
720 |
680 |
10.00 |
0.72 |
0.75 |
0.86 |
| 12 |
A" |
287 |
271 |
2.76 |
4.36 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8333.6 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 7871.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.063 |
-1.471 |
0.000 |
| C2 |
0.000 |
-0.670 |
0.000 |
| C3 |
-0.366 |
0.582 |
0.000 |
| O4 |
-0.902 |
1.626 |
0.000 |
| H5 |
2.087 |
-1.100 |
0.000 |
| H6 |
0.951 |
-2.552 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3310 | 2.5013 | 3.6677 | 1.0886 | 1.0867 |
C2 | 1.3310 | | 1.3042 | 2.4667 | 2.1305 | 2.1084 | C3 | 2.5013 | 1.3042 | | 1.1736 | 2.9742 | 3.3991 | O4 | 3.6677 | 2.4667 | 1.1736 | | 4.0452 | 4.5702 | H5 | 1.0886 | 2.1305 | 2.9742 | 4.0452 | | 1.8432 | H6 | 1.0867 | 2.1084 | 3.3991 | 4.5702 | 1.8432 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
143.316 |
|
C2 |
C1 |
H5 |
123.089 |
| C2 |
C1 |
H6 |
121.068 |
|
C2 |
C3 |
O4 |
169.153 |
| H5 |
C1 |
H6 |
115.842 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
